Jump to
S1C2
Energy calculated at mPW1PW91/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -209.158431 |
| Energy at 298.15K | -209.164508 |
| Nuclear repulsion energy | 118.012007 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3887 |
3728 |
95.68 |
|
|
|
| 2 |
A |
3158 |
3029 |
11.01 |
|
|
|
| 3 |
A |
3112 |
2985 |
14.10 |
|
|
|
| 4 |
A |
3049 |
2925 |
17.02 |
|
|
|
| 5 |
A |
1764 |
1692 |
0.97 |
|
|
|
| 6 |
A |
1483 |
1423 |
12.88 |
|
|
|
| 7 |
A |
1453 |
1394 |
9.76 |
|
|
|
| 8 |
A |
1401 |
1344 |
23.65 |
|
|
|
| 9 |
A |
1297 |
1244 |
52.81 |
|
|
|
| 10 |
A |
1161 |
1113 |
3.76 |
|
|
|
| 11 |
A |
1045 |
1002 |
162.47 |
|
|
|
| 12 |
A |
923 |
885 |
7.18 |
|
|
|
| 13 |
A |
574 |
550 |
13.49 |
|
|
|
| 14 |
A |
327 |
314 |
3.32 |
|
|
|
| 15 |
A |
3104 |
2977 |
12.82 |
|
|
|
| 16 |
A |
1477 |
1417 |
8.69 |
|
|
|
| 17 |
A |
1084 |
1040 |
1.18 |
|
|
|
| 18 |
A |
922 |
884 |
9.20 |
|
|
|
| 19 |
A |
417 |
400 |
128.79 |
|
|
|
| 20 |
A |
293 |
281 |
0.44 |
|
|
|
| 21 |
A |
208 |
200 |
0.32 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16068.3 cm
-1
Scaled (by 0.9592) Zero Point Vibrational Energy (zpe) 15412.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.290 |
1.297 |
0.000 |
| C2 |
0.000 |
0.561 |
0.000 |
| N3 |
0.010 |
-0.704 |
0.000 |
| O4 |
1.303 |
-1.208 |
0.000 |
| H5 |
1.162 |
-2.156 |
0.000 |
| H6 |
-2.124 |
0.599 |
0.000 |
| H7 |
-1.366 |
1.942 |
0.878 |
| H8 |
-1.366 |
1.942 |
-0.878 |
| H9 |
0.939 |
1.114 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.4846 | 2.3854 | 3.6047 | 4.2343 | 1.0876 | 1.0920 | 1.0920 | 2.2360 |
C2 | 1.4846 | | 1.2650 | 2.1970 | 2.9549 | 2.1243 | 2.1314 | 2.1314 | 1.0899 | N3 | 2.3854 | 1.2650 | | 1.3882 | 1.8537 | 2.4998 | 3.1084 | 3.1084 | 2.0420 | O4 | 3.6047 | 2.1970 | 1.3882 | | 0.9582 | 3.8741 | 4.2207 | 4.2207 | 2.3505 | H5 | 4.2343 | 2.9549 | 1.8537 | 0.9582 | | 4.2879 | 4.8940 | 4.8940 | 3.2776 | H6 | 1.0876 | 2.1243 | 2.4998 | 3.8741 | 4.2879 | | 1.7743 | 1.7743 | 3.1060 | H7 | 1.0920 | 2.1314 | 3.1084 | 4.2207 | 4.8940 | 1.7743 | | 1.7553 | 2.6015 | H8 | 1.0920 | 2.1314 | 3.1084 | 4.2207 | 4.8940 | 1.7743 | 1.7553 | | 2.6015 | H9 | 2.2360 | 1.0899 | 2.0420 | 2.3505 | 3.2776 | 3.1060 | 2.6015 | 2.6015 | |
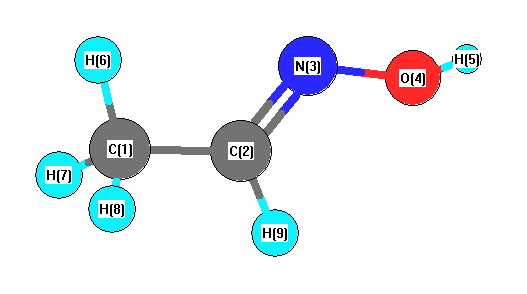 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
120.134 |
|
C1 |
C2 |
H9 |
119.792 |
| C2 |
C1 |
H6 |
110.398 |
|
C2 |
C1 |
H7 |
110.701 |
| C2 |
C1 |
H8 |
110.701 |
|
C2 |
N3 |
O4 |
111.720 |
| N3 |
C2 |
H9 |
120.075 |
|
N3 |
O4 |
H5 |
102.832 |
| H6 |
C1 |
H7 |
108.991 |
|
H6 |
C1 |
H8 |
108.991 |
| H7 |
C1 |
H8 |
106.972 |
|
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.226 |
0.000 |
|
|
| 2 |
C |
-0.023 |
0.000 |
|
|
| 3 |
N |
-0.063 |
0.000 |
|
|
| 4 |
O |
-0.241 |
0.000 |
|
|
| 5 |
H |
0.210 |
0.000 |
|
|
| 6 |
H |
0.097 |
0.000 |
|
|
| 7 |
H |
0.090 |
0.000 |
|
|
| 8 |
H |
0.090 |
0.000 |
|
|
| 9 |
H |
0.066 |
0.000 |
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.329 |
0.559 |
0.000 |
0.648 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.122 |
-2.095 |
0.000 |
| y |
-2.095 |
-18.679 |
0.000 |
| z |
0.000 |
0.000 |
-24.996 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.285 |
-2.095 |
0.000 |
| y |
-2.095 |
6.381 |
0.000 |
| z |
0.000 |
0.000 |
-3.096 |
|
| Polar |
|---|
| 3z2-r2 | -6.191 |
|---|
| x2-y2 | -6.444 |
|---|
| xy | -2.095 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.793 |
-1.387 |
0.000 |
| y |
-1.387 |
7.211 |
0.000 |
| z |
0.000 |
0.000 |
3.805 |
<r2> (average value of r
2) Å
2
| <r2> |
90.805 |
| (<r2>)1/2 |
9.529 |
Jump to
S1C1
Energy calculated at mPW1PW91/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -209.159196 |
| Energy at 298.15K | -209.165191 |
| HF Energy | -209.159196 |
| Nuclear repulsion energy | 120.495479 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3892 |
3733 |
96.46 |
|
|
|
| 2 |
A' |
3187 |
3057 |
7.20 |
|
|
|
| 3 |
A' |
3167 |
3038 |
5.25 |
|
|
|
| 4 |
A' |
3057 |
2932 |
11.19 |
|
|
|
| 5 |
A' |
1768 |
1696 |
8.16 |
|
|
|
| 6 |
A' |
1480 |
1420 |
19.40 |
|
|
|
| 7 |
A' |
1407 |
1350 |
25.52 |
|
|
|
| 8 |
A' |
1390 |
1333 |
30.78 |
|
|
|
| 9 |
A' |
1349 |
1294 |
36.75 |
|
|
|
| 10 |
A' |
1152 |
1105 |
10.07 |
|
|
|
| 11 |
A' |
969 |
929 |
149.36 |
|
|
|
| 12 |
A' |
928 |
890 |
2.00 |
|
|
|
| 13 |
A' |
683 |
655 |
12.60 |
|
|
|
| 14 |
A' |
313 |
300 |
1.35 |
|
|
|
| 15 |
A" |
3109 |
2982 |
10.40 |
|
|
|
| 16 |
A" |
1485 |
1425 |
9.39 |
|
|
|
| 17 |
A" |
1067 |
1024 |
0.62 |
|
|
|
| 18 |
A" |
873 |
837 |
14.15 |
|
|
|
| 19 |
A" |
515 |
494 |
35.59 |
|
|
|
| 20 |
A" |
403 |
386 |
81.04 |
|
|
|
| 21 |
A" |
32 |
31 |
0.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16112.4 cm
-1
Scaled (by 0.9592) Zero Point Vibrational Energy (zpe) 15455.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.441 |
0.487 |
0.000 |
| C2 |
0.000 |
0.855 |
0.000 |
| N3 |
1.007 |
0.084 |
0.000 |
| O4 |
0.658 |
-1.261 |
0.000 |
| H5 |
1.513 |
-1.694 |
0.000 |
| H6 |
-1.579 |
-0.589 |
0.000 |
| H7 |
-1.936 |
0.910 |
0.877 |
| H8 |
-1.936 |
0.910 |
-0.877 |
| H9 |
0.271 |
1.906 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.4877 | 2.4812 | 2.7318 | 3.6718 | 1.0850 | 1.0917 | 1.0917 | 2.2240 |
C2 | 1.4877 | | 1.2680 | 2.2156 | 2.9638 | 2.1397 | 2.1256 | 2.1256 | 1.0851 | N3 | 2.4812 | 1.2680 | | 1.3894 | 1.8486 | 2.6719 | 3.1795 | 3.1795 | 1.9642 | O4 | 2.7318 | 2.2156 | 1.3894 | | 0.9579 | 2.3356 | 3.4942 | 3.4942 | 3.1898 | H5 | 3.6718 | 2.9638 | 1.8486 | 0.9579 | | 3.2828 | 4.4091 | 4.4091 | 3.8074 | H6 | 1.0850 | 2.1397 | 2.6719 | 2.3356 | 3.2828 | | 1.7732 | 1.7732 | 3.1060 | H7 | 1.0917 | 2.1256 | 3.1795 | 3.4942 | 4.4091 | 1.7732 | | 1.7532 | 2.5749 | H8 | 1.0917 | 2.1256 | 3.1795 | 3.4942 | 4.4091 | 1.7732 | 1.7532 | | 2.5749 | H9 | 2.2240 | 1.0851 | 1.9642 | 3.1898 | 3.8074 | 3.1060 | 2.5749 | 2.5749 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
128.247 |
|
C1 |
C2 |
H9 |
118.806 |
| C2 |
C1 |
H6 |
111.594 |
|
C2 |
C1 |
H7 |
110.041 |
| C2 |
C1 |
H8 |
110.041 |
|
C2 |
N3 |
O4 |
112.900 |
| N3 |
C2 |
H9 |
112.947 |
|
N3 |
O4 |
H5 |
102.351 |
| H6 |
C1 |
H7 |
109.105 |
|
H6 |
C1 |
H8 |
109.105 |
| H7 |
C1 |
H8 |
106.828 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.249 |
|
|
|
| 2 |
C |
-0.071 |
|
|
|
| 3 |
N |
-0.067 |
|
|
|
| 4 |
O |
-0.239 |
|
|
|
| 5 |
H |
0.209 |
|
|
|
| 6 |
H |
0.109 |
|
|
|
| 7 |
H |
0.096 |
|
|
|
| 8 |
H |
0.096 |
|
|
|
| 9 |
H |
0.116 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.714 |
0.157 |
0.000 |
0.731 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-22.877 |
-3.856 |
0.000 |
| y |
-3.856 |
-21.061 |
0.000 |
| z |
0.000 |
0.000 |
-25.011 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.159 |
-3.856 |
0.000 |
| y |
-3.856 |
2.883 |
0.000 |
| z |
0.000 |
0.000 |
-3.042 |
|
| Polar |
|---|
| 3z2-r2 | -6.083 |
|---|
| x2-y2 | -1.816 |
|---|
| xy | -3.856 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.224 |
-1.174 |
0.000 |
| y |
-1.174 |
6.280 |
0.000 |
| z |
0.000 |
0.000 |
3.800 |
<r2> (average value of r
2) Å
2
| <r2> |
78.345 |
| (<r2>)1/2 |
8.851 |
