Jump to
S1C2
Vibrational Frequencies calculated at B1B95/cc-pVTZ
Geometric Data calculated at B1B95/cc-pVTZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -271.734197 |
| Energy at 298.15K | -271.744651 |
| Nuclear repulsion energy | 237.385462 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3178 |
3042 |
9.44 |
|
|
|
| 2 |
A |
3128 |
2994 |
34.81 |
|
|
|
| 3 |
A |
3126 |
2992 |
32.41 |
|
|
|
| 4 |
A |
3125 |
2991 |
29.64 |
|
|
|
| 5 |
A |
3100 |
2967 |
0.66 |
|
|
|
| 6 |
A |
3076 |
2944 |
17.06 |
|
|
|
| 7 |
A |
3062 |
2930 |
7.13 |
|
|
|
| 8 |
A |
3062 |
2930 |
4.85 |
|
|
|
| 9 |
A |
3048 |
2917 |
28.27 |
|
|
|
| 10 |
A |
3035 |
2904 |
14.35 |
|
|
|
| 11 |
A |
1826 |
1748 |
155.52 |
|
|
|
| 12 |
A |
1507 |
1443 |
6.51 |
|
|
|
| 13 |
A |
1500 |
1435 |
6.88 |
|
|
|
| 14 |
A |
1490 |
1426 |
2.06 |
|
|
|
| 15 |
A |
1475 |
1411 |
10.69 |
|
|
|
| 16 |
A |
1464 |
1401 |
20.00 |
|
|
|
| 17 |
A |
1448 |
1386 |
2.65 |
|
|
|
| 18 |
A |
1414 |
1353 |
14.93 |
|
|
|
| 19 |
A |
1409 |
1348 |
33.94 |
|
|
|
| 20 |
A |
1383 |
1324 |
36.61 |
|
|
|
| 21 |
A |
1335 |
1278 |
0.09 |
|
|
|
| 22 |
A |
1330 |
1273 |
17.82 |
|
|
|
| 23 |
A |
1252 |
1198 |
0.04 |
|
|
|
| 24 |
A |
1189 |
1138 |
57.24 |
|
|
|
| 25 |
A |
1141 |
1092 |
0.91 |
|
|
|
| 26 |
A |
1140 |
1091 |
0.04 |
|
|
|
| 27 |
A |
1072 |
1026 |
0.21 |
|
|
|
| 28 |
A |
975 |
933 |
0.84 |
|
|
|
| 29 |
A |
962 |
921 |
3.59 |
|
|
|
| 30 |
A |
919 |
879 |
9.67 |
|
|
|
| 31 |
A |
834 |
798 |
1.61 |
|
|
|
| 32 |
A |
833 |
798 |
0.72 |
|
|
|
| 33 |
A |
720 |
690 |
3.89 |
|
|
|
| 34 |
A |
599 |
573 |
12.47 |
|
|
|
| 35 |
A |
476 |
455 |
0.04 |
|
|
|
| 36 |
A |
387 |
370 |
1.47 |
|
|
|
| 37 |
A |
340 |
325 |
2.78 |
|
|
|
| 38 |
A |
245 |
234 |
0.03 |
|
|
|
| 39 |
A |
176 |
169 |
4.43 |
|
|
|
| 40 |
A |
105 |
100 |
0.74 |
|
|
|
| 41 |
A |
102 |
97 |
0.27 |
|
|
|
| 42 |
A |
29 |
28 |
0.26 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31006.0 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 29675.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-2.731 |
-0.289 |
0.000 |
| C2 |
-1.392 |
0.425 |
0.001 |
| C3 |
-0.223 |
-0.536 |
-0.001 |
| C4 |
1.125 |
0.143 |
-0.000 |
| C5 |
2.331 |
-0.758 |
0.000 |
| O6 |
1.231 |
1.343 |
-0.000 |
| H7 |
-3.556 |
0.420 |
0.001 |
| H8 |
-2.842 |
-0.923 |
-0.879 |
| H9 |
-2.841 |
-0.925 |
0.878 |
| H10 |
-1.311 |
1.080 |
-0.866 |
| H11 |
-1.311 |
1.079 |
0.868 |
| H12 |
-0.262 |
-1.203 |
0.865 |
| H13 |
-0.262 |
-1.202 |
-0.868 |
| H14 |
3.237 |
-0.163 |
-0.002 |
| H15 |
2.318 |
-1.405 |
0.877 |
| H16 |
2.316 |
-1.409 |
-0.873 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
O6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
| C1 | | 1.5174 | 2.5202 | 3.8799 | 5.0837 | 4.2849 | 1.0880 | 1.0897 | 1.0897 | 2.1543 | 2.1543 | 2.7713 | 2.7712 | 5.9695 | 5.2443 | 5.2428 |
C2 | 1.5174 | | 1.5139 | 2.5330 | 3.9068 | 2.7792 | 2.1642 | 2.1664 | 2.1664 | 1.0890 | 1.0890 | 2.1627 | 2.1627 | 4.6667 | 4.2286 | 4.2283 | C3 | 2.5202 | 1.5139 | | 1.5090 | 2.5636 | 2.3760 | 3.4679 | 2.7893 | 2.7893 | 2.1319 | 2.1319 | 1.0938 | 1.0938 | 3.4803 | 2.8247 | 2.8230 | C4 | 3.8799 | 2.5330 | 1.5090 | | 1.5052 | 1.2052 | 4.6895 | 4.2003 | 4.2004 | 2.7498 | 2.7499 | 2.1176 | 2.1175 | 2.1343 | 2.1417 | 2.1420 | C5 | 5.0837 | 3.9068 | 2.5636 | 1.5052 | | 2.3716 | 6.0041 | 5.2496 | 5.2492 | 4.1704 | 4.1699 | 2.7697 | 2.7706 | 1.0843 | 1.0896 | 1.0896 | O6 | 4.2849 | 2.7792 | 2.3760 | 1.2052 | 2.3716 | | 4.8757 | 4.7429 | 4.7434 | 2.6981 | 2.6989 | 3.0761 | 3.0754 | 2.5084 | 3.0826 | 3.0844 | H7 | 1.0880 | 2.1642 | 3.4679 | 4.6895 | 6.0041 | 4.8757 | | 1.7574 | 1.7574 | 2.4960 | 2.4960 | 3.7728 | 3.7727 | 6.8186 | 6.2130 | 6.2123 | H8 | 1.0897 | 2.1664 | 2.7893 | 4.2003 | 5.2496 | 4.7429 | 1.7574 | | 1.7574 | 2.5213 | 3.0666 | 3.1268 | 2.5943 | 6.1889 | 5.4712 | 5.1805 | H9 | 1.0897 | 2.1664 | 2.7893 | 4.2004 | 5.2492 | 4.7434 | 1.7574 | 1.7574 | | 3.0666 | 2.5213 | 2.5945 | 3.1268 | 6.1893 | 5.1815 | 5.4682 | H10 | 2.1543 | 1.0890 | 2.1319 | 2.7498 | 4.1704 | 2.6981 | 2.4960 | 2.5213 | 3.0666 | | 1.7338 | 3.0514 | 2.5111 | 4.7934 | 4.7306 | 4.3988 | H11 | 2.1543 | 1.0890 | 2.1319 | 2.7499 | 4.1699 | 2.6989 | 2.4960 | 3.0666 | 2.5213 | 1.7338 | | 2.5112 | 3.0514 | 4.7938 | 4.3970 | 4.7299 | H12 | 2.7713 | 2.1627 | 1.0938 | 2.1176 | 2.7697 | 3.0761 | 3.7728 | 3.1268 | 2.5945 | 3.0514 | 2.5112 | | 1.7332 | 3.7524 | 2.5876 | 3.1163 | H13 | 2.7712 | 2.1627 | 1.0938 | 2.1175 | 2.7706 | 3.0754 | 3.7727 | 2.5943 | 3.1268 | 2.5111 | 3.0514 | 1.7332 | | 3.7519 | 3.1211 | 2.5866 | H14 | 5.9695 | 4.6667 | 3.4803 | 2.1343 | 1.0843 | 2.5084 | 6.8186 | 6.1889 | 6.1893 | 4.7934 | 4.7938 | 3.7524 | 3.7519 | | 1.7781 | 1.7782 | H15 | 5.2443 | 4.2286 | 2.8247 | 2.1417 | 1.0896 | 3.0826 | 6.2130 | 5.4712 | 5.1815 | 4.7306 | 4.3970 | 2.5876 | 3.1211 | 1.7781 | | 1.7499 | H16 | 5.2428 | 4.2283 | 2.8230 | 2.1420 | 1.0896 | 3.0844 | 6.2123 | 5.1805 | 5.4682 | 4.3988 | 4.7299 | 3.1163 | 2.5866 | 1.7782 | 1.7499 | |
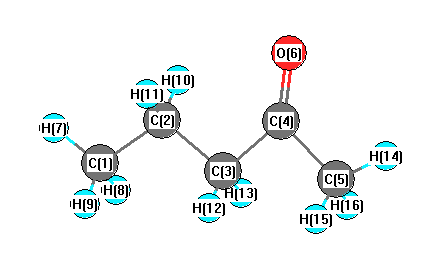 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
112.486 |
|
C1 |
C2 |
H10 |
110.411 |
| C1 |
C2 |
H11 |
110.410 |
|
C2 |
C1 |
H7 |
111.271 |
| C2 |
C1 |
H8 |
111.340 |
|
C2 |
C1 |
H9 |
111.342 |
| C2 |
C3 |
C4 |
113.842 |
|
C2 |
C3 |
H12 |
111.047 |
| C2 |
C3 |
H13 |
111.042 |
|
C3 |
C2 |
H10 |
108.884 |
| C3 |
C2 |
H11 |
108.885 |
|
C3 |
C4 |
C5 |
116.527 |
| C3 |
C4 |
O6 |
121.781 |
|
C4 |
C3 |
H12 |
107.826 |
| C4 |
C3 |
H13 |
107.818 |
|
C4 |
C5 |
H14 |
109.951 |
| C4 |
C5 |
H15 |
110.225 |
|
C4 |
C5 |
H16 |
110.250 |
| C5 |
C4 |
O6 |
121.693 |
|
H7 |
C1 |
H8 |
107.604 |
| H7 |
C1 |
H9 |
107.604 |
|
H8 |
C1 |
H9 |
107.482 |
| H10 |
C2 |
H11 |
105.511 |
|
H12 |
C3 |
H13 |
104.797 |
| H14 |
C5 |
H15 |
109.753 |
|
H14 |
C5 |
H16 |
109.769 |
| H15 |
C5 |
H16 |
106.843 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.366 |
|
|
|
| 2 |
C |
-0.212 |
|
|
|
| 3 |
C |
-0.234 |
|
|
|
| 4 |
C |
0.210 |
|
|
|
| 5 |
C |
-0.353 |
|
|
|
| 6 |
O |
-0.267 |
|
|
|
| 7 |
H |
0.123 |
|
|
|
| 8 |
H |
0.109 |
|
|
|
| 9 |
H |
0.109 |
|
|
|
| 10 |
H |
0.128 |
|
|
|
| 11 |
H |
0.128 |
|
|
|
| 12 |
H |
0.124 |
|
|
|
| 13 |
H |
0.124 |
|
|
|
| 14 |
H |
0.135 |
|
|
|
| 15 |
H |
0.120 |
|
|
|
| 16 |
H |
0.120 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.336 |
-2.670 |
0.001 |
2.691 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.639 |
-4.052 |
0.002 |
| y |
-4.052 |
-41.537 |
-0.000 |
| z |
0.002 |
-0.000 |
-36.980 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.620 |
-4.052 |
0.002 |
| y |
-4.052 |
-4.228 |
-0.000 |
| z |
0.002 |
-0.000 |
2.608 |
|
| Polar |
|---|
| 3z2-r2 | 5.216 |
|---|
| x2-y2 | 3.898 |
|---|
| xy | -4.052 |
|---|
| xz | 0.002 |
|---|
| yz | -0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.923 |
-0.170 |
0.000 |
| y |
-0.170 |
9.290 |
0.000 |
| z |
0.000 |
0.000 |
7.524 |
<r2> (average value of r
2) Å
2
| <r2> |
225.247 |
| (<r2>)1/2 |
15.008 |
