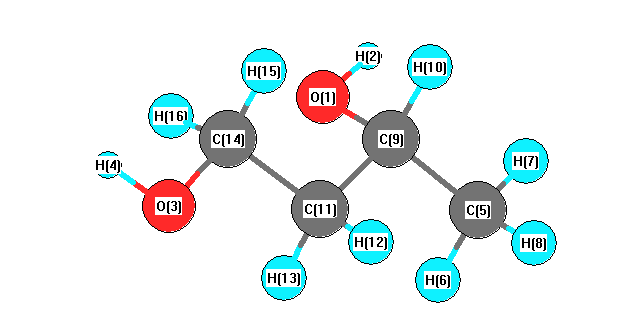
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3893 |
3726 |
31.65 |
|
|
|
| 2 |
A |
3877 |
3710 |
22.59 |
|
|
|
| 3 |
A |
3138 |
3004 |
30.78 |
|
|
|
| 4 |
A |
3120 |
2986 |
32.42 |
|
|
|
| 5 |
A |
3116 |
2982 |
26.72 |
|
|
|
| 6 |
A |
3068 |
2936 |
29.78 |
|
|
|
| 7 |
A |
3065 |
2934 |
13.87 |
|
|
|
| 8 |
A |
3049 |
2918 |
20.21 |
|
|
|
| 9 |
A |
2995 |
2867 |
79.42 |
|
|
|
| 10 |
A |
2982 |
2854 |
32.44 |
|
|
|
| 11 |
A |
1531 |
1465 |
2.99 |
|
|
|
| 12 |
A |
1497 |
1433 |
4.70 |
|
|
|
| 13 |
A |
1491 |
1427 |
5.38 |
|
|
|
| 14 |
A |
1480 |
1416 |
2.23 |
|
|
|
| 15 |
A |
1464 |
1401 |
9.11 |
|
|
|
| 16 |
A |
1431 |
1370 |
19.56 |
|
|
|
| 17 |
A |
1404 |
1344 |
14.47 |
|
|
|
| 18 |
A |
1380 |
1321 |
4.17 |
|
|
|
| 19 |
A |
1326 |
1269 |
4.86 |
|
|
|
| 20 |
A |
1321 |
1264 |
23.34 |
|
|
|
| 21 |
A |
1291 |
1236 |
35.05 |
|
|
|
| 22 |
A |
1251 |
1197 |
47.97 |
|
|
|
| 23 |
A |
1222 |
1169 |
4.72 |
|
|
|
| 24 |
A |
1162 |
1113 |
36.86 |
|
|
|
| 25 |
A |
1142 |
1093 |
35.42 |
|
|
|
| 26 |
A |
1101 |
1053 |
32.40 |
|
|
|
| 27 |
A |
1080 |
1033 |
80.65 |
|
|
|
| 28 |
A |
1042 |
998 |
9.32 |
|
|
|
| 29 |
A |
988 |
946 |
8.44 |
|
|
|
| 30 |
A |
950 |
909 |
9.09 |
|
|
|
| 31 |
A |
867 |
829 |
15.92 |
|
|
|
| 32 |
A |
801 |
766 |
3.29 |
|
|
|
| 33 |
A |
496 |
475 |
18.74 |
|
|
|
| 34 |
A |
477 |
456 |
5.23 |
|
|
|
| 35 |
A |
388 |
371 |
5.88 |
|
|
|
| 36 |
A |
340 |
325 |
7.31 |
|
|
|
| 37 |
A |
276 |
264 |
131.42 |
|
|
|
| 38 |
A |
264 |
252 |
72.10 |
|
|
|
| 39 |
A |
242 |
232 |
0.67 |
|
|
|
| 40 |
A |
179 |
172 |
3.71 |
|
|
|
| 41 |
A |
122 |
117 |
5.46 |
|
|
|
| 42 |
A |
92 |
88 |
7.16 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31199.1 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 29860.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.346 |
|
|
|
| 2 |
H |
0.205 |
|
|
|
| 3 |
O |
-0.337 |
|
|
|
| 4 |
H |
0.203 |
|
|
|
| 5 |
C |
-0.355 |
|
|
|
| 6 |
H |
0.123 |
|
|
|
| 7 |
H |
0.103 |
|
|
|
| 8 |
H |
0.109 |
|
|
|
| 9 |
C |
0.055 |
|
|
|
| 10 |
H |
0.084 |
|
|
|
| 11 |
C |
-0.209 |
|
|
|
| 12 |
H |
0.119 |
|
|
|
| 13 |
H |
0.123 |
|
|
|
| 14 |
C |
-0.032 |
|
|
|
| 15 |
H |
0.064 |
|
|
|
| 16 |
H |
0.091 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.149 |
0.754 |
1.359 |
1.933 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.037 |
-2.688 |
-0.316 |
| y |
-2.688 |
-38.552 |
0.744 |
| z |
-0.316 |
0.744 |
-38.691 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.585 |
-2.688 |
-0.316 |
| y |
-2.688 |
-3.188 |
0.744 |
| z |
-0.316 |
0.744 |
-3.397 |
|
| Polar |
|---|
| 3z2-r2 | -6.794 |
|---|
| x2-y2 | 6.515 |
|---|
| xy | -2.688 |
|---|
| xz | -0.316 |
|---|
| yz | 0.744 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.012 |
-0.198 |
0.041 |
| y |
-0.198 |
8.184 |
0.120 |
| z |
0.041 |
0.120 |
7.495 |
<r2> (average value of r
2) Å
2
| <r2> |
220.886 |
| (<r2>)1/2 |
14.862 |