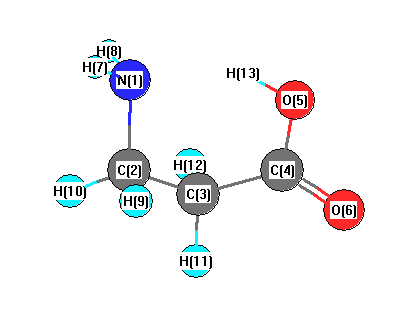
Vibrational Frequencies calculated at mPW1PW91/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3630 |
3482 |
9.45 |
|
|
|
| 2 |
A |
3545 |
3400 |
1.98 |
|
|
|
| 3 |
A |
3265 |
3132 |
758.20 |
|
|
|
| 4 |
A |
3142 |
3014 |
5.80 |
|
|
|
| 5 |
A |
3097 |
2971 |
18.85 |
|
|
|
| 6 |
A |
3049 |
2924 |
4.93 |
|
|
|
| 7 |
A |
3032 |
2908 |
58.17 |
|
|
|
| 8 |
A |
1867 |
1791 |
421.86 |
|
|
|
| 9 |
A |
1657 |
1589 |
29.90 |
|
|
|
| 10 |
A |
1517 |
1455 |
43.04 |
|
|
|
| 11 |
A |
1503 |
1442 |
208.23 |
|
|
|
| 12 |
A |
1462 |
1403 |
9.42 |
|
|
|
| 13 |
A |
1422 |
1364 |
10.89 |
|
|
|
| 14 |
A |
1375 |
1319 |
10.64 |
|
|
|
| 15 |
A |
1319 |
1265 |
6.99 |
|
|
|
| 16 |
A |
1304 |
1251 |
25.77 |
|
|
|
| 17 |
A |
1260 |
1209 |
64.15 |
|
|
|
| 18 |
A |
1154 |
1107 |
6.83 |
|
|
|
| 19 |
A |
1095 |
1050 |
7.02 |
|
|
|
| 20 |
A |
1026 |
984 |
8.91 |
|
|
|
| 21 |
A |
1003 |
962 |
82.16 |
|
|
|
| 22 |
A |
966 |
927 |
8.95 |
|
|
|
| 23 |
A |
920 |
882 |
19.40 |
|
|
|
| 24 |
A |
873 |
837 |
42.85 |
|
|
|
| 25 |
A |
820 |
787 |
27.06 |
|
|
|
| 26 |
A |
704 |
675 |
9.44 |
|
|
|
| 27 |
A |
583 |
559 |
2.20 |
|
|
|
| 28 |
A |
493 |
473 |
8.74 |
|
|
|
| 29 |
A |
408 |
391 |
11.75 |
|
|
|
| 30 |
A |
326 |
313 |
7.86 |
|
|
|
| 31 |
A |
287 |
275 |
9.71 |
|
|
|
| 32 |
A |
200 |
191 |
4.68 |
|
|
|
| 33 |
A |
87 |
84 |
0.49 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24194.1 cm
-1
Scaled (by 0.9592) Zero Point Vibrational Energy (zpe) 23207.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.283 |
|
|
|
| 2 |
C |
-0.093 |
|
|
|
| 3 |
C |
-0.205 |
|
|
|
| 4 |
C |
0.280 |
|
|
|
| 5 |
O |
-0.255 |
|
|
|
| 6 |
O |
-0.306 |
|
|
|
| 7 |
H |
0.136 |
|
|
|
| 8 |
H |
0.129 |
|
|
|
| 9 |
H |
0.084 |
|
|
|
| 10 |
H |
0.103 |
|
|
|
| 11 |
H |
0.103 |
|
|
|
| 12 |
H |
0.100 |
|
|
|
| 13 |
H |
0.206 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
6.435 |
-1.184 |
0.561 |
6.567 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.034 |
1.584 |
-0.262 |
| y |
1.584 |
-37.087 |
0.066 |
| z |
-0.262 |
0.066 |
-33.411 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.785 |
1.584 |
-0.262 |
| y |
1.584 |
-0.864 |
0.066 |
| z |
-0.262 |
0.066 |
4.650 |
|
| Polar |
|---|
| 3z2-r2 | 9.300 |
|---|
| x2-y2 | -1.947 |
|---|
| xy | 1.584 |
|---|
| xz | -0.262 |
|---|
| yz | 0.066 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.857 |
0.352 |
0.087 |
| y |
0.352 |
7.278 |
-0.012 |
| z |
0.087 |
-0.012 |
5.917 |
<r2> (average value of r
2) Å
2
| <r2> |
170.408 |
| (<r2>)1/2 |
13.054 |