return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3NHCH2COOH (Sarcosine)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-323.255485
Energy at 298.15K-323.265072
HF Energy-321.980102
Nuclear repulsion energy245.352437
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3566 3386 7.83      
2 A 3511 3333 321.83      
3 A 3182 3021 14.83      
4 A 3145 2986 12.51      
5 A 3137 2978 12.23      
6 A 3097 2940 14.22      
7 A 3059 2904 55.74      
8 A 1867 1773 309.89      
9 A 1546 1467 9.80      
10 A 1526 1449 14.58      
11 A 1505 1429 15.05      
12 A 1484 1409 9.45      
13 A 1472 1397 5.61      
14 A 1436 1363 368.12      
15 A 1358 1290 10.63      
16 A 1306 1240 4.07      
17 A 1252 1188 15.91      
18 A 1204 1143 30.07      
19 A 1172 1113 24.20      
20 A 1153 1094 10.56      
21 A 1016 964 13.43      
22 A 1000 949 19.51      
23 A 909 863 54.49      
24 A 898 853 22.91      
25 A 793 753 64.32      
26 A 656 623 3.34      
27 A 591 561 7.41      
28 A 485 460 9.58      
29 A 380 361 5.08      
30 A 281 267 4.06      
31 A 208 198 2.43      
32 A 133 126 2.83      
33 A 78 74 5.87      

Unscaled Zero Point Vibrational Energy (zpe) 24201.4 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 22976.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
0.30598 0.06810 0.05835

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 1.192 0.276 -0.314
C2 2.512 -0.084 0.181
C3 0.168 -0.703 0.009
C4 -1.210 -0.070 0.022
O5 -1.176 1.264 0.114
O6 -2.229 -0.707 -0.013
H7 1.233 0.394 -1.317
H8 0.140 -1.572 -0.644
H9 2.852 -1.060 -0.166
H10 3.231 0.665 -0.128
H11 0.348 -1.063 1.018
H12 2.487 -0.098 1.265
H13 -0.218 1.475 0.098

Atom - Atom Distances (Å)
  N1 C2 C3 C4 O5 O6 H7 H8 H9 H10 H11 H12 H13
N11.45511.45292.45032.60103.57181.01092.15182.13662.08442.06922.07651.8960
C21.45512.43013.72583.92694.78522.02722.91871.09061.08372.51821.08473.1449
C31.45292.43011.51702.38452.39692.02381.08752.71343.35751.08702.70562.2136
C42.45033.72581.51701.33751.20162.82462.12704.18594.50472.10003.90101.8377
O52.60103.92692.38451.33752.23822.93373.21704.65904.45422.92534.07390.9808
O63.57184.78522.39691.20162.23823.85952.59975.09555.63092.79854.92352.9691
H71.01092.02722.02382.82462.93373.85952.34782.46242.34122.89182.91292.2970
H82.15182.91871.08752.12703.21702.59972.34782.80093.85021.75113.36513.1563
H92.13661.09062.71344.18594.65905.09552.46242.80091.76692.77001.76283.9905
H102.08441.08373.35754.50474.45425.63092.34123.85021.76693.55161.75473.5504
H112.06922.51821.08702.10002.92532.79852.89181.75112.77003.55162.35922.7588
H122.07651.08472.70563.90104.07394.92352.91293.36511.76281.75472.35923.3399
H131.89603.14492.21361.83770.98082.96912.29703.15633.99053.55042.75883.3399

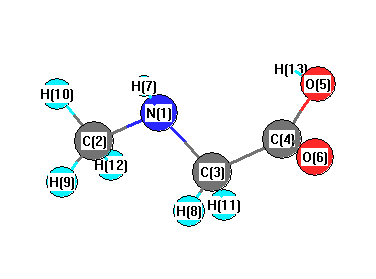
picture of Sarcosine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C2 H9 113.364 N1 C2 H10 109.512
N1 C2 H12 108.812 N1 C3 C4 111.164
N1 C3 H8 115.026 N1 C3 H11 108.250
C2 N1 C3 113.370 C2 N1 H7 109.256
C3 N1 H7 109.132 C3 C4 O5 113.150
C3 C4 O6 123.268 C4 C3 H8 108.378
C4 C3 H11 106.323 C4 O5 H13 103.808
O5 C4 O6 123.556 H8 C3 H11 107.281
H9 C2 H10 108.714 H9 C2 H12 108.267
H10 C2 H12 108.035
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability