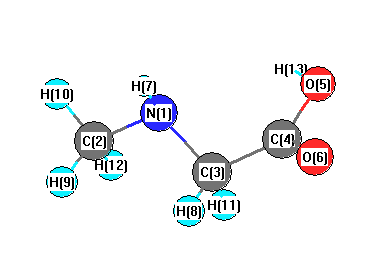
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3551 |
3399 |
4.93 |
|
|
|
| 2 |
A |
3494 |
3344 |
310.42 |
|
|
|
| 3 |
A |
3132 |
2998 |
20.16 |
|
|
|
| 4 |
A |
3095 |
2962 |
0.59 |
|
|
|
| 5 |
A |
3091 |
2958 |
30.91 |
|
|
|
| 6 |
A |
3048 |
2917 |
19.55 |
|
|
|
| 7 |
A |
3004 |
2875 |
69.81 |
|
|
|
| 8 |
A |
1883 |
1802 |
376.34 |
|
|
|
| 9 |
A |
1515 |
1450 |
12.74 |
|
|
|
| 10 |
A |
1493 |
1429 |
15.53 |
|
|
|
| 11 |
A |
1483 |
1420 |
14.80 |
|
|
|
| 12 |
A |
1468 |
1405 |
6.51 |
|
|
|
| 13 |
A |
1452 |
1389 |
6.96 |
|
|
|
| 14 |
A |
1422 |
1361 |
389.26 |
|
|
|
| 15 |
A |
1340 |
1282 |
9.15 |
|
|
|
| 16 |
A |
1287 |
1231 |
3.73 |
|
|
|
| 17 |
A |
1233 |
1180 |
13.28 |
|
|
|
| 18 |
A |
1186 |
1135 |
32.42 |
|
|
|
| 19 |
A |
1156 |
1107 |
24.28 |
|
|
|
| 20 |
A |
1136 |
1087 |
9.44 |
|
|
|
| 21 |
A |
1002 |
959 |
10.40 |
|
|
|
| 22 |
A |
981 |
939 |
24.42 |
|
|
|
| 23 |
A |
892 |
854 |
54.64 |
|
|
|
| 24 |
A |
888 |
849 |
17.00 |
|
|
|
| 25 |
A |
771 |
738 |
64.85 |
|
|
|
| 26 |
A |
645 |
617 |
3.25 |
|
|
|
| 27 |
A |
579 |
554 |
7.91 |
|
|
|
| 28 |
A |
478 |
457 |
9.91 |
|
|
|
| 29 |
A |
372 |
356 |
5.34 |
|
|
|
| 30 |
A |
280 |
268 |
4.98 |
|
|
|
| 31 |
A |
203 |
194 |
2.11 |
|
|
|
| 32 |
A |
134 |
128 |
2.63 |
|
|
|
| 33 |
A |
72 |
69 |
5.91 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23880.9 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 22856.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.255 |
|
|
|
| 2 |
C |
-0.259 |
|
|
|
| 3 |
C |
-0.175 |
|
|
|
| 4 |
C |
0.256 |
|
|
|
| 5 |
O |
-0.227 |
|
|
|
| 6 |
O |
-0.299 |
|
|
|
| 7 |
H |
0.150 |
|
|
|
| 8 |
H |
0.124 |
|
|
|
| 9 |
H |
0.100 |
|
|
|
| 10 |
H |
0.121 |
|
|
|
| 11 |
H |
0.133 |
|
|
|
| 12 |
H |
0.120 |
|
|
|
| 13 |
H |
0.211 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
5.349 |
-0.104 |
-0.609 |
5.385 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-43.079 |
-0.868 |
-0.455 |
| y |
-0.868 |
-37.294 |
-0.576 |
| z |
-0.455 |
-0.576 |
-33.236 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-7.814 |
-0.868 |
-0.455 |
| y |
-0.868 |
0.863 |
-0.576 |
| z |
-0.455 |
-0.576 |
6.951 |
|
| Polar |
|---|
| 3z2-r2 | 13.902 |
|---|
| x2-y2 | -5.785 |
|---|
| xy | -0.868 |
|---|
| xz | -0.455 |
|---|
| yz | -0.576 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.248 |
0.337 |
0.044 |
| y |
0.337 |
7.342 |
0.016 |
| z |
0.044 |
0.016 |
5.837 |
<r2> (average value of r
2) Å
2
| <r2> |
191.699 |
| (<r2>)1/2 |
13.846 |