return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H5CH3 (toluene)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-271.066050
Energy at 298.15K 
HF Energy-269.829866
Nuclear repulsion energy270.703455
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3251 3086 8.88      
2 A' 3224 3061 6.50      
3 A' 3203 3040 4.49      
4 A' 3152 2993 11.48      
5 A' 3084 2928 22.11      
6 A' 1668 1584 5.38      
7 A' 1539 1461 13.52      
8 A' 1513 1436 6.37      
9 A' 1415 1343 0.91      
10 A' 1256 1192 0.38      
11 A' 1216 1155 0.16      
12 A' 1070 1015 7.40      
13 A' 1061 1007 2.47      
14 A' 1030 978 0.04      
15 A' 995 945 0.04      
16 A' 931 884 0.63      
17 A' 808 767 0.79      
18 A' 764 725 53.21      
19 A' 719 683 17.14      
20 A' 527 500 0.67      
21 A' 480 456 8.51      
22 A' 214 203 2.08      
23 A" 3234 3071 25.33      
24 A" 3209 3047 2.54      
25 A" 3172 3011 10.23      
26 A" 1644 1561 0.01      
27 A" 1521 1444 13.37      
28 A" 1489 1413 0.64      
29 A" 1465 1391 0.23      
30 A" 1351 1283 0.09      
31 A" 1196 1136 0.03      
32 A" 1122 1065 5.24      
33 A" 1007 956 0.13      
34 A" 987 937 0.00      
35 A" 875 831 0.03      
36 A" 629 597 0.07      
37 A" 417 396 0.00      
38 A" 340 323 0.23      
39 A" 29i 28i 0.19      

Unscaled Zero Point Vibrational Energy (zpe) 28373.2 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 26937.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
0.18703 0.08509 0.05912

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.006 0.907 0.000
C2 0.007 0.196 1.194
C3 0.007 -1.191 1.199
C4 0.006 -1.890 0.000
C5 0.007 -1.191 -1.199
C6 0.007 0.196 -1.194
C7 -0.027 2.403 0.000
H8 0.011 0.733 2.129
H9 0.011 -1.723 2.135
H10 0.009 -2.967 0.000
H11 0.011 -1.723 -2.135
H12 0.011 0.733 -2.129
H13 -1.053 2.764 0.000
H14 0.466 2.803 0.880
H15 0.466 2.803 -0.880

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 H8 H9 H10 H11 H12 H13 H14 H15
C11.39012.41672.79772.41671.39011.49572.13633.38803.87393.38802.13632.13732.14012.1401
C21.39011.38732.40432.76632.38892.50921.07802.13763.38093.84323.36673.02362.66553.3631
C32.41671.38731.38792.39772.76633.78852.13751.07692.14243.37603.84434.26634.03304.5261
C42.79772.40431.38791.38792.40434.29313.37892.14151.07632.14153.37894.77314.79734.7973
C52.41672.76632.39771.38791.38733.78853.84433.37602.14241.07692.13754.26634.52614.0330
C61.39012.38892.76632.40431.38732.50923.36673.84323.38092.13761.07803.02363.36312.6655
C71.49572.50923.78854.29313.78852.50922.70584.64585.36934.64582.70581.08751.08541.0854
H82.13631.07802.13753.37893.84433.36672.70582.45674.26884.92124.25833.12862.45983.6807
H93.38802.13761.07692.14153.37603.84324.64582.45672.47064.26994.92125.08194.71915.4577
H103.87393.38092.14241.07632.14243.38095.36934.26882.47062.47064.26885.82815.85425.8542
H113.38803.84323.37602.14151.07692.13764.64584.92124.26992.47062.45675.08195.45774.7191
H122.13633.36673.84433.37892.13751.07802.70584.25834.92124.26882.45673.12863.68072.4598
H132.13733.02364.26634.77314.26633.02361.08753.12865.08195.82815.08193.12861.75561.7556
H142.14012.66554.03304.79734.52613.36311.08542.45984.71915.85425.45773.68071.75561.7605
H152.14013.36314.52614.79734.03302.66551.08543.68075.45775.85424.71912.45981.75561.7605

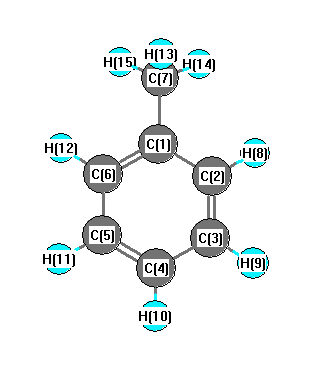
picture of toluene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 120.947 C1 C2 H8 119.352
C1 C6 C5 120.947 C1 C6 H12 119.352
C1 C7 H13 110.674 C1 C7 H14 111.025
C1 C7 H15 111.025 C2 C1 C6 118.467
C2 C1 C7 120.759 C2 C3 C4 120.074
C2 C3 H9 119.805 C3 C2 H8 119.700
C3 C4 C5 119.490 C3 C4 H10 120.255
C4 C3 H9 120.121 C4 C5 C6 120.074
C4 C5 H11 120.121 C5 C4 H10 120.255
C5 C6 H12 119.700 C6 C1 C7 120.759
C6 C5 H11 119.805 H13 C7 H14 107.799
H13 C7 H15 107.799 H14 C7 H15 108.390
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability