return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H5Cl (chlorobenzene)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-690.975483
Energy at 298.15K-690.980873
HF Energy-689.715596
Nuclear repulsion energy317.368144
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3265 3100 1.67      
2 A1 3250 3085 7.22      
3 A1 3218 3055 0.08      
4 A1 1642 1559 22.16      
5 A1 1518 1441 41.39      
6 A1 1209 1148 0.15      
7 A1 1133 1076 38.51      
8 A1 1054 1000 11.11      
9 A1 1025 974 3.51      
10 A1 720 684 18.22      
11 A1 424 403 2.06      
12 A2 993 943 0.00      
13 A2 865 821 0.00      
14 A2 421 399 0.00      
15 B1 1004 953 0.12      
16 B1 941 893 3.28      
17 B1 779 740 58.02      
18 B1 717 681 20.26      
19 B1 489 465 8.30      
20 B1 193 183 0.15      
21 B2 3261 3096 2.77      
22 B2 3231 3068 5.49      
23 B2 1641 1558 2.34      
24 B2 1486 1411 4.41      
25 B2 1470 1396 1.77      
26 B2 1332 1264 0.36      
27 B2 1198 1137 0.15      
28 B2 1105 1050 4.90      
29 B2 620 589 0.23      
30 B2 297 282 0.11      

Unscaled Zero Point Vibrational Energy (zpe) 20250.9 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 19226.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
0.19180 0.05302 0.04154

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
Cl1 0.000 0.000 2.239
C2 0.000 0.000 0.505
C3 0.000 1.208 -0.172
C4 0.000 -1.208 -0.172
C5 0.000 1.201 -1.560
C6 0.000 -1.201 -1.560
C7 0.000 0.000 -2.256
H8 0.000 2.131 0.378
H9 0.000 -2.131 0.378
H10 0.000 2.136 -2.093
H11 0.000 -2.136 -2.093
H12 0.000 0.000 -3.332

Atom - Atom Distances (Å)
  Cl1 C2 C3 C4 C5 C6 C7 H8 H9 H10 H11 H12
Cl11.73382.69672.69673.98373.98374.49502.82892.82894.82944.82945.5710
C21.73381.38461.38462.38832.38832.76122.13462.13463.36293.36293.8371
C32.69671.38462.41511.38732.77912.40851.07493.38352.13283.85553.3827
C42.69671.38462.41512.77911.38732.40853.38351.07493.85552.13283.3827
C53.98372.38831.38732.77912.40101.38802.14953.85401.07643.37862.1408
C63.98372.38832.77911.38732.40101.38803.85402.14953.37861.07642.1408
C74.49502.76122.40852.40851.38801.38803.38823.38822.14202.14201.0759
H82.82892.13461.07493.38352.14953.85403.38824.26172.47074.93034.2786
H92.82892.13463.38351.07493.85402.14953.38824.26174.93032.47074.2786
H104.82943.36292.13283.85551.07643.37862.14202.47074.93034.27152.4694
H114.82943.36293.85552.13283.37861.07642.14204.93032.47074.27152.4694
H125.57103.83713.38273.38272.14082.14081.07594.27864.27862.46942.4694

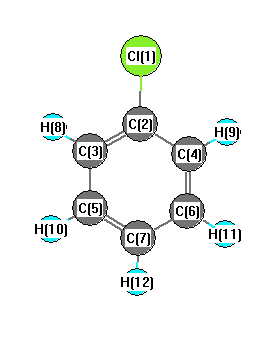
picture of chlorobenzene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
Cl1 C2 C3 119.290 Cl1 C2 C4 119.290
C2 C3 C5 118.999 C2 C3 H8 119.907
C2 C4 C6 118.999 C2 C4 H9 119.907
C3 C2 C4 121.421 C3 C5 C7 120.414
C3 C5 H10 119.387 C4 C6 C7 120.414
C4 C6 H11 119.387 C5 C3 H8 121.094
C5 C7 C6 119.753 C5 C7 H12 120.123
C6 C4 H9 121.094 C6 C7 H12 120.123
C7 C5 H10 120.199 C7 C6 H11 120.199
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability