All results from a given calculation for C6H13N (cyclohexanamine)
using model chemistry: MP2=FULL/cc-pVTZ
19 10 17 12 22
States and conformations
| State |
Conformation |
minimum conformation |
conformer description |
state description |
| 1 |
1 |
yes |
C1 |
1A |
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -290.699798 |
| Energy at 298.15K | |
| HF Energy | -289.333249 |
| Nuclear repulsion energy | 333.090913 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.860 |
0.011 |
0.292 |
| C2 |
1.168 |
-1.241 |
-0.221 |
| C3 |
-0.302 |
-1.250 |
0.163 |
| C4 |
-1.018 |
-0.009 |
-0.332 |
| C5 |
-0.315 |
1.238 |
0.184 |
| C6 |
1.154 |
1.259 |
-0.208 |
| N7 |
-2.428 |
-0.100 |
0.024 |
| H8 |
2.904 |
0.018 |
-0.009 |
| H9 |
1.843 |
0.006 |
1.382 |
| H10 |
1.251 |
-1.273 |
-1.307 |
| H11 |
1.661 |
-2.132 |
0.159 |
| H12 |
-0.807 |
-2.131 |
-0.223 |
| H13 |
-0.389 |
-1.281 |
1.252 |
| H14 |
-0.964 |
-0.003 |
-1.421 |
| H15 |
-0.401 |
1.248 |
1.272 |
| H16 |
-0.826 |
2.126 |
-0.185 |
| H17 |
1.637 |
2.153 |
0.177 |
| H18 |
1.232 |
1.301 |
-1.295 |
| H19 |
-2.911 |
0.737 |
-0.274 |
| H20 |
-2.509 |
-0.121 |
1.033 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
N7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
H19 |
H20 |
| C1 | | 1.5194 | 2.5059 | 2.9447 | 2.4993 | 1.5192 | 4.2976 | 1.0870 | 1.0905 | 2.1391 | 2.1556 | 3.4586 | 2.7654 | 3.3030 | 2.7574 | 3.4516 | 2.1570 | 2.1394 | 4.8592 | 4.4330 |
C2 | 1.5194 | | 1.5191 | 2.5114 | 2.9166 | 2.5003 | 3.7803 | 2.1551 | 2.1402 | 1.0903 | 1.0868 | 2.1657 | 2.1433 | 2.7419 | 3.2993 | 3.9128 | 3.4494 | 2.7603 | 4.5336 | 4.0426 | C3 | 2.5059 | 1.5191 | | 1.5158 | 2.4880 | 2.9245 | 2.4210 | 3.4521 | 2.7686 | 2.1392 | 2.1521 | 1.0862 | 1.0926 | 2.1223 | 2.7350 | 3.4338 | 3.9168 | 3.3145 | 3.3088 | 2.6269 | C4 | 2.9447 | 2.5114 | 1.5158 | | 1.5216 | 2.5177 | 1.4571 | 3.9354 | 3.3353 | 2.7747 | 3.4532 | 2.1349 | 2.1264 | 1.0912 | 2.1294 | 2.1484 | 3.4617 | 2.7759 | 2.0358 | 2.0243 | C5 | 2.4993 | 2.9166 | 2.4880 | 1.5216 | | 1.5198 | 2.5065 | 3.4478 | 2.7589 | 3.3137 | 3.9063 | 3.4286 | 2.7372 | 2.1304 | 1.0920 | 1.0887 | 2.1557 | 2.1408 | 2.6839 | 2.7171 | C6 | 1.5192 | 2.5003 | 2.9245 | 2.5177 | 1.5198 | | 3.8379 | 2.1553 | 2.1393 | 2.7622 | 3.4483 | 3.9159 | 3.3113 | 2.7475 | 2.1467 | 2.1608 | 1.0869 | 1.0905 | 4.0987 | 4.1060 | N7 | 4.2976 | 3.7803 | 2.4210 | 1.4571 | 2.5065 | 3.8379 | | 5.3336 | 4.4833 | 4.0848 | 4.5680 | 2.6099 | 2.6570 | 2.0597 | 2.7361 | 2.7507 | 4.6504 | 4.1350 | 1.0118 | 1.0126 | H8 | 1.0870 | 2.1551 | 3.4521 | 3.9354 | 3.4478 | 2.1553 | 5.3336 | | 1.7500 | 2.4663 | 2.4887 | 4.2934 | 3.7581 | 4.1178 | 3.7523 | 4.2879 | 2.4899 | 2.4688 | 5.8657 | 5.5142 | H9 | 1.0905 | 2.1402 | 2.7686 | 3.3353 | 2.7589 | 2.1393 | 4.4833 | 1.7500 | | 3.0363 | 2.4694 | 3.7636 | 2.5800 | 3.9675 | 2.5674 | 3.7516 | 2.4709 | 3.0362 | 5.0877 | 4.3679 | H10 | 2.1391 | 1.0903 | 2.1392 | 2.7747 | 3.3137 | 2.7622 | 4.0848 | 2.4663 | 3.0363 | | 1.7481 | 2.4794 | 3.0399 | 2.5560 | 3.9674 | 4.1383 | 3.7538 | 2.5739 | 4.7365 | 4.5763 | H11 | 2.1556 | 1.0868 | 2.1521 | 3.4532 | 3.9063 | 3.4483 | 4.5680 | 2.4887 | 2.4694 | 1.7481 | | 2.4975 | 2.4741 | 3.7310 | 4.1127 | 4.9426 | 4.2849 | 3.7525 | 5.4153 | 4.7111 | H12 | 3.4586 | 2.1657 | 1.0862 | 2.1349 | 3.4286 | 3.9159 | 2.6099 | 4.2934 | 3.7636 | 2.4794 | 2.4975 | | 1.7527 | 2.4469 | 3.7172 | 4.2567 | 4.9481 | 4.1329 | 3.5574 | 2.9176 | H13 | 2.7654 | 2.1433 | 1.0926 | 2.1264 | 2.7372 | 3.3113 | 2.6570 | 3.7581 | 2.5800 | 3.0399 | 2.4741 | 1.7527 | | 3.0185 | 2.5293 | 3.7232 | 4.1296 | 3.9725 | 3.5727 | 2.4262 | H14 | 3.3030 | 2.7419 | 2.1223 | 1.0912 | 2.1304 | 2.7475 | 2.0597 | 4.1178 | 3.9675 | 2.5560 | 3.7310 | 2.4469 | 3.0185 | | 3.0232 | 2.4658 | 3.7377 | 2.5569 | 2.3783 | 2.9028 | H15 | 2.7574 | 3.2993 | 2.7350 | 2.1294 | 1.0920 | 2.1467 | 2.7361 | 3.7523 | 2.5674 | 3.9674 | 4.1127 | 3.7172 | 2.5293 | 3.0232 | | 1.7534 | 2.4842 | 3.0431 | 2.9927 | 2.5251 | H16 | 3.4516 | 3.9128 | 3.4338 | 2.1484 | 1.0887 | 2.1608 | 2.7507 | 4.2879 | 3.7516 | 4.1383 | 4.9426 | 4.2567 | 3.7232 | 2.4658 | 1.7534 | | 2.4895 | 2.4793 | 2.5071 | 3.0603 | H17 | 2.1570 | 3.4494 | 3.9168 | 3.4617 | 2.1557 | 1.0869 | 4.6504 | 2.4899 | 2.4709 | 3.7538 | 4.2849 | 4.9481 | 4.1296 | 3.7377 | 2.4842 | 2.4895 | | 1.7488 | 4.7851 | 4.8056 | H18 | 2.1394 | 2.7603 | 3.3145 | 2.7759 | 2.1408 | 1.0905 | 4.1350 | 2.4688 | 3.0362 | 2.5739 | 3.7525 | 4.1329 | 3.9725 | 2.5569 | 3.0431 | 2.4793 | 1.7488 | | 4.3042 | 4.6299 | H19 | 4.8592 | 4.5336 | 3.3088 | 2.0358 | 2.6839 | 4.0987 | 1.0118 | 5.8657 | 5.0877 | 4.7365 | 5.4153 | 3.5574 | 3.5727 | 2.3783 | 2.9927 | 2.5071 | 4.7851 | 4.3042 | | 1.6151 | H20 | 4.4330 | 4.0426 | 2.6269 | 2.0243 | 2.7171 | 4.1060 | 1.0126 | 5.5142 | 4.3679 | 4.5763 | 4.7111 | 2.9176 | 2.4262 | 2.9028 | 2.5251 | 3.0603 | 4.8056 | 4.6299 | 1.6151 | |
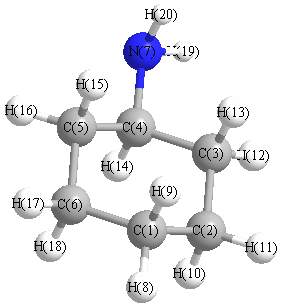 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
111.119 |
|
C1 |
C2 |
H10 |
108.997 |
| C1 |
C2 |
H11 |
110.508 |
|
C1 |
C6 |
C5 |
110.655 |
| C1 |
C6 |
H17 |
110.626 |
|
C1 |
C6 |
H18 |
109.028 |
| C2 |
C1 |
C6 |
110.742 |
|
C2 |
C1 |
H8 |
110.458 |
| C2 |
C1 |
H9 |
109.072 |
|
C2 |
C3 |
C4 |
111.691 |
| C2 |
C3 |
H12 |
111.380 |
|
C2 |
C3 |
H13 |
109.214 |
| C3 |
C2 |
H10 |
109.024 |
|
C3 |
C2 |
H11 |
110.250 |
| C3 |
C4 |
C5 |
109.999 |
|
C3 |
C4 |
N7 |
109.032 |
| C3 |
C4 |
H14 |
107.889 |
|
C4 |
C3 |
H12 |
109.157 |
| C4 |
C3 |
H13 |
108.124 |
|
C4 |
C5 |
C6 |
111.751 |
| C4 |
C5 |
H15 |
107.994 |
|
C4 |
C5 |
H16 |
109.667 |
| C4 |
N7 |
H19 |
109.779 |
|
C4 |
N7 |
H20 |
108.768 |
| C5 |
C4 |
N7 |
114.576 |
|
C5 |
C4 |
H14 |
108.122 |
| C5 |
C6 |
H17 |
110.479 |
|
C5 |
C6 |
H18 |
109.093 |
| C6 |
C1 |
H8 |
110.487 |
|
C6 |
C1 |
H9 |
109.015 |
| C6 |
C5 |
H15 |
109.462 |
|
C6 |
C5 |
H16 |
110.776 |
| N7 |
C4 |
H14 |
106.982 |
|
H8 |
C1 |
H9 |
106.970 |
| H10 |
C2 |
H11 |
106.826 |
|
H12 |
C3 |
H13 |
107.116 |
| H15 |
C5 |
H16 |
107.038 |
|
H17 |
C6 |
H18 |
106.862 |
| H19 |
N7 |
H20 |
105.838 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
