Jump to
S1C2
Vibrational Frequencies calculated at B1B95/cc-pVTZ
Geometric Data calculated at B1B95/cc-pVTZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -196.476200 |
| Energy at 298.15K | -196.486474 |
| HF Energy | -196.476200 |
| Nuclear repulsion energy | 174.325409 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3240 |
3101 |
16.68 |
|
|
|
| 2 |
A |
3155 |
3019 |
3.97 |
|
|
|
| 3 |
A |
3140 |
3005 |
28.02 |
|
|
|
| 4 |
A |
3133 |
2998 |
34.65 |
|
|
|
| 5 |
A |
3122 |
2988 |
39.33 |
|
|
|
| 6 |
A |
3087 |
2954 |
25.35 |
|
|
|
| 7 |
A |
3078 |
2946 |
23.95 |
|
|
|
| 8 |
A |
3053 |
2922 |
24.81 |
|
|
|
| 9 |
A |
3046 |
2915 |
25.20 |
|
|
|
| 10 |
A |
3035 |
2905 |
23.50 |
|
|
|
| 11 |
A |
1730 |
1656 |
12.15 |
|
|
|
| 12 |
A |
1507 |
1442 |
4.81 |
|
|
|
| 13 |
A |
1501 |
1437 |
7.75 |
|
|
|
| 14 |
A |
1489 |
1425 |
0.50 |
|
|
|
| 15 |
A |
1478 |
1415 |
8.64 |
|
|
|
| 16 |
A |
1453 |
1391 |
1.56 |
|
|
|
| 17 |
A |
1417 |
1356 |
5.57 |
|
|
|
| 18 |
A |
1374 |
1315 |
2.34 |
|
|
|
| 19 |
A |
1355 |
1297 |
0.53 |
|
|
|
| 20 |
A |
1324 |
1267 |
0.90 |
|
|
|
| 21 |
A |
1288 |
1233 |
0.33 |
|
|
|
| 22 |
A |
1258 |
1204 |
1.69 |
|
|
|
| 23 |
A |
1195 |
1143 |
0.25 |
|
|
|
| 24 |
A |
1105 |
1058 |
2.21 |
|
|
|
| 25 |
A |
1084 |
1038 |
5.00 |
|
|
|
| 26 |
A |
1041 |
996 |
9.65 |
|
|
|
| 27 |
A |
1014 |
971 |
1.62 |
|
|
|
| 28 |
A |
954 |
913 |
43.51 |
|
|
|
| 29 |
A |
943 |
903 |
0.27 |
|
|
|
| 30 |
A |
885 |
847 |
0.22 |
|
|
|
| 31 |
A |
875 |
838 |
3.33 |
|
|
|
| 32 |
A |
774 |
740 |
4.12 |
|
|
|
| 33 |
A |
647 |
619 |
9.33 |
|
|
|
| 34 |
A |
433 |
414 |
0.45 |
|
|
|
| 35 |
A |
383 |
367 |
1.31 |
|
|
|
| 36 |
A |
290 |
278 |
0.17 |
|
|
|
| 37 |
A |
231 |
221 |
0.02 |
|
|
|
| 38 |
A |
119 |
114 |
0.02 |
|
|
|
| 39 |
A |
91 |
87 |
0.05 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30162.1 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 28868.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-2.091 |
0.562 |
0.118 |
| C2 |
-1.175 |
-0.241 |
-0.397 |
| C3 |
-0.068 |
-0.890 |
0.363 |
| C4 |
1.319 |
-0.518 |
-0.160 |
| C5 |
1.655 |
0.950 |
0.026 |
| H6 |
-2.870 |
0.999 |
-0.490 |
| H7 |
-2.097 |
0.806 |
1.172 |
| H8 |
-1.206 |
-0.457 |
-1.461 |
| H9 |
-0.184 |
-1.974 |
0.300 |
| H10 |
-0.147 |
-0.625 |
1.419 |
| H11 |
2.061 |
-1.133 |
0.348 |
| H12 |
1.381 |
-0.779 |
-1.219 |
| H13 |
2.646 |
1.181 |
-0.361 |
| H14 |
0.937 |
1.587 |
-0.486 |
| H15 |
1.639 |
1.222 |
1.081 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
| C1 | | 1.3228 | 2.5023 | 3.5879 | 3.7681 | 1.0801 | 1.0821 | 2.0769 | 3.1784 | 2.6232 | 4.4913 | 3.9549 | 4.8016 | 3.2538 | 3.9090 |
C2 | 1.3228 | | 1.4912 | 2.5200 | 3.0995 | 2.1020 | 2.0998 | 1.0858 | 2.1146 | 2.1215 | 3.4387 | 2.7379 | 4.0768 | 2.7944 | 3.4988 | C3 | 2.5023 | 1.4912 | | 1.5283 | 2.5433 | 3.4849 | 2.7655 | 2.1929 | 1.0926 | 1.0912 | 2.1434 | 2.1480 | 3.4898 | 2.8046 | 2.8086 | C4 | 3.5879 | 2.5200 | 1.5283 | | 1.5173 | 4.4669 | 3.8983 | 2.8405 | 2.1429 | 2.1575 | 1.0902 | 1.0917 | 2.1652 | 2.1640 | 2.1611 | C5 | 3.7681 | 3.0995 | 2.5433 | 1.5173 | | 4.5548 | 3.9261 | 3.5178 | 3.4656 | 2.7696 | 2.1463 | 2.1478 | 1.0882 | 1.0883 | 1.0897 | H6 | 1.0801 | 2.1020 | 3.4849 | 4.4669 | 4.5548 | | 1.8432 | 2.4142 | 4.0836 | 3.7006 | 5.4373 | 4.6646 | 5.5201 | 3.8519 | 4.7800 | H7 | 1.0821 | 2.0998 | 2.7655 | 3.8983 | 3.9261 | 1.8432 | | 3.0530 | 3.4860 | 2.4316 | 4.6618 | 4.5082 | 4.9985 | 3.5444 | 3.7602 | H8 | 2.0769 | 1.0858 | 2.1929 | 2.8405 | 3.5178 | 2.4142 | 3.0530 | | 2.5390 | 3.0726 | 3.7954 | 2.6180 | 4.3274 | 3.1174 | 4.1679 | H9 | 3.1784 | 2.1146 | 1.0926 | 2.1429 | 3.4656 | 4.0836 | 3.4860 | 2.5390 | | 1.7531 | 2.3987 | 2.4869 | 4.2899 | 3.8156 | 3.7616 | H10 | 2.6232 | 2.1215 | 1.0912 | 2.1575 | 2.7696 | 3.7006 | 2.4316 | 3.0726 | 1.7531 | | 2.5065 | 3.0520 | 3.7726 | 3.1142 | 2.5916 | H11 | 4.4913 | 3.4387 | 2.1434 | 1.0902 | 2.1463 | 5.4373 | 4.6618 | 3.7954 | 2.3987 | 2.5065 | | 1.7446 | 2.4899 | 3.0593 | 2.5020 | H12 | 3.9549 | 2.7379 | 2.1480 | 1.0917 | 2.1478 | 4.6646 | 4.5082 | 2.6180 | 2.4869 | 3.0520 | 1.7446 | | 2.4854 | 2.5163 | 3.0591 | H13 | 4.8016 | 4.0768 | 3.4898 | 2.1652 | 1.0882 | 5.5201 | 4.9985 | 4.3274 | 4.2899 | 3.7726 | 2.4899 | 2.4854 | | 1.7609 | 1.7594 | H14 | 3.2538 | 2.7944 | 2.8046 | 2.1640 | 1.0883 | 3.8519 | 3.5444 | 3.1174 | 3.8156 | 3.1142 | 3.0593 | 2.5163 | 1.7609 | | 1.7557 | H15 | 3.9090 | 3.4988 | 2.8086 | 2.1611 | 1.0897 | 4.7800 | 3.7602 | 4.1679 | 3.7616 | 2.5916 | 2.5020 | 3.0591 | 1.7594 | 1.7557 | |
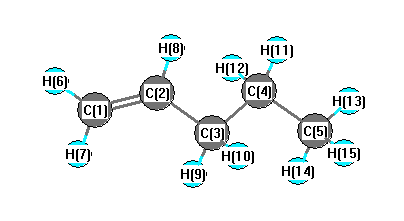 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
125.444 |
|
C1 |
C2 |
H8 |
118.824 |
| C2 |
C1 |
H6 |
121.708 |
|
C2 |
C1 |
H7 |
121.331 |
| C2 |
C3 |
C4 |
113.137 |
|
C2 |
C3 |
H9 |
108.872 |
| C2 |
C3 |
H10 |
109.500 |
|
C3 |
C2 |
H8 |
115.731 |
| C3 |
C4 |
C5 |
113.244 |
|
C3 |
C4 |
H11 |
108.729 |
| C3 |
C4 |
H12 |
108.998 |
|
C4 |
C3 |
H9 |
108.556 |
| C4 |
C3 |
H10 |
109.773 |
|
C4 |
C5 |
H13 |
111.335 |
| C4 |
C5 |
H14 |
111.231 |
|
C4 |
C5 |
H15 |
110.912 |
| C5 |
C4 |
H11 |
109.709 |
|
C5 |
C4 |
H12 |
109.736 |
| H6 |
C1 |
H7 |
116.960 |
|
H9 |
C3 |
H10 |
106.797 |
| H11 |
C4 |
H12 |
106.176 |
|
H13 |
C5 |
H14 |
108.004 |
| H13 |
C5 |
H15 |
107.763 |
|
H14 |
C5 |
H15 |
107.426 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.328 |
|
|
|
| 2 |
C |
-0.110 |
|
|
|
| 3 |
C |
-0.201 |
|
|
|
| 4 |
C |
-0.212 |
|
|
|
| 5 |
C |
-0.359 |
|
|
|
| 6 |
H |
0.132 |
|
|
|
| 7 |
H |
0.126 |
|
|
|
| 8 |
H |
0.138 |
|
|
|
| 9 |
H |
0.125 |
|
|
|
| 10 |
H |
0.109 |
|
|
|
| 11 |
H |
0.116 |
|
|
|
| 12 |
H |
0.112 |
|
|
|
| 13 |
H |
0.119 |
|
|
|
| 14 |
H |
0.121 |
|
|
|
| 15 |
H |
0.111 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.326 |
-0.243 |
-0.030 |
0.407 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.966 |
-1.235 |
0.324 |
| y |
-1.235 |
-34.023 |
0.402 |
| z |
0.324 |
0.402 |
-32.188 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.861 |
-1.235 |
0.324 |
| y |
-1.235 |
-0.946 |
0.402 |
| z |
0.324 |
0.402 |
1.807 |
|
| Polar |
|---|
| 3z2-r2 | 3.613 |
|---|
| x2-y2 | 0.057 |
|---|
| xy | -1.235 |
|---|
| xz | 0.324 |
|---|
| yz | 0.402 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.324 |
-1.099 |
-0.276 |
| y |
-1.099 |
8.518 |
0.365 |
| z |
-0.276 |
0.365 |
8.056 |
<r2> (average value of r
2) Å
2
| <r2> |
151.579 |
| (<r2>)1/2 |
12.312 |
