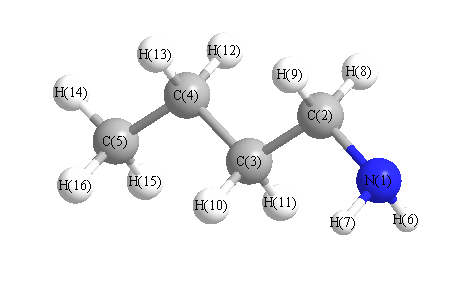
Vibrational Frequencies calculated at mPW1PW91/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3525 |
3381 |
0.93 |
|
|
|
| 2 |
A' |
3120 |
2993 |
37.59 |
|
|
|
| 3 |
A' |
3045 |
2921 |
44.86 |
|
|
|
| 4 |
A' |
3042 |
2918 |
75.51 |
|
|
|
| 5 |
A' |
3034 |
2910 |
2.54 |
|
|
|
| 6 |
A' |
3018 |
2895 |
11.58 |
|
|
|
| 7 |
A' |
1666 |
1598 |
22.14 |
|
|
|
| 8 |
A' |
1511 |
1449 |
6.73 |
|
|
|
| 9 |
A' |
1500 |
1439 |
0.64 |
|
|
|
| 10 |
A' |
1490 |
1429 |
0.45 |
|
|
|
| 11 |
A' |
1483 |
1422 |
0.23 |
|
|
|
| 12 |
A' |
1413 |
1356 |
2.21 |
|
|
|
| 13 |
A' |
1406 |
1349 |
8.08 |
|
|
|
| 14 |
A' |
1381 |
1325 |
3.46 |
|
|
|
| 15 |
A' |
1290 |
1237 |
2.96 |
|
|
|
| 16 |
A' |
1153 |
1106 |
5.68 |
|
|
|
| 17 |
A' |
1108 |
1063 |
13.24 |
|
|
|
| 18 |
A' |
1080 |
1036 |
3.74 |
|
|
|
| 19 |
A' |
1028 |
986 |
18.43 |
|
|
|
| 20 |
A' |
920 |
882 |
27.45 |
|
|
|
| 21 |
A' |
856 |
821 |
163.22 |
|
|
|
| 22 |
A' |
426 |
409 |
5.17 |
|
|
|
| 23 |
A' |
390 |
374 |
0.59 |
|
|
|
| 24 |
A' |
176 |
169 |
2.08 |
|
|
|
| 25 |
A" |
3605 |
3458 |
0.18 |
|
|
|
| 26 |
A" |
3114 |
2987 |
57.31 |
|
|
|
| 27 |
A" |
3085 |
2959 |
62.95 |
|
|
|
| 28 |
A" |
3067 |
2941 |
1.25 |
|
|
|
| 29 |
A" |
3044 |
2919 |
1.27 |
|
|
|
| 30 |
A" |
1499 |
1438 |
7.50 |
|
|
|
| 31 |
A" |
1393 |
1337 |
0.60 |
|
|
|
| 32 |
A" |
1334 |
1280 |
0.33 |
|
|
|
| 33 |
A" |
1311 |
1257 |
0.00 |
|
|
|
| 34 |
A" |
1231 |
1181 |
0.01 |
|
|
|
| 35 |
A" |
1050 |
1007 |
0.41 |
|
|
|
| 36 |
A" |
920 |
882 |
0.03 |
|
|
|
| 37 |
A" |
785 |
753 |
0.29 |
|
|
|
| 38 |
A" |
735 |
705 |
4.12 |
|
|
|
| 39 |
A" |
277 |
266 |
41.55 |
|
|
|
| 40 |
A" |
245 |
235 |
0.11 |
|
|
|
| 41 |
A" |
123 |
118 |
0.27 |
|
|
|
| 42 |
A" |
109 |
105 |
2.63 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 32993.9 cm
-1
Scaled (by 0.9592) Zero Point Vibrational Energy (zpe) 31647.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.303 |
|
|
|
| 2 |
C |
-0.062 |
|
|
|
| 3 |
C |
-0.149 |
|
|
|
| 4 |
C |
-0.150 |
|
|
|
| 5 |
C |
-0.267 |
|
|
|
| 6 |
H |
0.115 |
|
|
|
| 7 |
H |
0.115 |
|
|
|
| 8 |
H |
0.080 |
|
|
|
| 9 |
H |
0.080 |
|
|
|
| 10 |
H |
0.067 |
|
|
|
| 11 |
H |
0.067 |
|
|
|
| 12 |
H |
0.077 |
|
|
|
| 13 |
H |
0.077 |
|
|
|
| 14 |
H |
0.088 |
|
|
|
| 15 |
H |
0.083 |
|
|
|
| 16 |
H |
0.083 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.009 |
0.756 |
0.000 |
1.260 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.331 |
2.903 |
0.000 |
| y |
2.903 |
-32.996 |
0.000 |
| z |
0.000 |
0.000 |
-32.033 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-7.817 |
2.903 |
0.000 |
| y |
2.903 |
3.187 |
0.000 |
| z |
0.000 |
0.000 |
4.630 |
|
| Polar |
|---|
| 3z2-r2 | 9.260 |
|---|
| x2-y2 | -7.336 |
|---|
| xy | 2.903 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.062 |
0.245 |
0.000 |
| y |
0.245 |
8.198 |
0.000 |
| z |
0.000 |
0.000 |
7.714 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |