return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CH2CH2CN (Butanenitrile)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-211.009506
Energy at 298.15K-211.016837
HF Energy-210.067743
Nuclear repulsion energy154.948332
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3175 3015 17.99      
2 A' 3114 2957 18.56      
3 A' 3100 2943 0.30      
4 A' 3092 2936 15.39      
5 A' 2227 2114 0.17      
6 A' 1533 1456 7.70      
7 A' 1519 1442 1.34      
8 A' 1492 1417 3.40      
9 A' 1430 1357 3.10      
10 A' 1401 1330 1.97      
11 A' 1313 1247 1.32      
12 A' 1137 1079 2.31      
13 A' 1088 1033 0.14      
14 A' 983 933 1.48      
15 A' 898 853 1.69      
16 A' 530 503 0.50      
17 A' 347 329 0.03      
18 A' 163 155 5.75      
19 A" 3167 3007 31.80      
20 A" 3148 2989 0.16      
21 A" 3129 2971 0.00      
22 A" 1529 1452 8.37      
23 A" 1349 1281 0.02      
24 A" 1277 1213 0.03      
25 A" 1140 1083 0.06      
26 A" 883 838 0.21      
27 A" 756 718 3.10      
28 A" 382 362 0.30      
29 A" 260 247 0.01      
30 A" 105 99 4.16      

Unscaled Zero Point Vibrational Energy (zpe) 22833.2 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 21677.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
0.80448 0.07645 0.07260

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 -2.602 0.334 0.000
C2 -1.444 0.481 0.000
C3 0.000 0.630 0.000
C4 0.720 -0.713 0.000
C5 2.223 -0.526 0.000
H6 0.287 1.208 0.875
H7 0.287 1.208 -0.875
H8 0.409 -1.280 0.872
H9 0.409 -1.280 -0.872
H10 2.737 -1.481 0.000
H11 2.545 0.027 0.878
H12 2.545 0.027 -0.878

Atom - Atom Distances (Å)
  N1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12
N11.16652.61843.48304.90123.14233.14233.52573.52575.63865.22995.2299
C21.16651.45212.47213.80352.07182.07182.70132.70134.61854.10984.1098
C32.61841.45211.52412.50581.08701.08702.13932.13933.45612.75872.7587
C43.48302.47211.52411.51522.15512.15511.08581.08582.15782.15622.1562
C54.90123.80352.50581.51522.74222.74222.15022.15021.08431.08621.0862
H63.14232.07181.08702.15512.74221.74922.49133.04273.74102.54783.0924
H73.14232.07181.08702.15512.74221.74923.04272.49133.74103.09242.5478
H83.52572.70132.13931.08582.15022.49133.04271.74442.49402.50443.0554
H93.52572.70132.13931.08582.15023.04272.49131.74442.49403.05542.5044
H105.63864.61853.45612.15781.08433.74103.74102.49402.49401.75551.7555
H115.22994.10982.75872.15621.08622.54783.09242.50443.05541.75551.7561
H125.22994.10982.75872.15621.08623.09242.54783.05542.50441.75551.7561

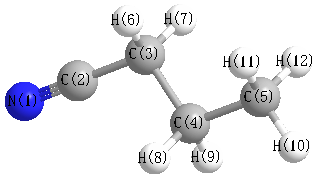
picture of Butanenitrile state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C2 C3 178.668 C2 C3 C4 112.299
C2 C3 H6 108.505 C2 C3 H7 108.505
C3 C4 C5 111.070 C3 C4 H8 108.951
C3 C4 H9 108.951 C4 C3 H6 110.118
C4 C3 H7 110.118 C4 C5 H10 111.131
C4 C5 H11 110.892 C4 C5 H12 110.892
C5 C4 H8 110.433 C5 C4 H9 110.433
H6 C3 H7 107.138 H8 C4 H9 106.892
H10 C5 H11 107.955 H10 C5 H12 107.955
H11 C5 H12 107.881
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability