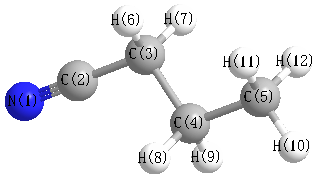
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3139 |
3004 |
23.89 |
|
|
|
| 2 |
A' |
3078 |
2946 |
21.59 |
|
|
|
| 3 |
A' |
3064 |
2933 |
0.59 |
|
|
|
| 4 |
A' |
3056 |
2925 |
19.55 |
|
|
|
| 5 |
A' |
2389 |
2286 |
13.81 |
|
|
|
| 6 |
A' |
1509 |
1444 |
8.21 |
|
|
|
| 7 |
A' |
1494 |
1430 |
1.28 |
|
|
|
| 8 |
A' |
1469 |
1406 |
3.76 |
|
|
|
| 9 |
A' |
1418 |
1357 |
2.80 |
|
|
|
| 10 |
A' |
1386 |
1326 |
2.49 |
|
|
|
| 11 |
A' |
1300 |
1245 |
1.59 |
|
|
|
| 12 |
A' |
1120 |
1072 |
2.77 |
|
|
|
| 13 |
A' |
1067 |
1022 |
0.28 |
|
|
|
| 14 |
A' |
974 |
933 |
1.47 |
|
|
|
| 15 |
A' |
888 |
850 |
1.88 |
|
|
|
| 16 |
A' |
539 |
515 |
0.77 |
|
|
|
| 17 |
A' |
342 |
328 |
0.07 |
|
|
|
| 18 |
A' |
164 |
156 |
6.26 |
|
|
|
| 19 |
A" |
3134 |
3000 |
42.43 |
|
|
|
| 20 |
A" |
3114 |
2980 |
0.31 |
|
|
|
| 21 |
A" |
3096 |
2963 |
0.28 |
|
|
|
| 22 |
A" |
1501 |
1436 |
8.37 |
|
|
|
| 23 |
A" |
1328 |
1271 |
0.01 |
|
|
|
| 24 |
A" |
1258 |
1204 |
0.03 |
|
|
|
| 25 |
A" |
1127 |
1078 |
0.16 |
|
|
|
| 26 |
A" |
871 |
834 |
0.15 |
|
|
|
| 27 |
A" |
742 |
710 |
4.11 |
|
|
|
| 28 |
A" |
392 |
375 |
0.40 |
|
|
|
| 29 |
A" |
246 |
236 |
0.02 |
|
|
|
| 30 |
A" |
98 |
94 |
4.52 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22650.8 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 21679.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.075 |
|
|
|
| 2 |
C |
-0.084 |
|
|
|
| 3 |
C |
-0.199 |
|
|
|
| 4 |
C |
-0.200 |
|
|
|
| 5 |
C |
-0.362 |
|
|
|
| 6 |
H |
0.147 |
|
|
|
| 7 |
H |
0.147 |
|
|
|
| 8 |
H |
0.132 |
|
|
|
| 9 |
H |
0.132 |
|
|
|
| 10 |
H |
0.130 |
|
|
|
| 11 |
H |
0.116 |
|
|
|
| 12 |
H |
0.116 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
4.175 |
0.022 |
0.000 |
4.175 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-42.616 |
0.978 |
0.000 |
| y |
0.978 |
-30.014 |
0.000 |
| z |
0.000 |
0.000 |
-30.214 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-12.502 |
0.978 |
0.000 |
| y |
0.978 |
6.401 |
0.000 |
| z |
0.000 |
0.000 |
6.101 |
|
| Polar |
|---|
| 3z2-r2 | 12.201 |
|---|
| x2-y2 | -12.602 |
|---|
| xy | 0.978 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.824 |
-0.345 |
0.000 |
| y |
-0.345 |
6.551 |
0.000 |
| z |
0.000 |
0.000 |
6.073 |
<r2> (average value of r
2) Å
2
| <r2> |
157.572 |
| (<r2>)1/2 |
12.553 |