Jump to
S1C2
Vibrational Frequencies calculated at B1B95/cc-pVTZ
Geometric Data calculated at B1B95/cc-pVTZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -269.517566 |
| Energy at 298.15K | -269.526945 |
| Nuclear repulsion energy | 194.477277 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3153 |
3018 |
19.85 |
|
|
|
| 2 |
A1 |
2997 |
2869 |
63.63 |
|
|
|
| 3 |
A1 |
2896 |
2772 |
63.86 |
|
|
|
| 4 |
A1 |
1550 |
1484 |
0.18 |
|
|
|
| 5 |
A1 |
1507 |
1442 |
4.64 |
|
|
|
| 6 |
A1 |
1474 |
1411 |
0.40 |
|
|
|
| 7 |
A1 |
1283 |
1228 |
15.82 |
|
|
|
| 8 |
A1 |
1188 |
1137 |
13.95 |
|
|
|
| 9 |
A1 |
1061 |
1016 |
54.98 |
|
|
|
| 10 |
A1 |
485 |
464 |
1.50 |
|
|
|
| 11 |
A1 |
211 |
202 |
1.87 |
|
|
|
| 12 |
A2 |
3044 |
2913 |
0.00 |
|
|
|
| 13 |
A2 |
1489 |
1425 |
0.00 |
|
|
|
| 14 |
A2 |
1249 |
1195 |
0.00 |
|
|
|
| 15 |
A2 |
1181 |
1130 |
0.00 |
|
|
|
| 16 |
A2 |
209 |
200 |
0.00 |
|
|
|
| 17 |
A2 |
76 |
72 |
0.00 |
|
|
|
| 18 |
B1 |
3044 |
2914 |
118.48 |
|
|
|
| 19 |
B1 |
2915 |
2790 |
124.00 |
|
|
|
| 20 |
B1 |
1490 |
1426 |
15.35 |
|
|
|
| 21 |
B1 |
1200 |
1149 |
21.84 |
|
|
|
| 22 |
B1 |
1151 |
1102 |
5.09 |
|
|
|
| 23 |
B1 |
219 |
210 |
4.68 |
|
|
|
| 24 |
B1 |
84 |
80 |
0.62 |
|
|
|
| 25 |
B2 |
3153 |
3017 |
24.77 |
|
|
|
| 26 |
B2 |
2996 |
2867 |
64.82 |
|
|
|
| 27 |
B2 |
1516 |
1451 |
0.72 |
|
|
|
| 28 |
B2 |
1491 |
1427 |
27.85 |
|
|
|
| 29 |
B2 |
1437 |
1376 |
55.83 |
|
|
|
| 30 |
B2 |
1236 |
1183 |
305.62 |
|
|
|
| 31 |
B2 |
1181 |
1131 |
159.15 |
|
|
|
| 32 |
B2 |
1039 |
995 |
2.53 |
|
|
|
| 33 |
B2 |
414 |
397 |
2.95 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24809.8 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 23745.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.300 |
| H2 |
-0.894 |
0.000 |
0.949 |
| H3 |
0.894 |
0.000 |
0.949 |
| O4 |
0.000 |
1.108 |
-0.528 |
| O5 |
0.000 |
-1.108 |
-0.528 |
| C6 |
0.000 |
2.303 |
0.202 |
| C7 |
0.000 |
-2.303 |
0.202 |
| H8 |
0.000 |
3.119 |
-0.513 |
| H9 |
0.000 |
-3.119 |
-0.513 |
| H10 |
-0.888 |
2.388 |
0.837 |
| H11 |
0.888 |
2.388 |
0.837 |
| H12 |
0.888 |
-2.388 |
0.837 |
| H13 |
-0.888 |
-2.388 |
0.837 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
H3 |
O4 |
O5 |
C6 |
C7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
| C1 | | 1.1049 | 1.1049 | 1.3831 | 1.3831 | 2.3049 | 2.3049 | 3.2230 | 3.2230 | 2.6040 | 2.6040 | 2.6040 | 2.6040 |
H2 | 1.1049 | | 1.7878 | 2.0516 | 2.0516 | 2.5806 | 2.5806 | 3.5586 | 3.5586 | 2.3909 | 2.9818 | 2.9818 | 2.3909 | H3 | 1.1049 | 1.7878 | | 2.0516 | 2.0516 | 2.5806 | 2.5806 | 3.5586 | 3.5586 | 2.9818 | 2.3909 | 2.3909 | 2.9818 | O4 | 1.3831 | 2.0516 | 2.0516 | | 2.2160 | 1.4004 | 3.4881 | 2.0110 | 4.2269 | 2.0715 | 2.0715 | 3.8569 | 3.8569 | O5 | 1.3831 | 2.0516 | 2.0516 | 2.2160 | | 3.4881 | 1.4004 | 4.2269 | 2.0110 | 3.8569 | 3.8569 | 2.0715 | 2.0715 | C6 | 2.3049 | 2.5806 | 2.5806 | 1.4004 | 3.4881 | | 4.6056 | 1.0851 | 5.4687 | 1.0948 | 1.0948 | 4.8164 | 4.8164 | C7 | 2.3049 | 2.5806 | 2.5806 | 3.4881 | 1.4004 | 4.6056 | | 5.4687 | 1.0851 | 4.8164 | 4.8164 | 1.0948 | 1.0948 | H8 | 3.2230 | 3.5586 | 3.5586 | 2.0110 | 4.2269 | 1.0851 | 5.4687 | | 6.2378 | 1.7733 | 1.7733 | 5.7393 | 5.7393 | H9 | 3.2230 | 3.5586 | 3.5586 | 4.2269 | 2.0110 | 5.4687 | 1.0851 | 6.2378 | | 5.7393 | 5.7393 | 1.7733 | 1.7733 | H10 | 2.6040 | 2.3909 | 2.9818 | 2.0715 | 3.8569 | 1.0948 | 4.8164 | 1.7733 | 5.7393 | | 1.7759 | 5.0960 | 4.7765 | H11 | 2.6040 | 2.9818 | 2.3909 | 2.0715 | 3.8569 | 1.0948 | 4.8164 | 1.7733 | 5.7393 | 1.7759 | | 4.7765 | 5.0960 | H12 | 2.6040 | 2.9818 | 2.3909 | 3.8569 | 2.0715 | 4.8164 | 1.0948 | 5.7393 | 1.7733 | 5.0960 | 4.7765 | | 1.7759 | H13 | 2.6040 | 2.3909 | 2.9818 | 3.8569 | 2.0715 | 4.8164 | 1.0948 | 5.7393 | 1.7733 | 4.7765 | 5.0960 | 1.7759 | |
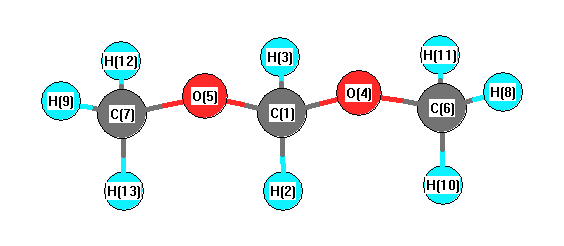 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O4 |
C6 |
111.795 |
|
C1 |
O5 |
C7 |
111.795 |
| H2 |
C1 |
H3 |
108.006 |
|
H2 |
C1 |
O4 |
110.597 |
| H2 |
C1 |
O5 |
110.597 |
|
H3 |
C1 |
O4 |
110.597 |
| H3 |
C1 |
O5 |
110.597 |
|
O4 |
C1 |
O5 |
106.467 |
| O4 |
C6 |
H8 |
107.335 |
|
O4 |
C6 |
H10 |
111.653 |
| O4 |
C6 |
H11 |
111.653 |
|
O5 |
C7 |
H9 |
107.335 |
| O5 |
C7 |
H12 |
111.653 |
|
O5 |
C7 |
H13 |
111.653 |
| H8 |
C6 |
H10 |
108.868 |
|
H8 |
C6 |
H11 |
108.868 |
| H9 |
C7 |
H12 |
108.868 |
|
H9 |
C7 |
H13 |
108.868 |
| H10 |
C6 |
H11 |
108.395 |
|
H12 |
C7 |
H13 |
108.395 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.174 |
|
|
|
| 2 |
H |
0.020 |
|
|
|
| 3 |
H |
0.020 |
|
|
|
| 4 |
O |
-0.226 |
|
|
|
| 5 |
O |
-0.226 |
|
|
|
| 6 |
C |
-0.164 |
|
|
|
| 7 |
C |
-0.164 |
|
|
|
| 8 |
H |
0.127 |
|
|
|
| 9 |
H |
0.127 |
|
|
|
| 10 |
H |
0.078 |
|
|
|
| 11 |
H |
0.078 |
|
|
|
| 12 |
H |
0.078 |
|
|
|
| 13 |
H |
0.078 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
2.272 |
2.272 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.801 |
0.000 |
0.000 |
| y |
0.000 |
-25.152 |
0.000 |
| z |
0.000 |
0.000 |
-33.406 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.521 |
0.000 |
0.000 |
| y |
0.000 |
7.452 |
0.000 |
| z |
0.000 |
0.000 |
-4.930 |
|
| Polar |
|---|
| 3z2-r2 | -9.860 |
|---|
| x2-y2 | -6.649 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.154 |
0.000 |
0.000 |
| y |
0.000 |
8.793 |
0.000 |
| z |
0.000 |
0.000 |
6.139 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |
