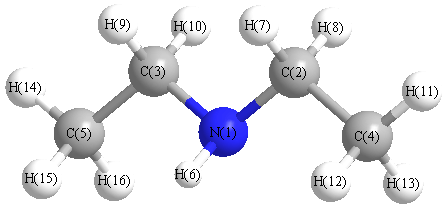
Vibrational Frequencies calculated at mPW1PW91/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3541 |
3397 |
0.07 |
|
|
|
| 2 |
A' |
3137 |
3009 |
39.96 |
|
|
|
| 3 |
A' |
3118 |
2991 |
63.46 |
|
|
|
| 4 |
A' |
3054 |
2930 |
63.34 |
|
|
|
| 5 |
A' |
3049 |
2924 |
3.43 |
|
|
|
| 6 |
A' |
2938 |
2818 |
172.72 |
|
|
|
| 7 |
A' |
1529 |
1467 |
2.33 |
|
|
|
| 8 |
A' |
1501 |
1439 |
2.44 |
|
|
|
| 9 |
A' |
1490 |
1430 |
14.70 |
|
|
|
| 10 |
A' |
1432 |
1373 |
3.46 |
|
|
|
| 11 |
A' |
1398 |
1341 |
1.24 |
|
|
|
| 12 |
A' |
1321 |
1268 |
3.91 |
|
|
|
| 13 |
A' |
1236 |
1185 |
1.72 |
|
|
|
| 14 |
A' |
1173 |
1125 |
13.76 |
|
|
|
| 15 |
A' |
1075 |
1032 |
7.35 |
|
|
|
| 16 |
A' |
905 |
868 |
4.76 |
|
|
|
| 17 |
A' |
827 |
793 |
1.21 |
|
|
|
| 18 |
A' |
757 |
726 |
75.27 |
|
|
|
| 19 |
A' |
426 |
408 |
0.32 |
|
|
|
| 20 |
A' |
254 |
244 |
0.76 |
|
|
|
| 21 |
A' |
183 |
175 |
0.82 |
|
|
|
| 22 |
A' |
109 |
104 |
1.04 |
|
|
|
| 23 |
A" |
3137 |
3009 |
11.99 |
|
|
|
| 24 |
A" |
3118 |
2990 |
13.83 |
|
|
|
| 25 |
A" |
3053 |
2929 |
10.02 |
|
|
|
| 26 |
A" |
3047 |
2923 |
20.33 |
|
|
|
| 27 |
A" |
2934 |
2815 |
15.18 |
|
|
|
| 28 |
A" |
1518 |
1456 |
7.94 |
|
|
|
| 29 |
A" |
1510 |
1448 |
12.23 |
|
|
|
| 30 |
A" |
1497 |
1436 |
8.51 |
|
|
|
| 31 |
A" |
1480 |
1420 |
12.61 |
|
|
|
| 32 |
A" |
1410 |
1353 |
13.98 |
|
|
|
| 33 |
A" |
1360 |
1305 |
24.02 |
|
|
|
| 34 |
A" |
1288 |
1235 |
3.95 |
|
|
|
| 35 |
A" |
1186 |
1137 |
52.26 |
|
|
|
| 36 |
A" |
1118 |
1072 |
8.18 |
|
|
|
| 37 |
A" |
1073 |
1029 |
2.71 |
|
|
|
| 38 |
A" |
954 |
915 |
0.19 |
|
|
|
| 39 |
A" |
809 |
776 |
0.45 |
|
|
|
| 40 |
A" |
425 |
408 |
0.47 |
|
|
|
| 41 |
A" |
254 |
243 |
0.69 |
|
|
|
| 42 |
A" |
116 |
111 |
0.85 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 32868.6 cm
-1
Scaled (by 0.9592) Zero Point Vibrational Energy (zpe) 31527.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.212 |
|
|
|
| 2 |
C |
-0.038 |
|
|
|
| 3 |
C |
-0.038 |
|
|
|
| 4 |
C |
-0.263 |
|
|
|
| 5 |
C |
-0.263 |
|
|
|
| 6 |
H |
0.091 |
|
|
|
| 7 |
H |
0.041 |
|
|
|
| 8 |
H |
0.072 |
|
|
|
| 9 |
H |
0.041 |
|
|
|
| 10 |
H |
0.072 |
|
|
|
| 11 |
H |
0.082 |
|
|
|
| 12 |
H |
0.076 |
|
|
|
| 13 |
H |
0.091 |
|
|
|
| 14 |
H |
0.082 |
|
|
|
| 15 |
H |
0.076 |
|
|
|
| 16 |
H |
0.091 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.733 |
0.344 |
0.000 |
0.809 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.628 |
1.630 |
0.000 |
| y |
1.630 |
-34.441 |
0.000 |
| z |
0.000 |
0.000 |
-33.351 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.268 |
1.630 |
0.000 |
| y |
1.630 |
-0.952 |
0.000 |
| z |
0.000 |
0.000 |
0.683 |
|
| Polar |
|---|
| 3z2-r2 | 1.367 |
|---|
| x2-y2 | 0.813 |
|---|
| xy | 1.630 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.681 |
-0.021 |
0.000 |
| y |
-0.021 |
8.291 |
0.000 |
| z |
0.000 |
0.000 |
10.559 |
<r2> (average value of r
2) Å
2
| <r2> |
185.959 |
| (<r2>)1/2 |
13.637 |