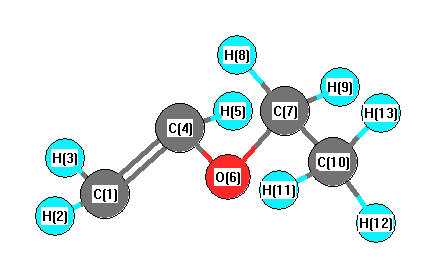
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3288 |
3147 |
5.43 |
|
|
|
| 2 |
A |
3191 |
3055 |
0.01 |
|
|
|
| 3 |
A |
3151 |
3016 |
24.63 |
|
|
|
| 4 |
A |
3149 |
3014 |
27.63 |
|
|
|
| 5 |
A |
3148 |
3013 |
13.62 |
|
|
|
| 6 |
A |
3070 |
2938 |
15.11 |
|
|
|
| 7 |
A |
3057 |
2926 |
36.81 |
|
|
|
| 8 |
A |
3020 |
2890 |
53.23 |
|
|
|
| 9 |
A |
1735 |
1660 |
182.92 |
|
|
|
| 10 |
A |
1526 |
1460 |
7.29 |
|
|
|
| 11 |
A |
1501 |
1436 |
2.30 |
|
|
|
| 12 |
A |
1482 |
1419 |
6.21 |
|
|
|
| 13 |
A |
1445 |
1383 |
2.86 |
|
|
|
| 14 |
A |
1418 |
1357 |
62.70 |
|
|
|
| 15 |
A |
1393 |
1334 |
24.66 |
|
|
|
| 16 |
A |
1346 |
1288 |
7.88 |
|
|
|
| 17 |
A |
1307 |
1251 |
0.79 |
|
|
|
| 18 |
A |
1245 |
1191 |
266.53 |
|
|
|
| 19 |
A |
1182 |
1131 |
4.52 |
|
|
|
| 20 |
A |
1169 |
1118 |
125.50 |
|
|
|
| 21 |
A |
1107 |
1060 |
11.35 |
|
|
|
| 22 |
A |
996 |
953 |
5.50 |
|
|
|
| 23 |
A |
989 |
947 |
18.01 |
|
|
|
| 24 |
A |
864 |
827 |
4.33 |
|
|
|
| 25 |
A |
858 |
822 |
53.78 |
|
|
|
| 26 |
A |
818 |
783 |
1.82 |
|
|
|
| 27 |
A |
712 |
682 |
5.51 |
|
|
|
| 28 |
A |
498 |
477 |
2.85 |
|
|
|
| 29 |
A |
434 |
415 |
1.44 |
|
|
|
| 30 |
A |
246 |
235 |
0.51 |
|
|
|
| 31 |
A |
205 |
196 |
0.21 |
|
|
|
| 32 |
A |
88 |
84 |
2.60 |
|
|
|
| 33 |
A |
36 |
35 |
0.09 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24835.3 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 23769.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.385 |
|
|
|
| 2 |
H |
0.133 |
|
|
|
| 3 |
H |
0.135 |
|
|
|
| 4 |
C |
0.060 |
|
|
|
| 5 |
H |
0.096 |
|
|
|
| 6 |
O |
-0.202 |
|
|
|
| 7 |
C |
-0.025 |
|
|
|
| 8 |
H |
0.090 |
|
|
|
| 9 |
H |
0.089 |
|
|
|
| 10 |
C |
-0.344 |
|
|
|
| 11 |
H |
0.123 |
|
|
|
| 12 |
H |
0.123 |
|
|
|
| 13 |
H |
0.108 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.189 |
1.258 |
0.001 |
1.731 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-29.792 |
-0.017 |
0.004 |
| y |
-0.017 |
-30.515 |
-0.012 |
| z |
0.004 |
-0.012 |
-33.200 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.066 |
-0.017 |
0.004 |
| y |
-0.017 |
0.981 |
-0.012 |
| z |
0.004 |
-0.012 |
-3.046 |
|
| Polar |
|---|
| 3z2-r2 | -6.093 |
|---|
| x2-y2 | 0.723 |
|---|
| xy | -0.017 |
|---|
| xz | 0.004 |
|---|
| yz | -0.012 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.056 |
0.742 |
-0.001 |
| y |
0.742 |
7.331 |
-0.003 |
| z |
-0.001 |
-0.003 |
5.853 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |