Jump to
S1C2
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -231.171084 |
| Energy at 298.15K | -231.176919 |
| Nuclear repulsion energy | 159.864600 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3292 |
3151 |
4.72 |
|
|
|
| 2 |
A1 |
3197 |
3060 |
1.20 |
|
|
|
| 3 |
A1 |
3171 |
3035 |
23.02 |
|
|
|
| 4 |
A1 |
1763 |
1687 |
4.11 |
|
|
|
| 5 |
A1 |
1434 |
1373 |
0.75 |
|
|
|
| 6 |
A1 |
1355 |
1296 |
4.81 |
|
|
|
| 7 |
A1 |
1223 |
1171 |
17.20 |
|
|
|
| 8 |
A1 |
872 |
834 |
2.90 |
|
|
|
| 9 |
A1 |
524 |
501 |
0.43 |
|
|
|
| 10 |
A1 |
229 |
219 |
0.01 |
|
|
|
| 11 |
A2 |
976 |
934 |
0.00 |
|
|
|
| 12 |
A2 |
872 |
835 |
0.00 |
|
|
|
| 13 |
A2 |
723 |
692 |
0.00 |
|
|
|
| 14 |
A2 |
123 |
117 |
0.00 |
|
|
|
| 15 |
B1 |
989 |
947 |
38.94 |
|
|
|
| 16 |
B1 |
875 |
838 |
110.69 |
|
|
|
| 17 |
B1 |
700 |
670 |
9.29 |
|
|
|
| 18 |
B1 |
88 |
85 |
1.33 |
|
|
|
| 19 |
B2 |
3292 |
3151 |
1.17 |
|
|
|
| 20 |
B2 |
3196 |
3059 |
0.48 |
|
|
|
| 21 |
B2 |
3166 |
3030 |
5.56 |
|
|
|
| 22 |
B2 |
1711 |
1638 |
489.52 |
|
|
|
| 23 |
B2 |
1426 |
1365 |
74.18 |
|
|
|
| 24 |
B2 |
1334 |
1277 |
16.28 |
|
|
|
| 25 |
B2 |
1245 |
1191 |
450.01 |
|
|
|
| 26 |
B2 |
1035 |
990 |
3.86 |
|
|
|
| 27 |
B2 |
489 |
468 |
7.20 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19649.4 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 18806.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.000 |
0.000 |
0.331 |
| C2 |
0.000 |
1.162 |
-0.367 |
| C3 |
0.000 |
-1.162 |
-0.367 |
| C4 |
0.000 |
2.339 |
0.230 |
| C5 |
0.000 |
-2.339 |
0.230 |
| H6 |
0.000 |
1.049 |
-1.444 |
| H7 |
0.000 |
-1.049 |
-1.444 |
| H8 |
0.000 |
3.235 |
-0.367 |
| H9 |
0.000 |
2.427 |
1.305 |
| H10 |
0.000 |
-3.235 |
-0.367 |
| H11 |
0.000 |
-2.427 |
1.305 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| O1 | | 1.3557 | 1.3557 | 2.3416 | 2.3416 | 2.0625 | 2.0625 | 3.3097 | 2.6150 | 3.3097 | 2.6150 |
C2 | 1.3557 | | 2.3242 | 1.3199 | 3.5520 | 1.0836 | 2.4598 | 2.0731 | 2.0962 | 4.3973 | 3.9593 | C3 | 1.3557 | 2.3242 | | 3.5520 | 1.3199 | 2.4598 | 1.0836 | 4.3973 | 3.9593 | 2.0731 | 2.0962 | C4 | 2.3416 | 1.3199 | 3.5520 | | 4.6788 | 2.1140 | 3.7796 | 1.0765 | 1.0782 | 5.6064 | 4.8861 | C5 | 2.3416 | 3.5520 | 1.3199 | 4.6788 | | 3.7796 | 2.1140 | 5.6064 | 4.8861 | 1.0765 | 1.0782 | H6 | 2.0625 | 1.0836 | 2.4598 | 2.1140 | 3.7796 | | 2.0980 | 2.4373 | 3.0753 | 4.4176 | 4.4318 | H7 | 2.0625 | 2.4598 | 1.0836 | 3.7796 | 2.1140 | 2.0980 | | 4.4176 | 4.4318 | 2.4373 | 3.0753 | H8 | 3.3097 | 2.0731 | 4.3973 | 1.0765 | 5.6064 | 2.4373 | 4.4176 | | 1.8567 | 6.4703 | 5.9039 | H9 | 2.6150 | 2.0962 | 3.9593 | 1.0782 | 4.8861 | 3.0753 | 4.4318 | 1.8567 | | 5.9039 | 4.8542 | H10 | 3.3097 | 4.3973 | 2.0731 | 5.6064 | 1.0765 | 4.4176 | 2.4373 | 6.4703 | 5.9039 | | 1.8567 | H11 | 2.6150 | 3.9593 | 2.0962 | 4.8861 | 1.0782 | 4.4318 | 3.0753 | 5.9039 | 4.8542 | 1.8567 | |
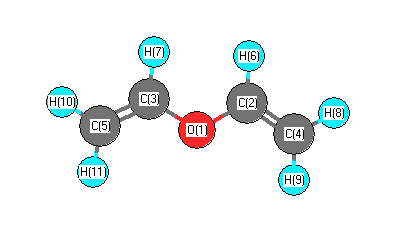 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C4 |
122.119 |
|
O1 |
C2 |
H6 |
115.005 |
| O1 |
C3 |
C5 |
122.119 |
|
O1 |
C3 |
H7 |
115.005 |
| C2 |
O1 |
C3 |
118.007 |
|
C2 |
C4 |
H8 |
119.435 |
| C2 |
C4 |
H9 |
121.550 |
|
C3 |
C5 |
H10 |
119.435 |
| C3 |
C5 |
H11 |
121.550 |
|
C4 |
C2 |
H6 |
122.876 |
| C5 |
C3 |
H7 |
122.876 |
|
H8 |
C4 |
H9 |
119.015 |
| H10 |
C5 |
H11 |
119.015 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.139 |
|
|
|
| 2 |
C |
0.056 |
|
|
|
| 3 |
C |
0.056 |
|
|
|
| 4 |
C |
-0.367 |
|
|
|
| 5 |
C |
-0.367 |
|
|
|
| 6 |
H |
0.100 |
|
|
|
| 7 |
H |
0.100 |
|
|
|
| 8 |
H |
0.140 |
|
|
|
| 9 |
H |
0.141 |
|
|
|
| 10 |
H |
0.140 |
|
|
|
| 11 |
H |
0.141 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-1.241 |
1.241 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.785 |
0.000 |
0.000 |
| y |
0.000 |
-27.780 |
0.000 |
| z |
0.000 |
0.000 |
-27.544 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.123 |
0.000 |
0.000 |
| y |
0.000 |
2.884 |
0.000 |
| z |
0.000 |
0.000 |
3.239 |
|
| Polar |
|---|
| 3z2-r2 | 6.477 |
|---|
| x2-y2 | -6.005 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.452 |
0.000 |
0.000 |
| y |
0.000 |
12.986 |
0.000 |
| z |
0.000 |
0.000 |
6.841 |
<r2> (average value of r
2) Å
2
| <r2> |
146.320 |
| (<r2>)1/2 |
12.096 |
Jump to
S1C1
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -231.170906 |
| Energy at 298.15K | -231.177042 |
| HF Energy | -231.170906 |
| Nuclear repulsion energy | 162.601252 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3289 |
3148 |
4.19 |
|
|
|
| 2 |
A |
3287 |
3146 |
5.40 |
|
|
|
| 3 |
A |
3222 |
3084 |
7.86 |
|
|
|
| 4 |
A |
3196 |
3059 |
2.89 |
|
|
|
| 5 |
A |
3193 |
3056 |
1.85 |
|
|
|
| 6 |
A |
3175 |
3039 |
7.19 |
|
|
|
| 7 |
A |
1735 |
1660 |
97.66 |
|
|
|
| 8 |
A |
1710 |
1636 |
206.41 |
|
|
|
| 9 |
A |
1441 |
1379 |
1.20 |
|
|
|
| 10 |
A |
1422 |
1361 |
22.83 |
|
|
|
| 11 |
A |
1369 |
1310 |
70.54 |
|
|
|
| 12 |
A |
1332 |
1275 |
6.53 |
|
|
|
| 13 |
A |
1263 |
1209 |
284.83 |
|
|
|
| 14 |
A |
1150 |
1101 |
15.42 |
|
|
|
| 15 |
A |
1022 |
978 |
40.30 |
|
|
|
| 16 |
A |
1015 |
971 |
2.36 |
|
|
|
| 17 |
A |
1000 |
957 |
29.88 |
|
|
|
| 18 |
A |
899 |
860 |
46.79 |
|
|
|
| 19 |
A |
888 |
849 |
19.19 |
|
|
|
| 20 |
A |
872 |
835 |
55.96 |
|
|
|
| 21 |
A |
743 |
711 |
6.90 |
|
|
|
| 22 |
A |
729 |
698 |
2.98 |
|
|
|
| 23 |
A |
593 |
568 |
2.70 |
|
|
|
| 24 |
A |
456 |
437 |
2.53 |
|
|
|
| 25 |
A |
296 |
283 |
1.32 |
|
|
|
| 26 |
A |
215 |
206 |
8.33 |
|
|
|
| 27 |
A |
97 |
93 |
0.25 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19803.3 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 18953.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
-0.044 |
-0.789 |
0.049 |
| C2 |
1.279 |
-0.492 |
0.006 |
| C3 |
-0.918 |
0.226 |
0.288 |
| C4 |
1.823 |
0.704 |
-0.146 |
| C5 |
-2.149 |
0.211 |
-0.188 |
| H6 |
1.866 |
-1.393 |
0.101 |
| H7 |
-0.535 |
1.006 |
0.935 |
| H8 |
2.897 |
0.791 |
-0.152 |
| H9 |
1.240 |
1.599 |
-0.287 |
| H10 |
-2.836 |
0.994 |
0.086 |
| H11 |
-2.490 |
-0.580 |
-0.837 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| O1 | | 1.3560 | 1.3610 | 2.3984 | 2.3421 | 2.0039 | 2.0604 | 3.3439 | 2.7319 | 3.3126 | 2.6102 |
C2 | 1.3560 | | 2.3289 | 1.3233 | 3.5042 | 1.0795 | 2.5290 | 2.0711 | 2.1127 | 4.3754 | 3.8632 | C3 | 1.3610 | 2.3289 | | 2.8167 | 1.3191 | 3.2266 | 1.0828 | 3.8818 | 2.6219 | 2.0751 | 2.0943 | C4 | 2.3984 | 1.3233 | 2.8167 | | 4.0028 | 2.1123 | 2.6117 | 1.0770 | 1.0780 | 4.6738 | 4.5534 | C5 | 2.3421 | 3.5042 | 1.3191 | 4.0028 | | 4.3329 | 2.1203 | 5.0789 | 3.6634 | 1.0770 | 1.0785 | H6 | 2.0039 | 1.0795 | 3.2266 | 2.1123 | 4.3329 | | 3.4948 | 2.4283 | 3.0821 | 5.2730 | 4.5298 | H7 | 2.0604 | 2.5290 | 1.0828 | 2.6117 | 2.1203 | 3.4948 | | 3.6063 | 2.2352 | 2.4524 | 3.0782 | H8 | 3.3439 | 2.0711 | 3.8818 | 1.0770 | 5.0789 | 2.4283 | 3.6063 | | 1.8487 | 5.7411 | 5.6007 | H9 | 2.7319 | 2.1127 | 2.6219 | 1.0780 | 3.6634 | 3.0821 | 2.2352 | 1.8487 | | 4.1371 | 4.3549 | H10 | 3.3126 | 4.3754 | 2.0751 | 4.6738 | 1.0770 | 5.2730 | 2.4524 | 5.7411 | 4.1371 | | 1.8566 | H11 | 2.6102 | 3.8632 | 2.0943 | 4.5534 | 1.0785 | 4.5298 | 3.0782 | 5.6007 | 4.3549 | 1.8566 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C4 |
127.046 |
|
O1 |
C2 |
H6 |
110.213 |
| O1 |
C3 |
C5 |
121.815 |
|
O1 |
C3 |
H7 |
114.460 |
| C2 |
O1 |
C3 |
117.997 |
|
C2 |
C4 |
H8 |
118.921 |
| C2 |
C4 |
H9 |
122.907 |
|
C3 |
C5 |
H10 |
119.659 |
| C3 |
C5 |
H11 |
121.406 |
|
C4 |
C2 |
H6 |
122.735 |
| C5 |
C3 |
H7 |
123.651 |
|
H8 |
C4 |
H9 |
118.158 |
| H10 |
C5 |
H11 |
118.930 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.159 |
|
|
|
| 2 |
C |
0.015 |
|
|
|
| 3 |
C |
0.046 |
|
|
|
| 4 |
C |
-0.379 |
|
|
|
| 5 |
C |
-0.354 |
|
|
|
| 6 |
H |
0.152 |
|
|
|
| 7 |
H |
0.120 |
|
|
|
| 8 |
H |
0.140 |
|
|
|
| 9 |
H |
0.137 |
|
|
|
| 10 |
H |
0.139 |
|
|
|
| 11 |
H |
0.143 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.337 |
0.593 |
0.241 |
0.724 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-27.282 |
-1.607 |
0.536 |
| y |
-1.607 |
-29.344 |
0.940 |
| z |
0.536 |
0.940 |
-32.461 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.620 |
-1.607 |
0.536 |
| y |
-1.607 |
0.528 |
0.940 |
| z |
0.536 |
0.940 |
-4.148 |
|
| Polar |
|---|
| 3z2-r2 | -8.296 |
|---|
| x2-y2 | 2.062 |
|---|
| xy | -1.607 |
|---|
| xz | 0.536 |
|---|
| yz | 0.940 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.079 |
0.438 |
0.477 |
| y |
0.438 |
7.048 |
0.137 |
| z |
0.477 |
0.137 |
5.038 |
<r2> (average value of r
2) Å
2
| <r2> |
129.016 |
| (<r2>)1/2 |
11.359 |
