return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for HCOOC2H5 (Ethyl formate)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS trans 1A'
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-267.976715
Energy at 298.15K-267.984014
HF Energy-266.929662
Nuclear repulsion energy178.667682
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3186 3025 13.73      
2 A' 3126 2968 8.40      
3 A' 3119 2961 56.26      
4 A' 3109 2951 11.11      
5 A' 1799 1708 273.35      
6 A' 1548 1469 7.82      
7 A' 1527 1450 2.65      
8 A' 1445 1372 16.66      
9 A' 1425 1352 2.73      
10 A' 1408 1336 0.79      
11 A' 1242 1179 410.32      
12 A' 1156 1097 6.52      
13 A' 1068 1014 16.57      
14 A' 879 835 10.63      
15 A' 799 759 3.12      
16 A' 387 367 5.29      
17 A' 232 221 6.69      
18 A" 3194 3033 21.71      
19 A" 3161 3001 5.31      
20 A" 1516 1439 6.64      
21 A" 1323 1256 1.54      
22 A" 1201 1141 3.86      
23 A" 1070 1016 0.03      
24 A" 825 783 0.65      
25 A" 357 339 18.57      
26 A" 257 244 3.23      
27 A" 76 72 0.55      

Unscaled Zero Point Vibrational Energy (zpe) 20217.5 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 19194.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
0.59678 0.09829 0.08711

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -2.161 -0.254 0.000
C2 -0.690 -0.555 0.000
O3 0.000 0.707 0.000
C4 1.333 0.611 0.000
O5 1.965 -0.412 0.000
H6 -2.726 -1.180 0.000
H7 -2.434 0.317 0.880
H8 -2.434 0.317 -0.880
H9 -0.389 -1.117 -0.877
H10 -0.389 -1.117 0.877
H11 1.764 1.611 0.000

Atom - Atom Distances (Å)
  C1 C2 O3 C4 O5 H6 H7 H8 H9 H10 H11
C11.50142.36523.59914.12951.08481.08381.08382.15732.15734.3453
C21.50141.43892.33502.65952.12952.13872.13871.08431.08433.2732
O32.36521.43891.33602.26173.31562.61712.61712.06152.06151.9817
C43.59912.33501.33601.20294.43623.87883.87882.59252.59251.0889
O54.12952.65952.26171.20294.75394.54494.54492.60972.60972.0329
H61.08482.12953.31564.43624.75391.76091.76092.49672.49675.2865
H71.08382.13872.61713.87884.54491.76091.76053.05342.49714.4796
H81.08382.13872.61713.87884.54491.76091.76052.49713.05344.4796
H92.15731.08432.06152.59252.60972.49673.05342.49711.75393.5843
H102.15731.08432.06152.59252.60972.49672.49713.05341.75393.5843
H114.34533.27321.98171.08892.03295.28654.47964.47963.58433.5843

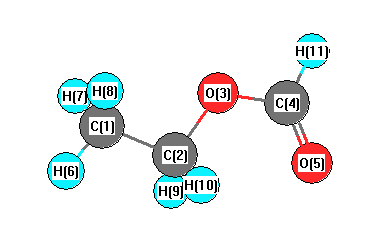
picture of Ethyl formate state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 O3 107.089 C1 C2 H9 112.088
C1 C2 H10 112.088 C2 C1 H6 109.806
C2 C1 H7 110.605 C2 C1 H8 110.605
C2 O3 C4 114.540 O3 C2 H9 108.764
O3 C2 H10 108.764 O3 C4 O5 125.868
O3 C4 H11 109.199 O5 C4 H11 124.933
H6 C1 H7 108.576 H6 C1 H8 108.576
H7 C1 H8 108.616 H9 C2 H10 107.960
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability