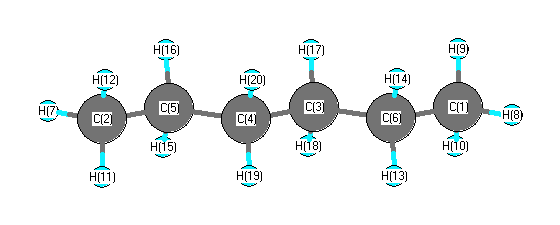
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3124 |
2990 |
0.00 |
|
|
|
| 2 |
Ag |
3049 |
2918 |
0.00 |
|
|
|
| 3 |
Ag |
3040 |
2910 |
0.00 |
|
|
|
| 4 |
Ag |
3023 |
2893 |
0.00 |
|
|
|
| 5 |
Ag |
1507 |
1443 |
0.00 |
|
|
|
| 6 |
Ag |
1491 |
1427 |
0.00 |
|
|
|
| 7 |
Ag |
1484 |
1420 |
0.00 |
|
|
|
| 8 |
Ag |
1411 |
1350 |
0.00 |
|
|
|
| 9 |
Ag |
1406 |
1346 |
0.00 |
|
|
|
| 10 |
Ag |
1335 |
1277 |
0.00 |
|
|
|
| 11 |
Ag |
1168 |
1118 |
0.00 |
|
|
|
| 12 |
Ag |
1095 |
1048 |
0.00 |
|
|
|
| 13 |
Ag |
1037 |
992 |
0.00 |
|
|
|
| 14 |
Ag |
909 |
870 |
0.00 |
|
|
|
| 15 |
Ag |
364 |
348 |
0.00 |
|
|
|
| 16 |
Ag |
285 |
273 |
0.00 |
|
|
|
| 17 |
Au |
3119 |
2985 |
117.95 |
|
|
|
| 18 |
Au |
3086 |
2954 |
70.19 |
|
|
|
| 19 |
Au |
3058 |
2927 |
0.33 |
|
|
|
| 20 |
Au |
1498 |
1434 |
14.68 |
|
|
|
| 21 |
Au |
1332 |
1275 |
0.92 |
|
|
|
| 22 |
Au |
1252 |
1198 |
0.01 |
|
|
|
| 23 |
Au |
1012 |
969 |
0.41 |
|
|
|
| 24 |
Au |
800 |
765 |
1.26 |
|
|
|
| 25 |
Au |
728 |
697 |
5.19 |
|
|
|
| 26 |
Au |
250 |
240 |
0.00 |
|
|
|
| 27 |
Au |
101 |
97 |
0.00 |
|
|
|
| 28 |
Au |
70 |
67 |
0.02 |
|
|
|
| 29 |
Bg |
3118 |
2985 |
0.00 |
|
|
|
| 30 |
Bg |
3075 |
2943 |
0.00 |
|
|
|
| 31 |
Bg |
3045 |
2914 |
0.00 |
|
|
|
| 32 |
Bg |
1498 |
1434 |
0.00 |
|
|
|
| 33 |
Bg |
1330 |
1273 |
0.00 |
|
|
|
| 34 |
Bg |
1305 |
1249 |
0.00 |
|
|
|
| 35 |
Bg |
1205 |
1154 |
0.00 |
|
|
|
| 36 |
Bg |
901 |
863 |
0.00 |
|
|
|
| 37 |
Bg |
741 |
709 |
0.00 |
|
|
|
| 38 |
Bg |
239 |
229 |
0.00 |
|
|
|
| 39 |
Bg |
145 |
139 |
0.00 |
|
|
|
| 40 |
Bu |
3124 |
2990 |
77.61 |
|
|
|
| 41 |
Bu |
3049 |
2918 |
93.84 |
|
|
|
| 42 |
Bu |
3043 |
2913 |
90.28 |
|
|
|
| 43 |
Bu |
3028 |
2898 |
10.95 |
|
|
|
| 44 |
Bu |
1513 |
1448 |
11.21 |
|
|
|
| 45 |
Bu |
1499 |
1435 |
1.71 |
|
|
|
| 46 |
Bu |
1484 |
1420 |
0.37 |
|
|
|
| 47 |
Bu |
1409 |
1348 |
6.58 |
|
|
|
| 48 |
Bu |
1387 |
1327 |
1.01 |
|
|
|
| 49 |
Bu |
1267 |
1212 |
1.77 |
|
|
|
| 50 |
Bu |
1089 |
1042 |
3.48 |
|
|
|
| 51 |
Bu |
1067 |
1021 |
1.11 |
|
|
|
| 52 |
Bu |
904 |
865 |
3.65 |
|
|
|
| 53 |
Bu |
453 |
433 |
0.05 |
|
|
|
| 54 |
Bu |
126 |
121 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 41538.7 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 39756.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.365 |
|
|
|
| 2 |
C |
-0.365 |
|
|
|
| 3 |
C |
-0.204 |
|
|
|
| 4 |
C |
-0.204 |
|
|
|
| 5 |
C |
-0.206 |
|
|
|
| 6 |
C |
-0.206 |
|
|
|
| 7 |
H |
0.118 |
|
|
|
| 8 |
H |
0.118 |
|
|
|
| 9 |
H |
0.111 |
|
|
|
| 10 |
H |
0.111 |
|
|
|
| 11 |
H |
0.111 |
|
|
|
| 12 |
H |
0.111 |
|
|
|
| 13 |
H |
0.110 |
|
|
|
| 14 |
H |
0.110 |
|
|
|
| 15 |
H |
0.110 |
|
|
|
| 16 |
H |
0.110 |
|
|
|
| 17 |
H |
0.108 |
|
|
|
| 18 |
H |
0.108 |
|
|
|
| 19 |
H |
0.108 |
|
|
|
| 20 |
H |
0.108 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.875 |
-0.056 |
0.000 |
| y |
-0.056 |
-43.098 |
0.000 |
| z |
0.000 |
0.000 |
-40.857 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.103 |
-0.056 |
0.000 |
| y |
-0.056 |
-1.733 |
0.000 |
| z |
0.000 |
0.000 |
1.630 |
|
| Polar |
|---|
| 3z2-r2 | 3.260 |
|---|
| x2-y2 | 1.223 |
|---|
| xy | -0.056 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.923 |
1.347 |
0.000 |
| y |
1.347 |
12.734 |
0.000 |
| z |
0.000 |
0.000 |
9.620 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |