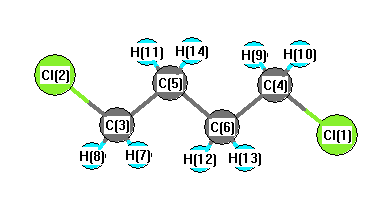
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3104 |
2970 |
0.00 |
|
|
|
| 2 |
Ag |
3061 |
2930 |
0.00 |
|
|
|
| 3 |
Ag |
1492 |
1428 |
0.00 |
|
|
|
| 4 |
Ag |
1490 |
1426 |
0.00 |
|
|
|
| 5 |
Ag |
1392 |
1332 |
0.00 |
|
|
|
| 6 |
Ag |
1290 |
1235 |
0.00 |
|
|
|
| 7 |
Ag |
1101 |
1054 |
0.00 |
|
|
|
| 8 |
Ag |
1052 |
1006 |
0.00 |
|
|
|
| 9 |
Ag |
776 |
742 |
0.00 |
|
|
|
| 10 |
Ag |
324 |
310 |
0.00 |
|
|
|
| 11 |
Ag |
221 |
211 |
0.00 |
|
|
|
| 12 |
Au |
3165 |
3029 |
23.53 |
|
|
|
| 13 |
Au |
3117 |
2983 |
16.02 |
|
|
|
| 14 |
Au |
1316 |
1259 |
1.09 |
|
|
|
| 15 |
Au |
1130 |
1082 |
1.26 |
|
|
|
| 16 |
Au |
897 |
859 |
0.52 |
|
|
|
| 17 |
Au |
745 |
713 |
4.45 |
|
|
|
| 18 |
Au |
113 |
109 |
1.33 |
|
|
|
| 19 |
Au |
62 |
60 |
4.73 |
|
|
|
| 20 |
Bg |
3163 |
3027 |
0.00 |
|
|
|
| 21 |
Bg |
3097 |
2964 |
0.00 |
|
|
|
| 22 |
Bg |
1324 |
1267 |
0.00 |
|
|
|
| 23 |
Bg |
1261 |
1207 |
0.00 |
|
|
|
| 24 |
Bg |
1074 |
1028 |
0.00 |
|
|
|
| 25 |
Bg |
778 |
745 |
0.00 |
|
|
|
| 26 |
Bg |
150 |
144 |
0.00 |
|
|
|
| 27 |
Bu |
3105 |
2972 |
42.99 |
|
|
|
| 28 |
Bu |
3070 |
2938 |
17.41 |
|
|
|
| 29 |
Bu |
1508 |
1444 |
9.22 |
|
|
|
| 30 |
Bu |
1488 |
1424 |
0.98 |
|
|
|
| 31 |
Bu |
1345 |
1287 |
55.30 |
|
|
|
| 32 |
Bu |
1231 |
1178 |
7.89 |
|
|
|
| 33 |
Bu |
1054 |
1009 |
17.40 |
|
|
|
| 34 |
Bu |
751 |
719 |
75.19 |
|
|
|
| 35 |
Bu |
414 |
396 |
12.80 |
|
|
|
| 36 |
Bu |
100 |
95 |
4.59 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25378.4 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 24289.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
Cl |
-0.170 |
|
|
|
| 2 |
Cl |
-0.170 |
|
|
|
| 3 |
C |
-0.157 |
|
|
|
| 4 |
C |
-0.157 |
|
|
|
| 5 |
C |
-0.235 |
|
|
|
| 6 |
C |
-0.235 |
|
|
|
| 7 |
H |
0.145 |
|
|
|
| 8 |
H |
0.145 |
|
|
|
| 9 |
H |
0.145 |
|
|
|
| 10 |
H |
0.145 |
|
|
|
| 11 |
H |
0.136 |
|
|
|
| 12 |
H |
0.136 |
|
|
|
| 13 |
H |
0.136 |
|
|
|
| 14 |
H |
0.136 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-50.212 |
-4.168 |
0.000 |
| y |
-4.168 |
-67.818 |
0.000 |
| z |
0.000 |
0.000 |
-50.155 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
8.775 |
-4.168 |
0.000 |
| y |
-4.168 |
-17.635 |
0.000 |
| z |
0.000 |
0.000 |
8.860 |
|
| Polar |
|---|
| 3z2-r2 | 17.720 |
|---|
| x2-y2 | 17.606 |
|---|
| xy | -4.168 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.340 |
1.239 |
0.000 |
| y |
1.239 |
14.764 |
0.000 |
| z |
0.000 |
0.000 |
8.154 |
<r2> (average value of r
2) Å
2
| <r2> |
511.787 |
| (<r2>)1/2 |
22.623 |