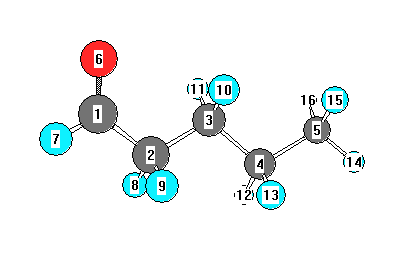
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3119 |
2985 |
34.73 |
|
|
|
| 2 |
A' |
3055 |
2924 |
33.04 |
|
|
|
| 3 |
A' |
3043 |
2912 |
24.15 |
|
|
|
| 4 |
A' |
3028 |
2898 |
25.03 |
|
|
|
| 5 |
A' |
3017 |
2888 |
21.04 |
|
|
|
| 6 |
A' |
2880 |
2756 |
155.87 |
|
|
|
| 7 |
A' |
1834 |
1755 |
160.01 |
|
|
|
| 8 |
A' |
1504 |
1439 |
8.66 |
|
|
|
| 9 |
A' |
1489 |
1425 |
0.72 |
|
|
|
| 10 |
A' |
1480 |
1417 |
0.99 |
|
|
|
| 11 |
A' |
1441 |
1380 |
14.82 |
|
|
|
| 12 |
A' |
1414 |
1353 |
11.98 |
|
|
|
| 13 |
A' |
1406 |
1345 |
2.87 |
|
|
|
| 14 |
A' |
1398 |
1338 |
6.90 |
|
|
|
| 15 |
A' |
1361 |
1302 |
15.34 |
|
|
|
| 16 |
A' |
1269 |
1215 |
4.36 |
|
|
|
| 17 |
A' |
1139 |
1090 |
10.03 |
|
|
|
| 18 |
A' |
1083 |
1037 |
0.37 |
|
|
|
| 19 |
A' |
1048 |
1003 |
1.64 |
|
|
|
| 20 |
A' |
923 |
884 |
1.94 |
|
|
|
| 21 |
A' |
897 |
858 |
13.90 |
|
|
|
| 22 |
A' |
692 |
662 |
13.86 |
|
|
|
| 23 |
A' |
387 |
370 |
1.57 |
|
|
|
| 24 |
A' |
280 |
268 |
4.30 |
|
|
|
| 25 |
A' |
126 |
121 |
5.19 |
|
|
|
| 26 |
A" |
3113 |
2979 |
56.98 |
|
|
|
| 27 |
A" |
3094 |
2961 |
20.46 |
|
|
|
| 28 |
A" |
3055 |
2924 |
4.98 |
|
|
|
| 29 |
A" |
3041 |
2910 |
8.31 |
|
|
|
| 30 |
A" |
1491 |
1427 |
7.50 |
|
|
|
| 31 |
A" |
1321 |
1264 |
0.24 |
|
|
|
| 32 |
A" |
1300 |
1244 |
0.09 |
|
|
|
| 33 |
A" |
1229 |
1177 |
0.01 |
|
|
|
| 34 |
A" |
1154 |
1105 |
0.22 |
|
|
|
| 35 |
A" |
972 |
930 |
0.58 |
|
|
|
| 36 |
A" |
848 |
812 |
0.13 |
|
|
|
| 37 |
A" |
737 |
706 |
2.25 |
|
|
|
| 38 |
A" |
661 |
632 |
3.50 |
|
|
|
| 39 |
A" |
229 |
219 |
0.01 |
|
|
|
| 40 |
A" |
187 |
179 |
1.32 |
|
|
|
| 41 |
A" |
108 |
103 |
1.24 |
|
|
|
| 42 |
A" |
62 |
59 |
1.83 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30954.9 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 29626.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.095 |
|
|
|
| 2 |
C |
-0.212 |
|
|
|
| 3 |
C |
-0.208 |
|
|
|
| 4 |
C |
-0.203 |
|
|
|
| 5 |
C |
-0.360 |
|
|
|
| 6 |
O |
-0.253 |
|
|
|
| 7 |
H |
0.078 |
|
|
|
| 8 |
H |
0.125 |
|
|
|
| 9 |
H |
0.125 |
|
|
|
| 10 |
H |
0.127 |
|
|
|
| 11 |
H |
0.127 |
|
|
|
| 12 |
H |
0.108 |
|
|
|
| 13 |
H |
0.108 |
|
|
|
| 14 |
H |
0.118 |
|
|
|
| 15 |
H |
0.113 |
|
|
|
| 16 |
H |
0.113 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.631 |
-0.065 |
0.000 |
2.632 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-44.850 |
3.980 |
0.000 |
| y |
3.980 |
-36.993 |
0.000 |
| z |
0.000 |
0.000 |
-37.202 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-7.753 |
3.980 |
0.000 |
| y |
3.980 |
4.033 |
0.000 |
| z |
0.000 |
0.000 |
3.719 |
|
| Polar |
|---|
| 3z2-r2 | 7.439 |
|---|
| x2-y2 | -7.857 |
|---|
| xy | 3.980 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.571 |
-0.527 |
0.000 |
| y |
-0.527 |
10.920 |
0.000 |
| z |
0.000 |
0.000 |
7.552 |
<r2> (average value of r
2) Å
2
| <r2> |
248.973 |
| (<r2>)1/2 |
15.779 |