Jump to
S1C2
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -235.811412 |
| Energy at 298.15K | -235.825862 |
| Nuclear repulsion energy | 257.639389 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1g |
3087 |
2954 |
0.00 |
|
|
|
| 2 |
A1g |
3027 |
2897 |
0.00 |
|
|
|
| 3 |
A1g |
1502 |
1438 |
0.00 |
|
|
|
| 4 |
A1g |
1180 |
1129 |
0.00 |
|
|
|
| 5 |
A1g |
824 |
789 |
0.00 |
|
|
|
| 6 |
A1g |
365 |
350 |
0.00 |
|
|
|
| 7 |
A1u |
1374 |
1315 |
0.00 |
|
|
|
| 8 |
A1u |
1130 |
1082 |
0.00 |
|
|
|
| 9 |
A1u |
1117 |
1069 |
0.00 |
|
|
|
| 10 |
A2g |
1345 |
1287 |
0.00 |
|
|
|
| 11 |
A2g |
1066 |
1020 |
0.00 |
|
|
|
| 12 |
A2u |
3092 |
2960 |
127.75 |
|
|
|
| 13 |
A2u |
3037 |
2906 |
81.89 |
|
|
|
| 14 |
A2u |
1487 |
1423 |
17.25 |
|
|
|
| 15 |
A2u |
1045 |
1000 |
2.14 |
|
|
|
| 16 |
A2u |
492 |
471 |
0.44 |
|
|
|
| 17 |
Eg |
3086 |
2954 |
0.00 |
|
|
|
| 17 |
Eg |
3086 |
2954 |
0.00 |
|
|
|
| 18 |
Eg |
3035 |
2905 |
0.00 |
|
|
|
| 18 |
Eg |
3035 |
2905 |
0.00 |
|
|
|
| 19 |
Eg |
1477 |
1414 |
0.00 |
|
|
|
| 19 |
Eg |
1477 |
1414 |
0.00 |
|
|
|
| 20 |
Eg |
1379 |
1320 |
0.00 |
|
|
|
| 20 |
Eg |
1379 |
1320 |
0.00 |
|
|
|
| 21 |
Eg |
1296 |
1240 |
0.00 |
|
|
|
| 21 |
Eg |
1296 |
1240 |
0.00 |
|
|
|
| 22 |
Eg |
1051 |
1006 |
0.00 |
|
|
|
| 22 |
Eg |
1051 |
1006 |
0.00 |
|
|
|
| 23 |
Eg |
786 |
752 |
0.00 |
|
|
|
| 23 |
Eg |
786 |
752 |
0.00 |
|
|
|
| 24 |
Eg |
411 |
393 |
0.00 |
|
|
|
| 24 |
Eg |
411 |
393 |
0.00 |
|
|
|
| 25 |
Eu |
3084 |
2951 |
110.20 |
|
|
|
| 25 |
Eu |
3084 |
2951 |
110.20 |
|
|
|
| 26 |
Eu |
3028 |
2898 |
29.14 |
|
|
|
| 26 |
Eu |
3028 |
2898 |
29.14 |
|
|
|
| 27 |
Eu |
1483 |
1419 |
5.63 |
|
|
|
| 27 |
Eu |
1483 |
1419 |
5.63 |
|
|
|
| 28 |
Eu |
1385 |
1325 |
0.03 |
|
|
|
| 28 |
Eu |
1385 |
1325 |
0.03 |
|
|
|
| 29 |
Eu |
1282 |
1227 |
1.68 |
|
|
|
| 29 |
Eu |
1282 |
1227 |
1.68 |
|
|
|
| 30 |
Eu |
911 |
872 |
1.18 |
|
|
|
| 30 |
Eu |
911 |
872 |
1.18 |
|
|
|
| 31 |
Eu |
880 |
842 |
3.14 |
|
|
|
| 31 |
Eu |
880 |
842 |
3.14 |
|
|
|
| 32 |
Eu |
220 |
210 |
0.01 |
|
|
|
| 32 |
Eu |
220 |
210 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 37376.3 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 35772.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is D3d
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
-1.449 |
0.231 |
| C2 |
-1.255 |
0.724 |
0.231 |
| C3 |
1.255 |
0.724 |
0.231 |
| C4 |
0.000 |
1.449 |
-0.231 |
| C5 |
-1.255 |
-0.724 |
-0.231 |
| C6 |
1.255 |
-0.724 |
-0.231 |
| H7 |
0.000 |
-1.502 |
1.323 |
| H8 |
-1.301 |
0.751 |
1.323 |
| H9 |
1.301 |
0.751 |
1.323 |
| H10 |
0.000 |
1.502 |
-1.323 |
| H11 |
-1.301 |
-0.751 |
-1.323 |
| H12 |
1.301 |
-0.751 |
-1.323 |
| H13 |
0.000 |
-2.477 |
-0.130 |
| H14 |
-2.145 |
1.239 |
-0.130 |
| H15 |
2.145 |
1.239 |
-0.130 |
| H16 |
0.000 |
2.477 |
0.130 |
| H17 |
-2.145 |
-1.239 |
0.130 |
| H18 |
2.145 |
-1.239 |
0.130 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
| C1 | | 2.5094 | 2.5094 | 2.9343 | 1.5208 | 1.5208 | 1.0933 | 2.7789 | 2.7789 | 3.3351 | 2.1437 | 2.1437 | 1.0900 | 3.4575 | 3.4575 | 3.9272 | 2.1579 | 2.1579 |
C2 | 2.5094 | | 2.5094 | 1.5208 | 1.5208 | 2.9343 | 2.7789 | 1.0933 | 2.7789 | 2.1437 | 2.1437 | 3.3351 | 3.4575 | 1.0900 | 3.4575 | 2.1579 | 2.1579 | 3.9272 | C3 | 2.5094 | 2.5094 | | 1.5208 | 2.9343 | 1.5208 | 2.7789 | 2.7789 | 1.0933 | 2.1437 | 3.3351 | 2.1437 | 3.4575 | 3.4575 | 1.0900 | 2.1579 | 3.9272 | 2.1579 | C4 | 2.9343 | 1.5208 | 1.5208 | | 2.5094 | 2.5094 | 3.3351 | 2.1437 | 2.1437 | 1.0933 | 2.7789 | 2.7789 | 3.9272 | 2.1579 | 2.1579 | 1.0900 | 3.4575 | 3.4575 | C5 | 1.5208 | 1.5208 | 2.9343 | 2.5094 | | 2.5094 | 2.1437 | 2.1437 | 3.3351 | 2.7789 | 1.0933 | 2.7789 | 2.1579 | 2.1579 | 3.9272 | 3.4575 | 1.0900 | 3.4575 | C6 | 1.5208 | 2.9343 | 1.5208 | 2.5094 | 2.5094 | | 2.1437 | 3.3351 | 2.1437 | 2.7789 | 2.7789 | 1.0933 | 2.1579 | 3.9272 | 2.1579 | 3.4575 | 3.4575 | 1.0900 | H7 | 1.0933 | 2.7789 | 2.7789 | 3.3351 | 2.1437 | 2.1437 | | 2.6012 | 2.6012 | 4.0033 | 3.0430 | 3.0430 | 1.7503 | 3.7715 | 3.7715 | 4.1540 | 2.4688 | 2.4688 | H8 | 2.7789 | 1.0933 | 2.7789 | 2.1437 | 2.1437 | 3.3351 | 2.6012 | | 2.6012 | 3.0430 | 3.0430 | 4.0033 | 3.7715 | 1.7503 | 3.7715 | 2.4688 | 2.4688 | 4.1540 | H9 | 2.7789 | 2.7789 | 1.0933 | 2.1437 | 3.3351 | 2.1437 | 2.6012 | 2.6012 | | 3.0430 | 4.0033 | 3.0430 | 3.7715 | 3.7715 | 1.7503 | 2.4688 | 4.1540 | 2.4688 | H10 | 3.3351 | 2.1437 | 2.1437 | 1.0933 | 2.7789 | 2.7789 | 4.0033 | 3.0430 | 3.0430 | | 2.6012 | 2.6012 | 4.1540 | 2.4688 | 2.4688 | 1.7503 | 3.7715 | 3.7715 | H11 | 2.1437 | 2.1437 | 3.3351 | 2.7789 | 1.0933 | 2.7789 | 3.0430 | 3.0430 | 4.0033 | 2.6012 | | 2.6012 | 2.4688 | 2.4688 | 4.1540 | 3.7715 | 1.7503 | 3.7715 | H12 | 2.1437 | 3.3351 | 2.1437 | 2.7789 | 2.7789 | 1.0933 | 3.0430 | 4.0033 | 3.0430 | 2.6012 | 2.6012 | | 2.4688 | 4.1540 | 2.4688 | 3.7715 | 3.7715 | 1.7503 | H13 | 1.0900 | 3.4575 | 3.4575 | 3.9272 | 2.1579 | 2.1579 | 1.7503 | 3.7715 | 3.7715 | 4.1540 | 2.4688 | 2.4688 | | 4.2905 | 4.2905 | 4.9611 | 2.4908 | 2.4908 | H14 | 3.4575 | 1.0900 | 3.4575 | 2.1579 | 2.1579 | 3.9272 | 3.7715 | 1.7503 | 3.7715 | 2.4688 | 2.4688 | 4.1540 | 4.2905 | | 4.2905 | 2.4908 | 2.4908 | 4.9611 | H15 | 3.4575 | 3.4575 | 1.0900 | 2.1579 | 3.9272 | 2.1579 | 3.7715 | 3.7715 | 1.7503 | 2.4688 | 4.1540 | 2.4688 | 4.2905 | 4.2905 | | 2.4908 | 4.9611 | 2.4908 | H16 | 3.9272 | 2.1579 | 2.1579 | 1.0900 | 3.4575 | 3.4575 | 4.1540 | 2.4688 | 2.4688 | 1.7503 | 3.7715 | 3.7715 | 4.9611 | 2.4908 | 2.4908 | | 4.2905 | 4.2905 | H17 | 2.1579 | 2.1579 | 3.9272 | 3.4575 | 1.0900 | 3.4575 | 2.4688 | 2.4688 | 4.1540 | 3.7715 | 1.7503 | 3.7715 | 2.4908 | 2.4908 | 4.9611 | 4.2905 | | 4.2905 | H18 | 2.1579 | 3.9272 | 2.1579 | 3.4575 | 3.4575 | 1.0900 | 2.4688 | 4.1540 | 2.4688 | 3.7715 | 3.7715 | 1.7503 | 2.4908 | 4.9611 | 2.4908 | 4.2905 | 4.2905 | |
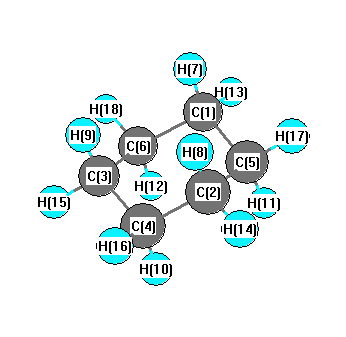 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C5 |
C2 |
111.174 |
|
C1 |
C5 |
H11 |
109.082 |
| C1 |
C5 |
H17 |
110.396 |
|
C1 |
C6 |
C3 |
111.174 |
| C1 |
C6 |
H12 |
109.082 |
|
C1 |
C6 |
H18 |
110.396 |
| C2 |
C4 |
C3 |
111.174 |
|
C2 |
C4 |
H10 |
109.082 |
| C2 |
C4 |
H16 |
110.396 |
|
C2 |
C5 |
H11 |
109.082 |
| C2 |
C5 |
H17 |
110.396 |
|
C3 |
C4 |
H10 |
109.082 |
| C3 |
C4 |
H16 |
110.396 |
|
C3 |
C6 |
H12 |
109.082 |
| C3 |
C6 |
H18 |
110.396 |
|
C4 |
C2 |
C5 |
111.174 |
| C4 |
C2 |
H8 |
109.082 |
|
C4 |
C2 |
H14 |
110.396 |
| C4 |
C3 |
C6 |
111.174 |
|
C4 |
C3 |
H9 |
109.082 |
| C4 |
C3 |
H15 |
110.396 |
|
C5 |
C1 |
C6 |
111.174 |
| C5 |
C1 |
H7 |
109.082 |
|
C5 |
C1 |
H13 |
110.396 |
| C5 |
C2 |
H8 |
109.082 |
|
C5 |
C2 |
H14 |
110.396 |
| C6 |
C1 |
H7 |
109.082 |
|
C6 |
C1 |
H13 |
110.396 |
| C6 |
C3 |
H9 |
109.082 |
|
C6 |
C3 |
H15 |
110.396 |
| H7 |
C1 |
H13 |
106.584 |
|
H8 |
C2 |
H14 |
106.584 |
| H9 |
C3 |
H15 |
106.584 |
|
H10 |
C4 |
H16 |
106.584 |
| H11 |
C5 |
H17 |
106.584 |
|
H12 |
C6 |
H18 |
106.584 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.227 |
|
|
|
| 2 |
C |
-0.227 |
|
|
|
| 3 |
C |
-0.227 |
|
|
|
| 4 |
C |
-0.227 |
|
|
|
| 5 |
C |
-0.227 |
|
|
|
| 6 |
C |
-0.227 |
|
|
|
| 7 |
H |
0.112 |
|
|
|
| 8 |
H |
0.112 |
|
|
|
| 9 |
H |
0.112 |
|
|
|
| 10 |
H |
0.112 |
|
|
|
| 11 |
H |
0.112 |
|
|
|
| 12 |
H |
0.112 |
|
|
|
| 13 |
H |
0.115 |
|
|
|
| 14 |
H |
0.115 |
|
|
|
| 15 |
H |
0.115 |
|
|
|
| 16 |
H |
0.115 |
|
|
|
| 17 |
H |
0.115 |
|
|
|
| 18 |
H |
0.115 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.188 |
0.000 |
0.000 |
| y |
0.000 |
-40.188 |
0.000 |
| z |
0.000 |
0.000 |
-39.620 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.284 |
0.000 |
0.000 |
| y |
0.000 |
-0.284 |
0.000 |
| z |
0.000 |
0.000 |
0.567 |
|
| Polar |
|---|
| 3z2-r2 | 1.135 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.604 |
0.000 |
0.000 |
| y |
0.000 |
10.605 |
0.000 |
| z |
0.000 |
0.000 |
9.248 |
<r2> (average value of r
2) Å
2
| <r2> |
163.429 |
| (<r2>)1/2 |
12.784 |
Jump to
S1C1
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -235.801944 |
| Energy at 298.15K | -235.816246 |
| Nuclear repulsion energy | 257.999068 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3099 |
2966 |
99.62 |
|
|
|
| 2 |
A |
3090 |
2958 |
0.00 |
|
|
|
| 3 |
A |
3071 |
2940 |
18.39 |
|
|
|
| 4 |
A |
3046 |
2915 |
0.00 |
|
|
|
| 5 |
A |
3041 |
2911 |
62.32 |
|
|
|
| 6 |
A |
3035 |
2905 |
0.00 |
|
|
|
| 7 |
A |
1511 |
1447 |
0.00 |
|
|
|
| 8 |
A |
1497 |
1433 |
12.55 |
|
|
|
| 9 |
A |
1481 |
1417 |
0.00 |
|
|
|
| 10 |
A |
1373 |
1315 |
0.00 |
|
|
|
| 11 |
A |
1349 |
1291 |
0.01 |
|
|
|
| 12 |
A |
1330 |
1273 |
0.49 |
|
|
|
| 13 |
A |
1289 |
1234 |
0.00 |
|
|
|
| 14 |
A |
1266 |
1212 |
0.00 |
|
|
|
| 15 |
A |
1253 |
1199 |
0.23 |
|
|
|
| 16 |
A |
1061 |
1015 |
0.57 |
|
|
|
| 17 |
A |
1036 |
991 |
0.00 |
|
|
|
| 18 |
A |
936 |
896 |
0.00 |
|
|
|
| 19 |
A |
898 |
860 |
0.03 |
|
|
|
| 20 |
A |
827 |
791 |
0.00 |
|
|
|
| 21 |
A |
778 |
745 |
1.85 |
|
|
|
| 22 |
A |
462 |
442 |
0.00 |
|
|
|
| 23 |
A |
422 |
404 |
0.30 |
|
|
|
| 24 |
A |
277 |
265 |
0.00 |
|
|
|
| 25 |
A |
127 |
121 |
0.04 |
|
|
|
| 26 |
B |
3094 |
2962 |
77.58 |
|
|
|
| 27 |
B |
3085 |
2953 |
61.63 |
|
|
|
| 28 |
B |
3074 |
2943 |
32.40 |
|
|
|
| 29 |
B |
3047 |
2917 |
67.07 |
|
|
|
| 30 |
B |
3036 |
2906 |
11.47 |
|
|
|
| 31 |
B |
3026 |
2896 |
23.16 |
|
|
|
| 32 |
B |
1493 |
1429 |
4.91 |
|
|
|
| 33 |
B |
1484 |
1420 |
4.04 |
|
|
|
| 34 |
B |
1478 |
1415 |
0.07 |
|
|
|
| 35 |
B |
1373 |
1314 |
0.00 |
|
|
|
| 36 |
B |
1369 |
1311 |
0.50 |
|
|
|
| 37 |
B |
1365 |
1306 |
0.47 |
|
|
|
| 38 |
B |
1306 |
1250 |
1.76 |
|
|
|
| 39 |
B |
1168 |
1118 |
1.79 |
|
|
|
| 40 |
B |
1138 |
1089 |
0.05 |
|
|
|
| 41 |
B |
1117 |
1069 |
0.00 |
|
|
|
| 42 |
B |
1104 |
1056 |
0.00 |
|
|
|
| 43 |
B |
1016 |
973 |
1.26 |
|
|
|
| 44 |
B |
885 |
847 |
2.28 |
|
|
|
| 45 |
B |
874 |
837 |
1.43 |
|
|
|
| 46 |
B |
756 |
724 |
1.89 |
|
|
|
| 47 |
B |
536 |
513 |
0.56 |
|
|
|
| 48 |
B |
239 |
229 |
0.06 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 37310.8 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 35710.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.203 |
-0.649 |
-0.396 |
| C2 |
0.000 |
1.513 |
-0.000 |
| C3 |
-1.203 |
-0.649 |
0.396 |
| C4 |
-1.203 |
0.649 |
-0.396 |
| C5 |
-0.000 |
-1.513 |
-0.000 |
| C6 |
1.203 |
0.649 |
0.396 |
| H7 |
1.156 |
-0.410 |
-1.461 |
| H8 |
0.270 |
2.166 |
-0.831 |
| H9 |
-2.131 |
-1.201 |
0.248 |
| H10 |
-1.156 |
0.410 |
-1.461 |
| H11 |
-0.270 |
-2.166 |
-0.831 |
| H12 |
2.131 |
1.201 |
0.248 |
| H13 |
2.131 |
-1.202 |
-0.248 |
| H14 |
-0.270 |
2.167 |
0.830 |
| H15 |
-1.156 |
-0.410 |
1.461 |
| H16 |
-2.131 |
1.202 |
-0.248 |
| H17 |
0.270 |
-2.167 |
0.830 |
| H18 |
1.156 |
0.410 |
1.461 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
| C1 | | 2.5055 | 2.5338 | 2.7348 | 1.5332 | 1.5205 | 1.0925 | 2.9977 | 3.4407 | 2.7973 | 2.1593 | 2.1678 | 1.0897 | 3.4062 | 3.0120 | 3.8165 | 2.1628 | 2.1382 |
C2 | 2.5055 | | 2.5056 | 1.5332 | 3.0252 | 1.5332 | 2.6773 | 1.0909 | 3.4596 | 2.1648 | 3.7811 | 2.1678 | 3.4596 | 1.0909 | 2.6773 | 2.1678 | 3.7813 | 2.1648 | C3 | 2.5338 | 2.5056 | | 1.5205 | 1.5332 | 2.7346 | 3.0122 | 3.4061 | 1.0897 | 2.1382 | 2.1628 | 3.8164 | 3.4407 | 2.9979 | 1.0925 | 2.1678 | 2.1593 | 2.7970 | C4 | 2.7348 | 1.5332 | 1.5205 | | 2.5055 | 2.5338 | 2.7973 | 2.1593 | 2.1678 | 1.0925 | 2.9977 | 3.4407 | 3.8165 | 2.1628 | 2.1382 | 1.0897 | 3.4062 | 3.0120 | C5 | 1.5332 | 3.0252 | 1.5332 | 2.5055 | | 2.5056 | 2.1648 | 3.7811 | 2.1678 | 2.6773 | 1.0909 | 3.4596 | 2.1678 | 3.7813 | 2.1648 | 3.4596 | 1.0909 | 2.6773 | C6 | 1.5205 | 1.5332 | 2.7346 | 2.5338 | 2.5056 | | 2.1382 | 2.1628 | 3.8164 | 3.0122 | 3.4061 | 1.0897 | 2.1678 | 2.1593 | 2.7970 | 3.4407 | 2.9979 | 1.0925 | H7 | 1.0925 | 2.6773 | 3.0122 | 2.7973 | 2.1648 | 2.1382 | | 2.7965 | 3.7886 | 2.4539 | 2.3486 | 2.5432 | 1.7458 | 3.7314 | 3.7262 | 3.8569 | 3.0199 | 3.0348 | H8 | 2.9977 | 1.0909 | 3.4061 | 2.1593 | 3.7811 | 2.1628 | 2.7965 | | 4.2744 | 2.3486 | 4.3660 | 2.3574 | 3.8916 | 1.7466 | 3.7314 | 2.6526 | 4.6402 | 3.0200 | H9 | 3.4407 | 3.4596 | 1.0897 | 2.1678 | 2.1678 | 3.8164 | 3.7886 | 4.2744 | | 2.5432 | 2.3574 | 4.8927 | 4.2908 | 3.8917 | 1.7458 | 2.4536 | 2.6524 | 3.8566 | H10 | 2.7973 | 2.1648 | 2.1382 | 1.0925 | 2.6773 | 3.0122 | 2.4539 | 2.3486 | 2.5432 | | 2.7965 | 3.7886 | 3.8569 | 3.0199 | 3.0348 | 1.7458 | 3.7314 | 3.7262 | H11 | 2.1593 | 3.7811 | 2.1628 | 2.9977 | 1.0909 | 3.4061 | 2.3486 | 4.3660 | 2.3574 | 2.7965 | | 4.2744 | 2.6526 | 4.6402 | 3.0200 | 3.8916 | 1.7466 | 3.7314 | H12 | 2.1678 | 2.1678 | 3.8164 | 3.4407 | 3.4596 | 1.0897 | 2.5432 | 2.3574 | 4.8927 | 3.7886 | 4.2744 | | 2.4536 | 2.6524 | 3.8566 | 4.2908 | 3.8917 | 1.7458 | H13 | 1.0897 | 3.4596 | 3.4407 | 3.8165 | 2.1678 | 2.1678 | 1.7458 | 3.8916 | 4.2908 | 3.8569 | 2.6526 | 2.4536 | | 4.2744 | 3.7883 | 4.8928 | 2.3573 | 2.5431 | H14 | 3.4062 | 1.0909 | 2.9979 | 2.1628 | 3.7813 | 2.1593 | 3.7314 | 1.7466 | 3.8917 | 3.0199 | 4.6402 | 2.6524 | 4.2744 | | 2.7967 | 2.3573 | 4.3665 | 2.3487 | H15 | 3.0120 | 2.6773 | 1.0925 | 2.1382 | 2.1648 | 2.7970 | 3.7262 | 3.7314 | 1.7458 | 3.0348 | 3.0200 | 3.8566 | 3.7883 | 2.7967 | | 2.5431 | 2.3487 | 2.4533 | H16 | 3.8165 | 2.1678 | 2.1678 | 1.0897 | 3.4596 | 3.4407 | 3.8569 | 2.6526 | 2.4536 | 1.7458 | 3.8916 | 4.2908 | 4.8928 | 2.3573 | 2.5431 | | 4.2744 | 3.7883 | H17 | 2.1628 | 3.7813 | 2.1593 | 3.4062 | 1.0909 | 2.9979 | 3.0199 | 4.6402 | 2.6524 | 3.7314 | 1.7466 | 3.8917 | 2.3573 | 4.3665 | 2.3487 | 4.2744 | | 2.7967 | H18 | 2.1382 | 2.1648 | 2.7970 | 3.0120 | 2.6773 | 1.0925 | 3.0348 | 3.0200 | 3.8566 | 3.7262 | 3.7314 | 1.7458 | 2.5431 | 2.3487 | 2.4533 | 3.7883 | 2.7967 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C5 |
C2 |
55.721 |
|
C1 |
C5 |
H11 |
109.590 |
| C1 |
C5 |
H17 |
109.863 |
|
C1 |
C6 |
C3 |
66.096 |
| C1 |
C6 |
H12 |
111.230 |
|
C1 |
C6 |
H18 |
108.719 |
| C2 |
C4 |
C3 |
110.265 |
|
C2 |
C4 |
H10 |
109.933 |
| C2 |
C4 |
H16 |
110.337 |
|
C2 |
C5 |
H11 |
126.811 |
| C2 |
C5 |
H17 |
126.828 |
|
C3 |
C4 |
H10 |
108.719 |
| C3 |
C4 |
H16 |
111.228 |
|
C3 |
C6 |
H12 |
171.830 |
| C3 |
C6 |
H18 |
81.837 |
|
C4 |
C2 |
C5 |
55.721 |
| C4 |
C2 |
H8 |
109.590 |
|
C4 |
C2 |
H14 |
109.863 |
| C4 |
C3 |
C6 |
66.096 |
|
C4 |
C3 |
H9 |
111.230 |
| C4 |
C3 |
H15 |
108.719 |
|
C5 |
C1 |
C6 |
110.265 |
| C5 |
C1 |
H7 |
109.933 |
|
C5 |
C1 |
H13 |
110.337 |
| C5 |
C2 |
H8 |
126.811 |
|
C5 |
C2 |
H14 |
126.828 |
| C6 |
C1 |
H7 |
108.719 |
|
C6 |
C1 |
H13 |
111.228 |
| C6 |
C3 |
H9 |
171.830 |
|
C6 |
C3 |
H15 |
81.837 |
| H7 |
C1 |
H13 |
106.265 |
|
H8 |
C2 |
H14 |
106.361 |
| H9 |
C3 |
H15 |
106.266 |
|
H10 |
C4 |
H16 |
106.265 |
| H11 |
C5 |
H17 |
106.361 |
|
H12 |
C6 |
H18 |
106.266 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.217 |
|
|
|
| 2 |
C |
-0.234 |
|
|
|
| 3 |
C |
-0.217 |
|
|
|
| 4 |
C |
-0.217 |
|
|
|
| 5 |
C |
-0.234 |
|
|
|
| 6 |
C |
-0.217 |
|
|
|
| 7 |
H |
0.111 |
|
|
|
| 8 |
H |
0.113 |
|
|
|
| 9 |
H |
0.110 |
|
|
|
| 10 |
H |
0.111 |
|
|
|
| 11 |
H |
0.113 |
|
|
|
| 12 |
H |
0.110 |
|
|
|
| 13 |
H |
0.110 |
|
|
|
| 14 |
H |
0.113 |
|
|
|
| 15 |
H |
0.111 |
|
|
|
| 16 |
H |
0.110 |
|
|
|
| 17 |
H |
0.113 |
|
|
|
| 18 |
H |
0.111 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.039 |
0.000 |
0.000 |
| y |
0.000 |
-40.358 |
0.000 |
| z |
0.000 |
0.000 |
-39.599 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.060 |
0.000 |
0.000 |
| y |
0.000 |
-0.539 |
0.000 |
| z |
0.000 |
0.000 |
0.599 |
|
| Polar |
|---|
| 3z2-r2 | 1.199 |
|---|
| x2-y2 | 0.319 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.451 |
0.000 |
0.000 |
| y |
0.000 |
10.607 |
0.000 |
| z |
0.000 |
0.000 |
9.219 |
<r2> (average value of r
2) Å
2
| <r2> |
161.636 |
| (<r2>)1/2 |
12.714 |
