return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H5N (Pyridine)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-247.852233
Energy at 298.15K-247.858434
HF Energy-246.777379
Nuclear repulsion energy207.085278
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3261 3096 3.60      
2 A1 3230 3067 3.71      
3 A1 3207 3045 3.40      
4 A1 1647 1563 14.77      
5 A1 1522 1445 3.47      
6 A1 1258 1194 1.70      
7 A1 1106 1050 3.95      
8 A1 1057 1004 6.55      
9 A1 1019 968 5.87      
10 A1 608 577 4.53      
11 A2 1024 972 0.00      
12 A2 927 880 0.00      
13 A2 389 369 0.00      
14 B1 1044 991 0.00      
15 B1 975 925 0.02      
16 B1 782 742 8.39      
17 B1 746 709 62.05      
18 B1 423 401 4.13      
19 B2 3251 3087 16.06      
20 B2 3205 3043 20.23      
21 B2 1635 1552 3.03      
22 B2 1491 1415 20.98      
23 B2 1416 1345 2.15      
24 B2 1392 1321 0.07      
25 B2 1193 1133 1.34      
26 B2 1091 1035 0.15      
27 B2 664 630 0.39      

Unscaled Zero Point Vibrational Energy (zpe) 19780.2 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 18779.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
0.20384 0.19578 0.09987

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 0.000 0.000 1.411
C2 0.000 0.000 -1.377
C3 0.000 1.138 0.718
C4 0.000 -1.138 0.718
C5 0.000 1.191 -0.669
C6 0.000 -1.191 -0.669
H7 0.000 0.000 -2.454
H8 0.000 2.050 1.296
H9 0.000 -2.050 1.296
H10 0.000 2.140 -1.175
H11 0.000 -2.140 -1.175

Atom - Atom Distances (Å)
  N1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11
N12.78891.33251.33252.39692.39693.86542.05362.05363.35663.3566
C22.78892.38492.38491.38581.38581.07663.36883.36882.14932.1493
C31.33252.38492.27621.38822.71073.37041.07943.24022.14183.7853
C41.33252.38492.27622.71071.38823.37043.24021.07943.78532.1418
C52.39691.38581.38822.71072.38162.14602.14413.78991.07543.3687
C62.39691.38582.71071.38822.38162.14603.78992.14413.36871.0754
H73.86541.07663.37043.37042.14602.14604.27354.27352.49292.4929
H82.05363.36881.07943.24022.14413.78994.27354.10062.47204.8640
H92.05363.36883.24021.07943.78992.14414.27354.10064.86402.4720
H103.35662.14932.14183.78531.07543.36872.49292.47204.86404.2794
H113.35662.14933.78532.14183.36871.07542.49294.86402.47204.2794

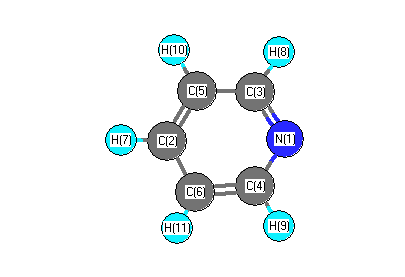
picture of Pyridine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C3 C5 123.514 N1 C3 H8 116.341
N1 C4 C6 123.514 N1 C4 H9 116.341
C2 C5 C3 118.582 C2 C5 H10 121.171
C2 C6 C4 118.582 C2 C6 H11 121.171
C3 N1 C4 117.325 C3 C5 H10 120.247
C4 C6 H11 120.247 C5 C2 C6 118.483
C5 C2 H7 120.759 C5 C3 H8 120.145
C6 C2 H7 120.759 C6 C4 H9 120.145
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability