Jump to
S1C2
Vibrational Frequencies calculated at mPW1PW91/cc-pVTZ
Geometric Data calculated at mPW1PW91/cc-pVTZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at mPW1PW91/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -251.935627 |
| Energy at 298.15K | -251.949931 |
| Nuclear repulsion energy | 259.885926 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3540 |
3396 |
0.38 |
|
|
|
| 2 |
A' |
3090 |
2964 |
67.57 |
|
|
|
| 3 |
A' |
3084 |
2958 |
38.92 |
|
|
|
| 4 |
A' |
3078 |
2953 |
47.86 |
|
|
|
| 5 |
A' |
3032 |
2909 |
54.64 |
|
|
|
| 6 |
A' |
3027 |
2904 |
42.59 |
|
|
|
| 7 |
A' |
3024 |
2901 |
1.37 |
|
|
|
| 8 |
A' |
1501 |
1439 |
3.10 |
|
|
|
| 9 |
A' |
1485 |
1424 |
11.80 |
|
|
|
| 10 |
A' |
1483 |
1423 |
3.16 |
|
|
|
| 11 |
A' |
1401 |
1344 |
2.63 |
|
|
|
| 12 |
A' |
1383 |
1326 |
2.58 |
|
|
|
| 13 |
A' |
1345 |
1290 |
0.24 |
|
|
|
| 14 |
A' |
1286 |
1234 |
6.91 |
|
|
|
| 15 |
A' |
1194 |
1145 |
2.30 |
|
|
|
| 16 |
A' |
1059 |
1016 |
1.61 |
|
|
|
| 17 |
A' |
1026 |
984 |
8.18 |
|
|
|
| 18 |
A' |
926 |
889 |
2.94 |
|
|
|
| 19 |
A' |
868 |
833 |
35.94 |
|
|
|
| 20 |
A' |
831 |
797 |
0.21 |
|
|
|
| 21 |
A' |
755 |
724 |
101.97 |
|
|
|
| 22 |
A' |
543 |
521 |
2.49 |
|
|
|
| 23 |
A' |
436 |
418 |
1.10 |
|
|
|
| 24 |
A' |
375 |
360 |
5.73 |
|
|
|
| 25 |
A' |
233 |
223 |
5.10 |
|
|
|
| 26 |
A" |
3088 |
2962 |
35.79 |
|
|
|
| 27 |
A" |
3077 |
2951 |
74.45 |
|
|
|
| 28 |
A" |
3029 |
2905 |
1.62 |
|
|
|
| 29 |
A" |
3021 |
2897 |
39.46 |
|
|
|
| 30 |
A" |
1500 |
1439 |
8.28 |
|
|
|
| 31 |
A" |
1481 |
1421 |
4.20 |
|
|
|
| 32 |
A" |
1471 |
1411 |
4.56 |
|
|
|
| 33 |
A" |
1385 |
1329 |
0.79 |
|
|
|
| 34 |
A" |
1371 |
1315 |
1.23 |
|
|
|
| 35 |
A" |
1353 |
1298 |
1.53 |
|
|
|
| 36 |
A" |
1305 |
1252 |
0.57 |
|
|
|
| 37 |
A" |
1227 |
1177 |
16.36 |
|
|
|
| 38 |
A" |
1147 |
1101 |
16.85 |
|
|
|
| 39 |
A" |
1125 |
1079 |
0.48 |
|
|
|
| 40 |
A" |
1077 |
1033 |
2.96 |
|
|
|
| 41 |
A" |
951 |
912 |
3.99 |
|
|
|
| 42 |
A" |
889 |
853 |
0.28 |
|
|
|
| 43 |
A" |
807 |
774 |
0.14 |
|
|
|
| 44 |
A" |
456 |
437 |
0.76 |
|
|
|
| 45 |
A" |
229 |
220 |
0.05 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 34996.4 cm
-1
Scaled (by 0.9592) Zero Point Vibrational Energy (zpe) 33568.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.637 |
1.317 |
0.000 |
| H2 |
0.575 |
2.407 |
0.000 |
| H3 |
1.703 |
1.067 |
0.000 |
| C4 |
-0.009 |
0.741 |
1.255 |
| C5 |
-0.009 |
0.741 |
-1.255 |
| C6 |
-0.009 |
-0.784 |
-1.207 |
| C7 |
-0.009 |
-0.784 |
1.207 |
| N8 |
-0.601 |
-1.340 |
0.000 |
| H9 |
-1.596 |
-1.161 |
0.000 |
| H10 |
0.510 |
1.090 |
2.151 |
| H11 |
0.510 |
1.090 |
-2.151 |
| H12 |
-1.042 |
1.097 |
1.331 |
| H13 |
-1.042 |
1.097 |
-1.331 |
| H14 |
1.024 |
-1.145 |
-1.264 |
| H15 |
1.024 |
-1.145 |
1.264 |
| H16 |
-0.531 |
-1.199 |
-2.071 |
| H17 |
-0.531 |
-1.199 |
2.071 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
H3 |
C4 |
C5 |
C6 |
C7 |
N8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
| C1 | | 1.0917 | 1.0952 | 1.5246 | 1.5246 | 2.5083 | 2.5083 | 2.9316 | 3.3359 | 2.1668 | 2.1668 | 2.1538 | 2.1538 | 2.7949 | 2.7949 | 3.4622 | 3.4622 |
H2 | 1.0917 | | 1.7518 | 2.1661 | 2.1661 | 3.4619 | 3.4619 | 3.9275 | 4.1766 | 2.5232 | 2.5232 | 2.4704 | 2.4704 | 3.7972 | 3.7972 | 4.3034 | 4.3034 | H3 | 1.0952 | 1.7518 | | 2.1478 | 2.1478 | 2.7960 | 2.7960 | 3.3323 | 3.9814 | 2.4601 | 2.4601 | 3.0509 | 3.0509 | 2.6372 | 2.6372 | 3.7970 | 3.7970 | C4 | 1.5246 | 2.1661 | 2.1478 | | 2.5099 | 2.8962 | 1.5262 | 2.5014 | 2.7769 | 1.0928 | 3.4630 | 1.0951 | 2.8070 | 3.3120 | 2.1505 | 3.8859 | 2.1687 | C5 | 1.5246 | 2.1661 | 2.1478 | 2.5099 | | 1.5262 | 2.8962 | 2.5014 | 2.7769 | 3.4630 | 1.0928 | 2.8070 | 1.0951 | 2.1505 | 3.3120 | 2.1687 | 3.8859 | C6 | 2.5083 | 3.4619 | 2.7960 | 2.8962 | 1.5262 | | 2.4140 | 1.4545 | 2.0293 | 3.8807 | 2.1620 | 3.3235 | 2.1496 | 1.0954 | 2.7022 | 1.0913 | 3.3453 | C7 | 2.5083 | 3.4619 | 2.7960 | 1.5262 | 2.8962 | 2.4140 | | 1.4545 | 2.0293 | 2.1620 | 3.8807 | 2.1496 | 3.3235 | 2.7022 | 1.0954 | 3.3453 | 1.0913 | N8 | 2.9316 | 3.9275 | 3.3323 | 2.5014 | 2.5014 | 1.4545 | 1.4545 | | 1.0120 | 3.4303 | 3.4303 | 2.8115 | 2.8115 | 2.0672 | 2.0672 | 2.0772 | 2.0772 | H9 | 3.3359 | 4.1766 | 3.9814 | 2.7769 | 2.7769 | 2.0293 | 2.0293 | 1.0120 | | 3.7589 | 3.7589 | 2.6783 | 2.6783 | 2.9090 | 2.9090 | 2.3297 | 2.3297 | H10 | 2.1668 | 2.5232 | 2.4601 | 1.0928 | 3.4630 | 3.8807 | 2.1620 | 3.4303 | 3.7589 | | 4.3023 | 1.7555 | 3.8121 | 4.1137 | 2.4592 | 4.9145 | 2.5159 | H11 | 2.1668 | 2.5232 | 2.4601 | 3.4630 | 1.0928 | 2.1620 | 3.8807 | 3.4303 | 3.7589 | 4.3023 | | 3.8121 | 1.7555 | 2.4592 | 4.1137 | 2.5159 | 4.9145 | H12 | 2.1538 | 2.4704 | 3.0509 | 1.0951 | 2.8070 | 3.3235 | 2.1496 | 2.8115 | 2.6783 | 1.7555 | 3.8121 | | 2.6614 | 4.0032 | 3.0492 | 4.1360 | 2.4661 | H13 | 2.1538 | 2.4704 | 3.0509 | 2.8070 | 1.0951 | 2.1496 | 3.3235 | 2.8115 | 2.6783 | 3.8121 | 1.7555 | 2.6614 | | 3.0492 | 4.0032 | 2.4661 | 4.1360 | H14 | 2.7949 | 3.7972 | 2.6372 | 3.3120 | 2.1505 | 1.0954 | 2.7022 | 2.0672 | 2.9090 | 4.1137 | 2.4592 | 4.0032 | 3.0492 | | 2.5278 | 1.7522 | 3.6799 | H15 | 2.7949 | 3.7972 | 2.6372 | 2.1505 | 3.3120 | 2.7022 | 1.0954 | 2.0672 | 2.9090 | 2.4592 | 4.1137 | 3.0492 | 4.0032 | 2.5278 | | 3.6799 | 1.7522 | H16 | 3.4622 | 4.3034 | 3.7970 | 3.8859 | 2.1687 | 1.0913 | 3.3453 | 2.0772 | 2.3297 | 4.9145 | 2.5159 | 4.1360 | 2.4661 | 1.7522 | 3.6799 | | 4.1425 | H17 | 3.4622 | 4.3034 | 3.7970 | 2.1687 | 3.8859 | 3.3453 | 1.0913 | 2.0772 | 2.3297 | 2.5159 | 4.9145 | 2.4661 | 4.1360 | 3.6799 | 1.7522 | 4.1425 | |
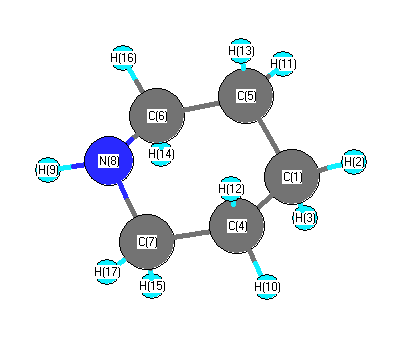 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C4 |
C7 |
110.607 |
|
C1 |
C4 |
H10 |
110.677 |
| C1 |
C4 |
H12 |
109.506 |
|
C1 |
C5 |
C6 |
110.607 |
| C1 |
C5 |
H11 |
110.677 |
|
C1 |
C5 |
H13 |
109.506 |
| H2 |
C1 |
H3 |
106.460 |
|
H2 |
C1 |
C4 |
110.687 |
| H2 |
C1 |
C5 |
110.687 |
|
H3 |
C1 |
C4 |
109.042 |
| H3 |
C1 |
C5 |
109.042 |
|
C4 |
C1 |
C5 |
110.802 |
| C4 |
C7 |
N8 |
114.085 |
|
C4 |
C7 |
H15 |
109.122 |
| C4 |
C7 |
H17 |
110.800 |
|
C5 |
C6 |
N8 |
114.085 |
| C5 |
C6 |
H14 |
109.122 |
|
C5 |
C6 |
H16 |
110.800 |
| C6 |
C5 |
H11 |
110.178 |
|
C6 |
C5 |
H13 |
109.069 |
| C6 |
N8 |
C7 |
112.161 |
|
C6 |
N8 |
H9 |
109.407 |
| C7 |
C4 |
H10 |
110.178 |
|
C7 |
C4 |
H12 |
109.069 |
| C7 |
N8 |
H9 |
109.407 |
|
N8 |
C6 |
H14 |
107.491 |
| N8 |
C6 |
H16 |
108.517 |
|
N8 |
C7 |
H15 |
107.491 |
| N8 |
C7 |
H17 |
108.517 |
|
H10 |
C4 |
H12 |
106.711 |
| H11 |
C5 |
H13 |
106.711 |
|
H14 |
C6 |
H16 |
106.505 |
| H15 |
C7 |
H17 |
106.505 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.151 |
|
|
|
| 2 |
H |
0.082 |
|
|
|
| 3 |
H |
0.079 |
|
|
|
| 4 |
C |
-0.161 |
|
|
|
| 5 |
C |
-0.161 |
|
|
|
| 6 |
C |
-0.078 |
|
|
|
| 7 |
C |
-0.078 |
|
|
|
| 8 |
N |
-0.206 |
|
|
|
| 9 |
H |
0.084 |
|
|
|
| 10 |
H |
0.075 |
|
|
|
| 11 |
H |
0.075 |
|
|
|
| 12 |
H |
0.071 |
|
|
|
| 13 |
H |
0.071 |
|
|
|
| 14 |
H |
0.071 |
|
|
|
| 15 |
H |
0.071 |
|
|
|
| 16 |
H |
0.079 |
|
|
|
| 17 |
H |
0.079 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.276 |
1.056 |
0.000 |
1.091 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.901 |
-0.217 |
0.000 |
| y |
-0.217 |
-43.187 |
0.000 |
| z |
0.000 |
0.000 |
-37.985 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.685 |
-0.217 |
0.000 |
| y |
-0.217 |
-5.744 |
0.000 |
| z |
0.000 |
0.000 |
2.059 |
|
| Polar |
|---|
| 3z2-r2 | 4.118 |
|---|
| x2-y2 | 6.286 |
|---|
| xy | -0.217 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.901 |
0.305 |
0.000 |
| y |
0.305 |
9.555 |
0.000 |
| z |
0.000 |
0.000 |
10.305 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |
