return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C8H18 (Octane)

using model chemistry: B1B95/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2H 1AG
Energy calculated at B1B95/cc-pVTZ
 hartrees
Energy at 0K-315.607418
Energy at 298.15K-315.626727
Nuclear repulsion energy376.434380
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 Ag 3114 2980 0.00      
2 Ag 3040 2909 0.00      
3 Ag 3030 2900 0.00      
4 Ag 3016 2887 0.00      
5 Ag 3014 2884 0.00      
6 Ag 1505 1440 0.00      
7 Ag 1491 1427 0.00      
8 Ag 1481 1418 0.00      
9 Ag 1478 1414 0.00      
10 Ag 1405 1344 0.00      
11 Ag 1400 1340 0.00      
12 Ag 1378 1319 0.00      
13 Ag 1291 1236 0.00      
14 Ag 1161 1111 0.00      
15 Ag 1093 1046 0.00      
16 Ag 1072 1026 0.00      
17 Ag 1017 973 0.00      
18 Ag 911 872 0.00      
19 Ag 455 436 0.00      
20 Ag 274 262 0.00      
21 Ag 185 177 0.00      
22 Au 3108 2975 118.51      
23 Au 3078 2946 130.40      
24 Au 3060 2929 0.71      
25 Au 3038 2907 0.18      
26 Au 1491 1427 14.44      
27 Au 1331 1274 0.68      
28 Au 1310 1254 0.35      
29 Au 1230 1177 0.00      
30 Au 1035 990 0.24      
31 Au 870 833 0.76      
32 Au 747 715 1.29      
33 Au 726 695 6.53      
34 Au 237 227 0.00      
35 Au 160 153 0.00      
36 Au 73 70 0.00      
37 Au 46 44 0.01      
38 Bg 3108 2975 0.00      
39 Bg 3070 2938 0.00      
40 Bg 3048 2917 0.00      
41 Bg 3033 2903 0.00      
42 Bg 1491 1427 0.00      
43 Bg 1330 1273 0.00      
44 Bg 1324 1267 0.00      
45 Bg 1274 1219 0.00      
46 Bg 1199 1148 0.00      
47 Bg 956 915 0.00      
48 Bg 795 761 0.00      
49 Bg 728 697 0.00      
50 Bg 240 230 0.00      
51 Bg 137 131 0.00      
52 Bg 107 103 0.00      
53 Bu 3114 2980 78.82      
54 Bu 3039 2909 92.98      
55 Bu 3032 2902 155.59      
56 Bu 3023 2894 12.48      
57 Bu 3013 2884 1.66      
58 Bu 1509 1444 13.25      
59 Bu 1498 1434 1.73      
60 Bu 1485 1421 0.85      
61 Bu 1477 1414 0.06      
62 Bu 1402 1342 5.52      
63 Bu 1399 1339 1.01      
64 Bu 1339 1282 1.41      
65 Bu 1241 1188 1.49      
66 Bu 1103 1056 4.63      
67 Bu 1084 1038 0.48      
68 Bu 1033 989 0.46      
69 Bu 894 855 3.68      
70 Bu 462 442 0.11      
71 Bu 323 309 0.02      
72 Bu 79 75 0.00      

Unscaled Zero Point Vibrational Energy (zpe) 53867.8 cm-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 51556.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B1B95/cc-pVTZ
ABC
0.36184 0.01650 0.01616

See section I.F.4 to change rotational constant units
Geometric Data calculated at B1B95/cc-pVTZ

Point Group is C2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.002 0.760 0.000
C2 -0.002 -0.760 0.000
C3 -1.391 1.368 0.000
C4 1.391 -1.368 0.000
C5 -1.391 2.888 0.000
C6 1.391 -2.888 0.000
C7 -2.788 3.483 0.000
C8 2.788 -3.483 0.000
H9 0.554 1.120 0.873
H10 0.554 1.120 -0.873
H11 -0.554 -1.120 0.873
H12 -0.554 -1.120 -0.873
H13 -1.943 1.008 -0.873
H14 -1.943 1.008 0.873
H15 1.943 -1.008 -0.873
H16 1.943 -1.008 0.873
H17 -0.839 3.246 0.873
H18 -0.839 3.246 -0.873
H19 0.839 -3.246 0.873
H20 0.839 -3.246 -0.873
H21 -2.761 4.572 0.000
H22 -3.349 3.165 -0.879
H23 -3.349 3.165 0.879
H24 2.761 -4.572 0.000
H25 3.349 -3.165 -0.879
H26 3.349 -3.165 0.879

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 C8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18 H19 H20 H21 H22 H23 H24 H25 H26
C11.51981.51962.54062.54323.90303.89845.07541.09381.09382.14592.14592.14612.14612.76612.76612.76582.76584.18444.18444.70834.21804.21806.00345.23295.2329
C21.51982.54061.51963.90302.54325.07543.89842.14592.14591.09381.09382.76612.76612.14612.14614.18444.18442.76582.76586.00345.23295.23294.70834.21804.2180
C31.51962.54063.90101.52005.08382.53486.40212.14592.14592.76602.76601.09391.09394.18514.18512.14332.14335.19805.19803.48502.80012.80017.24696.61726.6172
C42.54061.51963.90105.08381.52006.40212.53482.76602.76602.14592.14594.18514.18511.09391.09395.19805.19802.14332.14337.24696.61726.61723.48502.80012.8001
C52.54323.90301.52005.08386.41031.51867.61862.76932.76934.18614.18612.14522.14525.20065.20061.09281.09286.58456.58452.17152.16492.16498.53737.73817.7381
C63.90302.54325.08381.52006.41037.61861.51864.18614.18612.76932.76935.20065.20062.14522.14526.58456.58451.09281.09288.53737.73817.73812.17152.16492.1649
C73.89845.07542.53486.40211.51867.61868.92224.18484.18485.19005.19002.75732.75736.58046.58042.14852.14857.69357.69351.08931.09041.09049.78119.09039.0903
C85.07543.89846.40212.53487.61861.51868.92225.19005.19004.18484.18486.58046.58042.75732.75737.69357.69352.14852.14859.78119.09039.09031.08931.09041.0904
H91.09382.14592.14592.76602.76934.18614.18485.19001.74642.49863.04843.04872.49913.08282.54062.54173.08354.37524.71064.86524.74234.40686.16685.40795.1162
H101.09382.14592.14592.76602.76934.18614.18485.19001.74643.04842.49862.49913.04872.54063.08283.08352.54174.71064.37524.86524.40684.74236.16685.11625.4079
H112.14591.09382.76602.14594.18612.76935.19004.18482.49863.04841.74643.08282.54063.04872.49914.37524.71062.54173.08356.16685.40795.11624.86524.74234.4068
H122.14591.09382.76602.14594.18612.76935.19004.18483.04842.49861.74642.54063.08282.49913.04874.71064.37523.08352.54176.16685.11625.40794.86524.40684.7423
H132.14612.76611.09394.18512.14525.20062.75736.58043.04872.49913.08282.54061.74594.37684.71223.04572.49595.37375.08233.75982.57573.11507.34996.73936.9633
H142.14612.76611.09394.18512.14525.20062.75736.58042.49913.04872.54063.08281.74594.71224.37682.49593.04575.08235.37373.75983.11502.57577.34996.96336.7393
H152.76612.14614.18511.09395.20062.14526.58042.75733.08282.54063.04872.49914.37684.71221.74595.37375.08233.04572.49597.34996.73936.96333.75982.57573.1150
H162.76612.14614.18511.09395.20062.14526.58042.75732.54063.08282.49913.04874.71224.37681.74595.08235.37372.49593.04577.34996.96336.73933.75983.11502.5757
H172.76584.18442.14335.19801.09286.58452.14857.69352.54173.08354.37524.71063.04572.49595.37375.08231.74516.70536.92872.49323.06242.51218.65117.85577.6580
H182.76584.18442.14335.19801.09286.58452.14857.69353.08352.54174.71064.37522.49593.04575.08235.37371.74516.92876.70532.49322.51213.06248.65117.65807.8557
H194.18442.76585.19802.14336.58451.09287.69352.14854.37524.71062.54173.08355.37375.08233.04572.49596.70536.92871.74518.65117.85577.65802.49323.06242.5121
H204.18442.76585.19802.14336.58451.09287.69352.14854.71064.37523.08352.54175.08235.37372.49593.04576.92876.70531.74518.65117.65807.85572.49322.51213.0624
H214.70836.00343.48507.24692.17158.53731.08939.78114.86524.86526.16686.16683.75983.75987.34997.34992.49322.49328.65118.65111.75981.759810.68219.89839.8983
H224.21805.23292.80016.61722.16497.73811.09049.09034.74234.40685.40795.11622.57573.11506.73936.96333.06242.51217.85577.65801.75981.75789.89839.21689.3829
H234.21805.23292.80016.61722.16497.73811.09049.09034.40684.74235.11625.40793.11502.57576.96336.73932.51213.06247.65807.85571.75981.75789.89839.38299.2168
H246.00344.70837.24693.48508.53732.17159.78111.08936.16686.16684.86524.86527.34997.34993.75983.75988.65118.65112.49322.493210.68219.89839.89831.75981.7598
H255.23294.21806.61722.80017.73812.16499.09031.09045.40795.11624.74234.40686.73936.96332.57573.11507.85577.65803.06242.51219.89839.21689.38291.75981.7578
H265.23294.21806.61722.80017.73812.16499.09031.09045.11625.40794.40684.74236.96336.73933.11502.57577.65807.85572.51213.06249.89839.38299.21681.75981.7578

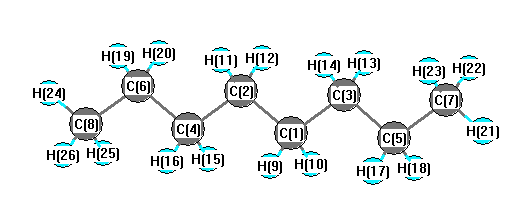
picture of Octane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 113.418 C1 C2 H11 109.298
C1 C2 H12 109.298 C1 C3 C5 113.581
C1 C3 H13 109.319 C1 C3 H14 109.319
C2 C1 C3 113.418 C2 C1 H9 109.298
C2 C1 H10 109.298 C2 C4 C6 113.581
C2 C4 H15 109.319 C2 C4 H16 109.319
C3 C1 H9 109.310 C3 C1 H10 109.310
C3 C5 C7 113.067 C3 C5 H17 109.138
C3 C5 H18 109.138 C4 C2 H11 109.310
C4 C2 H12 109.310 C4 C6 C8 113.067
C4 C6 H19 109.138 C4 C6 H20 109.138
C5 C3 H13 109.225 C5 C3 H14 109.225
C5 C7 H21 111.686 C5 C7 H22 111.091
C5 C7 H23 111.091 C6 C4 H15 109.225
C6 C4 H16 109.225 C6 C8 H24 111.686
C6 C8 H25 111.091 C6 C8 H26 111.091
C7 C5 H17 109.644 C7 C5 H18 109.644
C8 C6 H19 109.644 C8 C6 H20 109.644
H9 C1 H10 105.936 H11 C2 H12 105.936
H13 C3 H14 105.886 H15 C4 H16 105.886
H17 C5 H18 105.962 H19 C6 H20 105.962
H21 C7 H22 107.678 H21 C7 H23 107.678
H22 C7 H23 107.422 H24 C8 H25 107.678
H24 C8 H26 107.678 H25 C8 H26 107.422
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.213      
2 C -0.213      
3 C -0.204      
4 C -0.204      
5 C -0.204      
6 C -0.204      
7 C -0.364      
8 C -0.364      
9 H 0.108      
10 H 0.108      
11 H 0.108      
12 H 0.108      
13 H 0.107      
14 H 0.107      
15 H 0.107      
16 H 0.107      
17 H 0.108      
18 H 0.108      
19 H 0.108      
20 H 0.108      
21 H 0.117      
22 H 0.110      
23 H 0.110      
24 H 0.117      
25 H 0.110      
26 H 0.110      


Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section VII.A.3)
  x y z Total
  0.000 0.000 0.000 0.000
CHELPG        
AIM        
ESP        


Electric Quadrupole moment
Quadrupole components in D Å
Primitive
 xyz
x -56.655 0.613 0.000
y 0.613 -55.783 0.000
z 0.000 0.000 -53.937
Traceless
 xyz
x -1.795 0.613 0.000
y 0.613 -0.488 0.000
z 0.000 0.000 2.282
Polar
3z2-r24.565
x2-y2-0.872
xy0.613
xz0.000
yz0.000


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 15.213 -2.535 0.000
y -2.535 16.695 0.000
z 0.000 0.000 12.491


<r2> (average value of r2) Å2
<r2> 673.911
(<r2>)1/2 25.960