return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H8 (Cyclobutane, methylene-)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-194.920052
Energy at 298.15K-194.928639
HF Energy-194.009046
Nuclear repulsion energy169.037136
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3199 3037 4.57      
2 A' 3183 3022 31.23      
3 A' 3160 3000 7.77      
4 A' 3124 2966 32.28      
5 A' 3100 2943 11.10      
6 A' 1763 1674 18.14      
7 A' 1519 1442 0.07      
8 A' 1495 1419 3.49      
9 A' 1462 1388 1.35      
10 A' 1261 1197 0.60      
11 A' 1235 1172 0.16      
12 A' 1098 1042 0.71      
13 A' 996 946 0.50      
14 A' 933 885 42.04      
15 A' 916 869 0.30      
16 A' 760 721 2.29      
17 A' 665 631 1.92      
18 A' 389 369 5.75      
19 A' 134 127 0.32      
20 A" 3277 3111 6.34      
21 A" 3163 3003 20.75      
22 A" 3098 2942 28.29      
23 A" 1479 1404 4.54      
24 A" 1294 1229 0.01      
25 A" 1259 1195 0.27      
26 A" 1228 1166 0.61      
27 A" 1191 1131 1.66      
28 A" 1054 1000 0.22      
29 A" 943 896 1.49      
30 A" 930 883 0.72      
31 A" 822 780 0.07      
32 A" 662 628 0.08      
33 A" 356 338 0.09      

Unscaled Zero Point Vibrational Energy (zpe) 25572.8 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 24278.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
0.34726 0.15716 0.11927

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.252 -0.381 1.086
C2 0.252 -0.381 -1.086
C3 -0.121 0.601 0.000
C4 0.252 -1.483 0.000
C5 -0.669 1.806 0.000
H6 -0.448 -0.492 1.907
H7 -0.448 -0.492 -1.907
H8 1.243 -0.194 1.489
H9 1.243 -0.194 -1.489
H10 -0.904 2.313 0.921
H11 -0.904 2.313 -0.921
H12 1.094 -2.162 0.000
H13 -0.668 -2.055 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13
C12.17211.51111.54682.60951.08443.07591.08642.76592.93613.55292.24982.1971
C22.17211.51111.54682.60953.07591.08442.76591.08643.55292.93612.24982.1971
C31.51111.51112.11681.32332.22252.22252.17062.17062.09572.09573.01882.7120
C41.54681.54682.11683.41472.25992.25992.20472.20474.07324.07321.08261.0832
C52.60952.60951.32333.41472.99452.99453.14213.14211.07771.07774.34213.8610
H61.08443.07592.22252.25992.99453.81431.76703.80583.00834.00982.96722.4755
H73.07591.08442.22252.25992.99453.81433.80581.76704.00983.00832.96722.4755
H81.08642.76592.17062.20473.14211.76703.80582.97873.34924.08732.47243.0549
H92.76591.08642.17062.20473.14213.80581.76702.97874.08733.34922.47243.0549
H102.93613.55292.09574.07321.07773.00834.00983.34924.08731.84264.98694.4707
H113.55292.93612.09574.07321.07774.00983.00834.08733.34921.84264.98694.4707
H122.24982.24983.01881.08264.34212.96722.96722.47242.47244.98694.98691.7655
H132.19712.19712.71201.08323.86102.47552.47553.05493.05494.47074.47071.7655

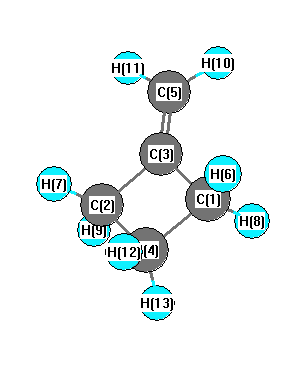
picture of Cyclobutane, methylene- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C3 C2 91.898 C1 C3 C5 133.938
C1 C4 C2 89.195 C1 C4 H12 116.551
C1 C4 H13 112.111 C2 C3 C5 133.938
C2 C4 H12 116.551 C2 C4 H13 112.111
C3 C1 C4 87.607 C3 C1 H6 116.852
C3 C1 H8 112.340 C3 C2 C4 87.607
C3 C2 H7 116.852 C3 C2 H9 112.340
C3 C5 H10 121.249 C3 C5 H11 121.249
C4 C1 H6 117.302 C4 C1 H8 112.529
C4 C2 H7 117.302 C4 C2 H9 112.529
H6 C1 H8 108.974 H7 C2 H9 108.974
H10 C5 H11 117.500 H12 C4 H13 109.204
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability