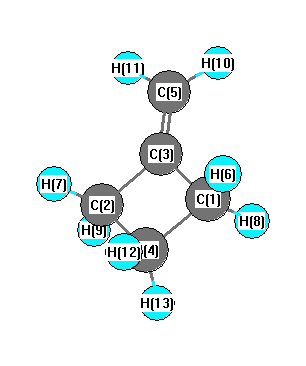
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3156 |
3021 |
6.47 |
|
|
|
| 2 |
A' |
3150 |
3015 |
40.38 |
|
|
|
| 3 |
A' |
3119 |
2986 |
12.40 |
|
|
|
| 4 |
A' |
3092 |
2960 |
41.46 |
|
|
|
| 5 |
A' |
3066 |
2935 |
11.21 |
|
|
|
| 6 |
A' |
1774 |
1698 |
30.11 |
|
|
|
| 7 |
A' |
1498 |
1434 |
0.06 |
|
|
|
| 8 |
A' |
1471 |
1408 |
2.53 |
|
|
|
| 9 |
A' |
1440 |
1378 |
1.24 |
|
|
|
| 10 |
A' |
1262 |
1207 |
0.01 |
|
|
|
| 11 |
A' |
1226 |
1173 |
0.29 |
|
|
|
| 12 |
A' |
1093 |
1046 |
0.45 |
|
|
|
| 13 |
A' |
993 |
950 |
0.49 |
|
|
|
| 14 |
A' |
916 |
877 |
42.77 |
|
|
|
| 15 |
A' |
902 |
863 |
0.60 |
|
|
|
| 16 |
A' |
753 |
721 |
2.05 |
|
|
|
| 17 |
A' |
662 |
634 |
2.19 |
|
|
|
| 18 |
A' |
387 |
371 |
6.41 |
|
|
|
| 19 |
A' |
115 |
110 |
0.10 |
|
|
|
| 20 |
A" |
3240 |
3101 |
11.24 |
|
|
|
| 21 |
A" |
3123 |
2989 |
21.22 |
|
|
|
| 22 |
A" |
3062 |
2931 |
39.23 |
|
|
|
| 23 |
A" |
1456 |
1394 |
5.21 |
|
|
|
| 24 |
A" |
1292 |
1236 |
0.08 |
|
|
|
| 25 |
A" |
1244 |
1191 |
0.20 |
|
|
|
| 26 |
A" |
1224 |
1171 |
0.82 |
|
|
|
| 27 |
A" |
1195 |
1143 |
1.80 |
|
|
|
| 28 |
A" |
1042 |
998 |
0.16 |
|
|
|
| 29 |
A" |
946 |
905 |
1.05 |
|
|
|
| 30 |
A" |
919 |
880 |
1.20 |
|
|
|
| 31 |
A" |
810 |
775 |
0.09 |
|
|
|
| 32 |
A" |
644 |
616 |
0.09 |
|
|
|
| 33 |
A" |
358 |
342 |
0.10 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25314.1 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 24228.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.223 |
|
|
|
| 2 |
C |
-0.223 |
|
|
|
| 3 |
C |
0.069 |
|
|
|
| 4 |
C |
-0.200 |
|
|
|
| 5 |
C |
-0.341 |
|
|
|
| 6 |
H |
0.108 |
|
|
|
| 7 |
H |
0.108 |
|
|
|
| 8 |
H |
0.116 |
|
|
|
| 9 |
H |
0.116 |
|
|
|
| 10 |
H |
0.125 |
|
|
|
| 11 |
H |
0.125 |
|
|
|
| 12 |
H |
0.107 |
|
|
|
| 13 |
H |
0.112 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.226 |
-0.533 |
0.000 |
0.579 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.824 |
-0.607 |
0.000 |
| y |
-0.607 |
-32.217 |
0.000 |
| z |
0.000 |
0.000 |
-30.677 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.377 |
-0.607 |
0.000 |
| y |
-0.607 |
-0.967 |
0.000 |
| z |
0.000 |
0.000 |
1.344 |
|
| Polar |
|---|
| 3z2-r2 | 2.687 |
|---|
| x2-y2 | 0.393 |
|---|
| xy | -0.607 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.182 |
-1.672 |
0.000 |
| y |
-1.672 |
9.996 |
0.000 |
| z |
0.000 |
0.000 |
8.186 |
<r2> (average value of r
2) Å
2
| <r2> |
114.400 |
| (<r2>)1/2 |
10.696 |