return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3OCH3 (Dimethyl ether)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-154.769826
Energy at 298.15K-154.776549
HF Energy-154.126462
Nuclear repulsion energy84.498095
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3187 3026 17.52 94.07 0.31 0.47
2 A1 3043 2889 55.85 264.90 0.02 0.05
3 A1 1549 1471 2.18 11.22 0.75 0.85
4 A1 1511 1435 0.09 2.85 0.74 0.85
5 A1 1284 1219 6.14 1.44 0.49 0.66
6 A1 978 928 33.97 7.60 0.39 0.56
7 A1 420 399 2.43 0.48 0.05 0.09
8 A2 3094 2938 0.00 14.19 0.75 0.86
9 A2 1515 1438 0.00 18.03 0.75 0.86
10 A2 1182 1122 0.00 2.71 0.75 0.86
11 A2 216 205 0.00 0.20 0.75 0.86
12 B1 3087 2931 111.65 93.84 0.75 0.86
13 B1 1524 1447 12.98 0.01 0.75 0.86
14 B1 1213 1152 7.82 0.91 0.75 0.86
15 B1 260 247 5.17 0.01 0.75 0.86
16 B2 3186 3025 26.58 63.71 0.75 0.86
17 B2 3036 2882 46.51 0.88 0.75 0.86
18 B2 1532 1455 11.65 2.11 0.75 0.86
19 B2 1476 1402 3.88 2.78 0.75 0.86
20 B2 1228 1166 112.26 0.51 0.75 0.86
21 B2 1143 1085 22.09 2.27 0.75 0.86

Unscaled Zero Point Vibrational Energy (zpe) 17832.0 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 16929.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
1.29725 0.34349 0.30232

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
O1 0.000 0.000 0.600
C2 0.000 1.155 -0.199
C3 0.000 -1.155 -0.199
H4 0.000 2.012 0.462
H5 0.000 -2.012 0.462
H6 0.885 1.197 -0.836
H7 -0.885 1.197 -0.836
H8 -0.885 -1.197 -0.836
H9 0.885 -1.197 -0.836

Atom - Atom Distances (Å)
  O1 C2 C3 H4 H5 H6 H7 H8 H9
O11.40411.40412.01622.01622.06862.06862.06862.0686
C21.40412.30911.08243.23431.09141.09142.59202.5920
C31.40412.30913.23431.08242.59202.59201.09141.0914
H42.01621.08243.23434.02301.77001.77003.57263.5726
H52.01623.23431.08244.02303.57263.57261.77001.7700
H62.06861.09142.59201.77003.57261.76972.97702.3939
H72.06861.09142.59201.77003.57261.76972.39392.9770
H82.06862.59201.09143.57261.77002.97702.39391.7697
H92.06862.59201.09143.57261.77002.39392.97701.7697

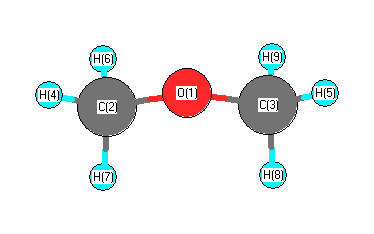
picture of Dimethyl ether state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
O1 C2 H4 107.659 O1 C2 H6 111.368
O1 C2 H7 111.368 O1 C3 H5 107.659
O1 C3 H8 111.368 O1 C3 H9 111.368
C2 O1 C3 110.621 H4 C2 H6 109.026
H4 C2 H7 109.026 H5 C3 H8 109.026
H5 C3 H9 109.026 H6 C2 H7 108.344
H8 C3 H9 108.344
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability