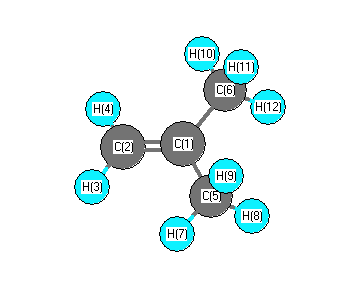
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3156 |
3021 |
8.68 |
|
|
|
| 2 |
A1 |
3143 |
3008 |
19.26 |
|
|
|
| 3 |
A1 |
3044 |
2913 |
35.71 |
|
|
|
| 4 |
A1 |
1744 |
1669 |
21.63 |
|
|
|
| 5 |
A1 |
1498 |
1434 |
14.65 |
|
|
|
| 6 |
A1 |
1442 |
1380 |
0.72 |
|
|
|
| 7 |
A1 |
1405 |
1345 |
0.51 |
|
|
|
| 8 |
A1 |
1079 |
1033 |
3.60 |
|
|
|
| 9 |
A1 |
831 |
795 |
0.21 |
|
|
|
| 10 |
A1 |
366 |
350 |
0.32 |
|
|
|
| 11 |
A2 |
3092 |
2960 |
0.00 |
|
|
|
| 12 |
A2 |
1468 |
1405 |
0.00 |
|
|
|
| 13 |
A2 |
1017 |
973 |
0.00 |
|
|
|
| 14 |
A2 |
706 |
675 |
0.00 |
|
|
|
| 15 |
A2 |
168 |
160 |
0.00 |
|
|
|
| 16 |
B1 |
3094 |
2961 |
39.67 |
|
|
|
| 17 |
B1 |
1486 |
1422 |
16.56 |
|
|
|
| 18 |
B1 |
1103 |
1056 |
0.09 |
|
|
|
| 19 |
B1 |
927 |
888 |
43.92 |
|
|
|
| 20 |
B1 |
438 |
419 |
7.59 |
|
|
|
| 21 |
B1 |
208 |
199 |
0.44 |
|
|
|
| 22 |
B2 |
3240 |
3101 |
18.25 |
|
|
|
| 23 |
B2 |
3142 |
3007 |
23.30 |
|
|
|
| 24 |
B2 |
3039 |
2909 |
18.83 |
|
|
|
| 25 |
B2 |
1480 |
1417 |
3.87 |
|
|
|
| 26 |
B2 |
1409 |
1348 |
12.27 |
|
|
|
| 27 |
B2 |
1306 |
1250 |
2.04 |
|
|
|
| 28 |
B2 |
989 |
946 |
0.34 |
|
|
|
| 29 |
B2 |
957 |
916 |
0.02 |
|
|
|
| 30 |
B2 |
427 |
409 |
1.69 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23700.8 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 22684.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.119 |
|
|
|
| 2 |
C |
-0.375 |
|
|
|
| 3 |
H |
0.121 |
|
|
|
| 4 |
H |
0.121 |
|
|
|
| 5 |
C |
-0.336 |
|
|
|
| 6 |
C |
-0.336 |
|
|
|
| 7 |
H |
0.114 |
|
|
|
| 8 |
H |
0.114 |
|
|
|
| 9 |
H |
0.114 |
|
|
|
| 10 |
H |
0.114 |
|
|
|
| 11 |
H |
0.114 |
|
|
|
| 12 |
H |
0.114 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.556 |
0.556 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-27.975 |
0.000 |
0.000 |
| y |
0.000 |
-25.623 |
0.000 |
| z |
0.000 |
0.000 |
-26.132 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.098 |
0.000 |
0.000 |
| y |
0.000 |
1.431 |
0.000 |
| z |
0.000 |
0.000 |
0.667 |
|
| Polar |
|---|
| 3z2-r2 | 1.334 |
|---|
| x2-y2 | -2.353 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.350 |
0.000 |
0.000 |
| y |
0.000 |
7.411 |
0.000 |
| z |
0.000 |
0.000 |
9.045 |
<r2> (average value of r
2) Å
2
| <r2> |
89.902 |
| (<r2>)1/2 |
9.482 |