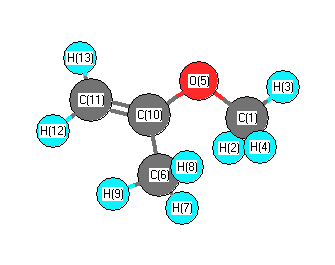
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3286 |
3145 |
6.54 |
|
|
|
| 2 |
A |
3191 |
3054 |
0.28 |
|
|
|
| 3 |
A |
3166 |
3030 |
11.55 |
|
|
|
| 4 |
A |
3154 |
3019 |
24.98 |
|
|
|
| 5 |
A |
3109 |
2976 |
14.32 |
|
|
|
| 6 |
A |
3092 |
2959 |
40.48 |
|
|
|
| 7 |
A |
3053 |
2922 |
21.62 |
|
|
|
| 8 |
A |
3028 |
2898 |
52.40 |
|
|
|
| 9 |
A |
1741 |
1666 |
138.98 |
|
|
|
| 10 |
A |
1519 |
1454 |
4.28 |
|
|
|
| 11 |
A |
1506 |
1441 |
13.40 |
|
|
|
| 12 |
A |
1490 |
1426 |
13.16 |
|
|
|
| 13 |
A |
1483 |
1420 |
6.84 |
|
|
|
| 14 |
A |
1479 |
1416 |
6.42 |
|
|
|
| 15 |
A |
1426 |
1365 |
8.85 |
|
|
|
| 16 |
A |
1405 |
1345 |
46.88 |
|
|
|
| 17 |
A |
1323 |
1266 |
178.10 |
|
|
|
| 18 |
A |
1213 |
1161 |
0.41 |
|
|
|
| 19 |
A |
1178 |
1127 |
9.06 |
|
|
|
| 20 |
A |
1154 |
1105 |
98.94 |
|
|
|
| 21 |
A |
1064 |
1019 |
1.43 |
|
|
|
| 22 |
A |
1020 |
976 |
4.41 |
|
|
|
| 23 |
A |
938 |
898 |
6.43 |
|
|
|
| 24 |
A |
860 |
823 |
56.23 |
|
|
|
| 25 |
A |
825 |
790 |
9.94 |
|
|
|
| 26 |
A |
743 |
711 |
2.75 |
|
|
|
| 27 |
A |
539 |
516 |
6.67 |
|
|
|
| 28 |
A |
488 |
467 |
3.05 |
|
|
|
| 29 |
A |
421 |
403 |
0.94 |
|
|
|
| 30 |
A |
345 |
330 |
0.60 |
|
|
|
| 31 |
A |
279 |
267 |
0.38 |
|
|
|
| 32 |
A |
219 |
210 |
2.08 |
|
|
|
| 33 |
A |
81 |
77 |
5.65 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24908.2 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 23839.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.179 |
|
|
|
| 2 |
H |
0.097 |
|
|
|
| 3 |
H |
0.128 |
|
|
|
| 4 |
H |
0.094 |
|
|
|
| 5 |
O |
-0.214 |
|
|
|
| 6 |
C |
-0.377 |
|
|
|
| 7 |
H |
0.121 |
|
|
|
| 8 |
H |
0.131 |
|
|
|
| 9 |
H |
0.125 |
|
|
|
| 10 |
C |
0.227 |
|
|
|
| 11 |
C |
-0.416 |
|
|
|
| 12 |
H |
0.130 |
|
|
|
| 13 |
H |
0.132 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.987 |
1.578 |
0.406 |
1.905 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-28.551 |
-0.262 |
0.291 |
| y |
-0.262 |
-31.494 |
-0.436 |
| z |
0.291 |
-0.436 |
-33.197 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.795 |
-0.262 |
0.291 |
| y |
-0.262 |
-0.620 |
-0.436 |
| z |
0.291 |
-0.436 |
-3.175 |
|
| Polar |
|---|
| 3z2-r2 | -6.350 |
|---|
| x2-y2 | 2.944 |
|---|
| xy | -0.262 |
|---|
| xz | 0.291 |
|---|
| yz | -0.436 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.907 |
0.978 |
-0.146 |
| y |
0.978 |
8.013 |
-0.115 |
| z |
-0.146 |
-0.115 |
5.754 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |