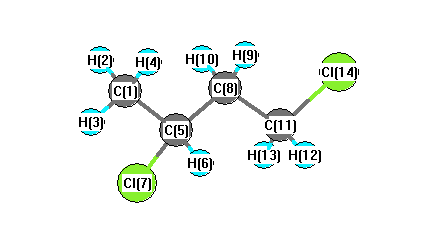
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3178 |
3042 |
4.24 |
|
|
|
| 2 |
A |
3159 |
3023 |
14.13 |
|
|
|
| 3 |
A |
3138 |
3004 |
19.87 |
|
|
|
| 4 |
A |
3120 |
2986 |
11.08 |
|
|
|
| 5 |
A |
3108 |
2975 |
9.06 |
|
|
|
| 6 |
A |
3099 |
2966 |
2.59 |
|
|
|
| 7 |
A |
3066 |
2935 |
18.70 |
|
|
|
| 8 |
A |
3061 |
2930 |
4.82 |
|
|
|
| 9 |
A |
1495 |
1431 |
3.06 |
|
|
|
| 10 |
A |
1494 |
1430 |
4.46 |
|
|
|
| 11 |
A |
1485 |
1422 |
9.58 |
|
|
|
| 12 |
A |
1473 |
1410 |
1.24 |
|
|
|
| 13 |
A |
1410 |
1349 |
7.03 |
|
|
|
| 14 |
A |
1393 |
1333 |
9.01 |
|
|
|
| 15 |
A |
1339 |
1281 |
6.64 |
|
|
|
| 16 |
A |
1320 |
1263 |
8.89 |
|
|
|
| 17 |
A |
1281 |
1226 |
15.09 |
|
|
|
| 18 |
A |
1262 |
1208 |
17.04 |
|
|
|
| 19 |
A |
1181 |
1130 |
8.69 |
|
|
|
| 20 |
A |
1130 |
1081 |
4.64 |
|
|
|
| 21 |
A |
1125 |
1076 |
3.76 |
|
|
|
| 22 |
A |
1075 |
1029 |
1.42 |
|
|
|
| 23 |
A |
1035 |
991 |
16.96 |
|
|
|
| 24 |
A |
960 |
919 |
1.36 |
|
|
|
| 25 |
A |
921 |
881 |
5.85 |
|
|
|
| 26 |
A |
782 |
748 |
17.03 |
|
|
|
| 27 |
A |
767 |
734 |
26.96 |
|
|
|
| 28 |
A |
635 |
608 |
26.60 |
|
|
|
| 29 |
A |
429 |
410 |
2.17 |
|
|
|
| 30 |
A |
407 |
390 |
5.26 |
|
|
|
| 31 |
A |
331 |
317 |
3.26 |
|
|
|
| 32 |
A |
250 |
239 |
0.24 |
|
|
|
| 33 |
A |
241 |
231 |
0.15 |
|
|
|
| 34 |
A |
146 |
140 |
2.26 |
|
|
|
| 35 |
A |
120 |
115 |
1.00 |
|
|
|
| 36 |
A |
74 |
71 |
3.29 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25243.5 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 24160.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.339 |
|
|
|
| 2 |
H |
0.133 |
|
|
|
| 3 |
H |
0.138 |
|
|
|
| 4 |
H |
0.118 |
|
|
|
| 5 |
C |
-0.077 |
|
|
|
| 6 |
H |
0.156 |
|
|
|
| 7 |
Cl |
-0.171 |
|
|
|
| 8 |
C |
-0.225 |
|
|
|
| 9 |
H |
0.139 |
|
|
|
| 10 |
H |
0.153 |
|
|
|
| 11 |
C |
-0.158 |
|
|
|
| 12 |
H |
0.141 |
|
|
|
| 13 |
H |
0.162 |
|
|
|
| 14 |
Cl |
-0.169 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.373 |
1.667 |
1.007 |
2.382 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-58.688 |
-3.570 |
0.074 |
| y |
-3.570 |
-50.814 |
0.128 |
| z |
0.074 |
0.128 |
-49.993 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-8.284 |
-3.570 |
0.074 |
| y |
-3.570 |
3.527 |
0.128 |
| z |
0.074 |
0.128 |
4.757 |
|
| Polar |
|---|
| 3z2-r2 | 9.514 |
|---|
| x2-y2 | -7.874 |
|---|
| xy | -3.570 |
|---|
| xz | 0.074 |
|---|
| yz | 0.128 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.916 |
0.204 |
-0.208 |
| y |
0.204 |
10.553 |
0.071 |
| z |
-0.208 |
0.071 |
8.497 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |