return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H5N ((Z)-2-Butenenitrile)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-209.787791
Energy at 298.15K-209.792391
HF Energy-208.878392
Nuclear repulsion energy142.500535
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3255 3090 2.52      
2 A' 3215 3052 4.90      
3 A' 3180 3019 1.89      
4 A' 3089 2933 6.65      
5 A' 2206 2095 0.11      
6 A' 1707 1620 3.70      
7 A' 1508 1432 13.37      
8 A' 1444 1371 4.62      
9 A' 1408 1337 3.42      
10 A' 1273 1209 0.24      
11 A' 1135 1078 0.22      
12 A' 981 931 12.87      
13 A' 924 877 1.86      
14 A' 661 628 1.76      
15 A' 389 369 0.13      
16 A' 155 147 4.53      
17 A" 3151 2991 5.65      
18 A" 1508 1432 7.72      
19 A" 1084 1029 1.14      
20 A" 1010 959 0.69      
21 A" 776 737 36.33      
22 A" 532 506 2.78      
23 A" 290 276 2.84      
24 A" 167 158 0.04      

Unscaled Zero Point Vibrational Energy (zpe) 17523.5 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 16636.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
0.40016 0.11830 0.09286

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.207 0.238 0.000
N2 -2.198 -0.372 0.000
C3 0.000 0.994 0.000
H4 -0.110 2.064 0.000
C5 1.204 0.422 0.000
H6 2.064 1.073 0.000
C7 1.467 -1.038 0.000
H8 0.546 -1.609 0.000
H9 2.051 -1.311 0.875
H10 2.051 -1.311 -0.875

Atom - Atom Distances (Å)
  C1 N2 C3 H4 C5 H6 C7 H8 H9 H10
C11.16341.42432.12992.41783.37552.96292.54663.71153.7115
N21.16342.58773.20773.49324.49983.72533.01034.43834.4383
C31.42432.58771.07521.33302.06522.50632.66023.20673.2067
H42.12993.20771.07522.10292.38903.47963.73124.10134.1013
C52.41783.49321.33302.10291.07841.48322.13512.11762.1176
H63.37554.49982.06522.38901.07842.19323.08172.53902.5390
C72.96293.72532.50633.47961.48322.19321.08411.08651.0865
H82.54663.01032.66023.73122.13513.08171.08411.76591.7659
H93.71154.43833.20674.10132.11762.53901.08651.76591.7501
H103.71154.43833.20674.10132.11762.53901.08651.76591.7501

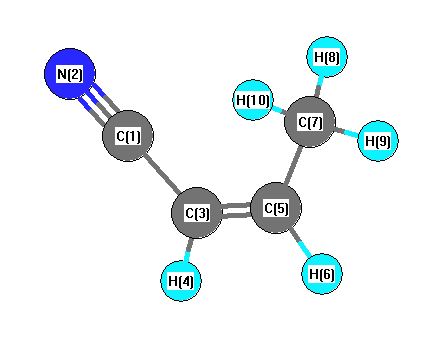
picture of (Z)-2-Butenenitrile state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C3 H4 116.192 C1 C3 C5 122.502
N2 C1 C3 179.517 C3 C5 H6 117.448
C3 C5 C7 125.654 H4 C3 C5 121.305
C5 C7 H8 111.589 C5 C7 H9 110.028
C5 C7 H10 110.028 H6 C5 C7 116.897
H8 C7 H9 108.895 H8 C7 H10 108.895
H9 C7 H10 107.297
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability