return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H6O (Cyclobutanone)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-230.842791
Energy at 298.15K-230.849686
HF Energy-229.877980
Nuclear repulsion energy168.748964
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3199 3037 16.17      
2 A' 3179 3018 2.28      
3 A' 3139 2981 18.51      
4 A' 3117 2959 4.49      
5 A' 1855 1761 237.92      
6 A' 1519 1442 0.62      
7 A' 1471 1397 3.89      
8 A' 1240 1177 7.54      
9 A' 1237 1174 5.11      
10 A' 1102 1046 0.19      
11 A' 999 949 1.94      
12 A' 889 844 1.11      
13 A' 741 703 3.53      
14 A' 684 649 2.87      
15 A' 404 383 0.47      
16 A' 114 108 2.87      
17 A" 3183 3022 7.40      
18 A" 3114 2956 6.58      
19 A" 1453 1379 16.96      
20 A" 1278 1213 6.93      
21 A" 1251 1188 3.10      
22 A" 1178 1118 0.82      
23 A" 1107 1051 68.16      
24 A" 957 909 0.57      
25 A" 942 895 0.10      
26 A" 651 618 0.27      
27 A" 469 446 3.37      

Unscaled Zero Point Vibrational Energy (zpe) 20235.1 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 19211.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
0.36060 0.16334 0.12244

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
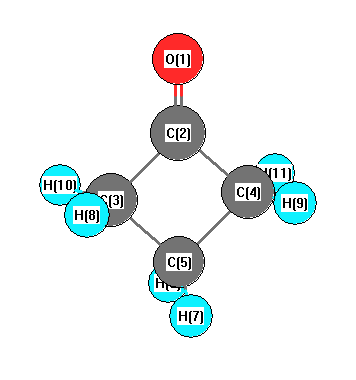
O1 -0.091 1.865 0.000
C2 -0.033 0.670 0.000
C3 0.035 -0.382 1.104
C4 0.035 -0.382 -1.104
C5 0.035 -1.466 0.000
H6 0.915 -2.094 0.000
H7 -0.846 -2.095 0.000
H8 -0.823 -0.374 1.768
H9 -0.823 -0.374 -1.768
H10 0.939 -0.316 1.700
H11 0.939 -0.316 -1.700

Atom - Atom Distances (Å)
  O1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11
O11.19602.50662.50663.33344.08454.03132.94532.94532.95122.9512
C21.19601.52681.52682.13752.92212.88232.20002.20002.19322.1932
C32.50661.52682.20881.54772.21932.22051.08432.99761.08542.9479
C42.50661.52682.20881.54772.21932.22052.99761.08432.94791.0854
C53.33342.13751.54771.54771.08091.08242.24802.24802.24332.2433
H64.08452.92212.21932.21931.08091.76103.01703.01702.46012.4601
H74.03132.88232.22052.22051.08241.76102.46722.46723.04043.0404
H82.94532.20001.08432.99762.24803.01702.46723.53581.76453.8908
H92.94532.20002.99761.08432.24803.01702.46723.53583.89081.7645
H102.95122.19321.08542.94792.24332.46013.04041.76453.89083.4009
H112.95122.19322.94791.08542.24332.46013.04043.89081.76453.4009

picture of Cyclobutanone state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
O1 C2 C3 133.663 O1 C2 C4 133.663
C2 C3 C5 88.087 C2 C3 H8 113.744
C2 C3 H10 113.112 C2 C4 C5 88.087
C2 C4 H9 113.744 C2 C4 H11 113.112
C3 C2 C4 92.660 C3 C5 C4 91.051
C3 C5 H6 114.009 C3 C5 H7 114.016
C4 C5 H6 114.009 C4 C5 H7 114.016
C5 C3 H8 116.208 C5 C3 H10 115.735
C5 C4 H9 116.208 C5 C4 H11 115.735
H6 C5 H7 108.979 H8 C3 H10 108.829
H9 C4 H11 108.829
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability