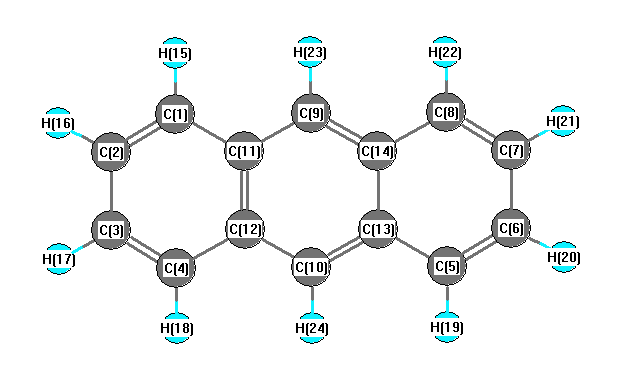
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3209 |
3071 |
0.00 |
|
|
|
| 2 |
Ag |
3183 |
3046 |
0.00 |
|
|
|
| 3 |
Ag |
3175 |
3039 |
0.00 |
|
|
|
| 4 |
Ag |
1617 |
1548 |
0.00 |
|
|
|
| 5 |
Ag |
1527 |
1462 |
0.00 |
|
|
|
| 6 |
Ag |
1457 |
1395 |
0.00 |
|
|
|
| 7 |
Ag |
1304 |
1248 |
0.00 |
|
|
|
| 8 |
Ag |
1183 |
1133 |
0.00 |
|
|
|
| 9 |
Ag |
1040 |
995 |
0.00 |
|
|
|
| 10 |
Ag |
771 |
738 |
0.00 |
|
|
|
| 11 |
Ag |
626 |
599 |
0.00 |
|
|
|
| 12 |
Ag |
398 |
381 |
0.00 |
|
|
|
| 13 |
Au |
1002 |
959 |
0.00 |
|
|
|
| 14 |
Au |
875 |
837 |
0.00 |
|
|
|
| 15 |
Au |
765 |
732 |
0.00 |
|
|
|
| 16 |
Au |
503 |
482 |
0.00 |
|
|
|
| 17 |
Au |
122 |
117 |
0.00 |
|
|
|
| 18 |
B1g |
979 |
937 |
0.00 |
|
|
|
| 19 |
B1g |
775 |
742 |
0.00 |
|
|
|
| 20 |
B1g |
485 |
464 |
0.00 |
|
|
|
| 21 |
B1g |
233 |
223 |
0.00 |
|
|
|
| 22 |
B1u |
3196 |
3059 |
55.42 |
|
|
|
| 23 |
B1u |
3177 |
3041 |
9.23 |
|
|
|
| 24 |
B1u |
3173 |
3037 |
14.81 |
|
|
|
| 25 |
B1u |
1692 |
1619 |
5.24 |
|
|
|
| 26 |
B1u |
1494 |
1430 |
1.45 |
|
|
|
| 27 |
B1u |
1345 |
1288 |
6.57 |
|
|
|
| 28 |
B1u |
1291 |
1235 |
4.90 |
|
|
|
| 29 |
B1u |
1172 |
1122 |
6.99 |
|
|
|
| 30 |
B1u |
915 |
875 |
1.54 |
|
|
|
| 31 |
B1u |
657 |
629 |
0.71 |
|
|
|
| 32 |
B1u |
225 |
216 |
1.27 |
|
|
|
| 33 |
B2g |
1003 |
959 |
0.00 |
|
|
|
| 34 |
B2g |
926 |
887 |
0.00 |
|
|
|
| 35 |
B2g |
853 |
816 |
0.00 |
|
|
|
| 36 |
B2g |
791 |
757 |
0.00 |
|
|
|
| 37 |
B2g |
595 |
570 |
0.00 |
|
|
|
| 38 |
B2g |
269 |
258 |
0.00 |
|
|
|
| 39 |
B2u |
3209 |
3071 |
51.64 |
|
|
|
| 40 |
B2u |
3182 |
3045 |
0.02 |
|
|
|
| 41 |
B2u |
1599 |
1530 |
6.41 |
|
|
|
| 42 |
B2u |
1488 |
1424 |
1.59 |
|
|
|
| 43 |
B2u |
1441 |
1379 |
2.18 |
|
|
|
| 44 |
B2u |
1392 |
1332 |
3.14 |
|
|
|
| 45 |
B2u |
1190 |
1139 |
1.97 |
|
|
|
| 46 |
B2u |
1160 |
1110 |
1.25 |
|
|
|
| 47 |
B2u |
1034 |
990 |
6.61 |
|
|
|
| 48 |
B2u |
831 |
795 |
0.01 |
|
|
|
| 49 |
B2u |
607 |
581 |
8.49 |
|
|
|
| 50 |
B3g |
3196 |
3059 |
0.00 |
|
|
|
| 51 |
B3g |
3177 |
3041 |
0.00 |
|
|
|
| 52 |
B3g |
1691 |
1618 |
0.00 |
|
|
|
| 53 |
B3g |
1643 |
1572 |
0.00 |
|
|
|
| 54 |
B3g |
1414 |
1354 |
0.00 |
|
|
|
| 55 |
B3g |
1293 |
1237 |
0.00 |
|
|
|
| 56 |
B3g |
1207 |
1155 |
0.00 |
|
|
|
| 57 |
B3g |
1131 |
1083 |
0.00 |
|
|
|
| 58 |
B3g |
925 |
885 |
0.00 |
|
|
|
| 59 |
B3g |
522 |
500 |
0.00 |
|
|
|
| 60 |
B3g |
382 |
365 |
0.00 |
|
|
|
| 61 |
B3u |
984 |
942 |
7.40 |
|
|
|
| 62 |
B3u |
910 |
871 |
54.66 |
|
|
|
| 63 |
B3u |
745 |
713 |
76.64 |
|
|
|
| 64 |
B3u |
478 |
458 |
21.85 |
|
|
|
| 65 |
B3u |
387 |
370 |
0.03 |
|
|
|
| 66 |
B3u |
89 |
85 |
1.20 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 42654.6 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 40824.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.146 |
|
|
|
| 2 |
C |
-0.156 |
|
|
|
| 3 |
C |
-0.156 |
|
|
|
| 4 |
C |
-0.146 |
|
|
|
| 5 |
C |
-0.146 |
|
|
|
| 6 |
C |
-0.156 |
|
|
|
| 7 |
C |
-0.156 |
|
|
|
| 8 |
C |
-0.146 |
|
|
|
| 9 |
C |
-0.224 |
|
|
|
| 10 |
C |
-0.224 |
|
|
|
| 11 |
C |
0.118 |
|
|
|
| 12 |
C |
0.118 |
|
|
|
| 13 |
C |
0.118 |
|
|
|
| 14 |
C |
0.118 |
|
|
|
| 15 |
H |
0.116 |
|
|
|
| 16 |
H |
0.126 |
|
|
|
| 17 |
H |
0.126 |
|
|
|
| 18 |
H |
0.116 |
|
|
|
| 19 |
H |
0.116 |
|
|
|
| 20 |
H |
0.126 |
|
|
|
| 21 |
H |
0.126 |
|
|
|
| 22 |
H |
0.116 |
|
|
|
| 23 |
H |
0.107 |
|
|
|
| 24 |
H |
0.107 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-86.893 |
0.000 |
0.000 |
| y |
0.000 |
-70.229 |
0.000 |
| z |
0.000 |
0.000 |
-70.881 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-16.338 |
0.000 |
0.000 |
| y |
0.000 |
8.658 |
0.000 |
| z |
0.000 |
0.000 |
7.680 |
|
| Polar |
|---|
| 3z2-r2 | 15.360 |
|---|
| x2-y2 | -16.664 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.110 |
0.000 |
0.000 |
| y |
0.000 |
41.485 |
0.000 |
| z |
0.000 |
0.000 |
23.561 |
<r2> (average value of r
2) Å
2
| <r2> |
789.556 |
| (<r2>)1/2 |
28.099 |