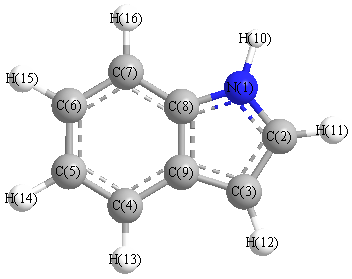
Vibrational Frequencies calculated at mPW1PW91/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3718 |
3567 |
72.08 |
|
|
|
| 2 |
A' |
3281 |
3147 |
2.62 |
|
|
|
| 3 |
A' |
3262 |
3129 |
0.79 |
|
|
|
| 4 |
A' |
3217 |
3085 |
15.70 |
|
|
|
| 5 |
A' |
3205 |
3075 |
27.11 |
|
|
|
| 6 |
A' |
3195 |
3064 |
2.81 |
|
|
|
| 7 |
A' |
3188 |
3058 |
1.08 |
|
|
|
| 8 |
A' |
1687 |
1618 |
2.53 |
|
|
|
| 9 |
A' |
1644 |
1577 |
1.48 |
|
|
|
| 10 |
A' |
1572 |
1507 |
6.89 |
|
|
|
| 11 |
A' |
1542 |
1480 |
3.58 |
|
|
|
| 12 |
A' |
1500 |
1439 |
23.63 |
|
|
|
| 13 |
A' |
1467 |
1407 |
14.18 |
|
|
|
| 14 |
A' |
1394 |
1337 |
22.30 |
|
|
|
| 15 |
A' |
1391 |
1334 |
16.87 |
|
|
|
| 16 |
A' |
1318 |
1264 |
11.51 |
|
|
|
| 17 |
A' |
1277 |
1225 |
9.44 |
|
|
|
| 18 |
A' |
1237 |
1187 |
3.15 |
|
|
|
| 19 |
A' |
1177 |
1129 |
1.01 |
|
|
|
| 20 |
A' |
1151 |
1104 |
0.25 |
|
|
|
| 21 |
A' |
1120 |
1075 |
26.12 |
|
|
|
| 22 |
A' |
1097 |
1052 |
4.33 |
|
|
|
| 23 |
A' |
1046 |
1003 |
6.20 |
|
|
|
| 24 |
A' |
917 |
880 |
6.30 |
|
|
|
| 25 |
A' |
892 |
856 |
0.37 |
|
|
|
| 26 |
A' |
783 |
751 |
2.68 |
|
|
|
| 27 |
A' |
619 |
594 |
1.18 |
|
|
|
| 28 |
A' |
554 |
531 |
0.05 |
|
|
|
| 29 |
A' |
405 |
389 |
4.21 |
|
|
|
| 30 |
A" |
991 |
951 |
0.04 |
|
|
|
| 31 |
A" |
952 |
914 |
2.07 |
|
|
|
| 32 |
A" |
880 |
845 |
2.42 |
|
|
|
| 33 |
A" |
867 |
832 |
0.67 |
|
|
|
| 34 |
A" |
790 |
758 |
15.41 |
|
|
|
| 35 |
A" |
760 |
729 |
86.43 |
|
|
|
| 36 |
A" |
737 |
707 |
27.44 |
|
|
|
| 37 |
A" |
625 |
599 |
6.35 |
|
|
|
| 38 |
A" |
593 |
568 |
0.65 |
|
|
|
| 39 |
A" |
435 |
417 |
0.82 |
|
|
|
| 40 |
A" |
412 |
395 |
67.59 |
|
|
|
| 41 |
A" |
248 |
238 |
0.14 |
|
|
|
| 42 |
A" |
216 |
207 |
10.72 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28679.6 cm
-1
Scaled (by 0.9592) Zero Point Vibrational Energy (zpe) 27509.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.056 |
|
|
|
| 2 |
C |
-0.071 |
|
|
|
| 3 |
C |
-0.226 |
|
|
|
| 4 |
C |
-0.124 |
|
|
|
| 5 |
C |
-0.122 |
|
|
|
| 6 |
C |
-0.106 |
|
|
|
| 7 |
C |
-0.171 |
|
|
|
| 8 |
C |
0.060 |
|
|
|
| 9 |
C |
0.122 |
|
|
|
| 10 |
H |
0.136 |
|
|
|
| 11 |
H |
0.108 |
|
|
|
| 12 |
H |
0.094 |
|
|
|
| 13 |
H |
0.088 |
|
|
|
| 14 |
H |
0.093 |
|
|
|
| 15 |
H |
0.097 |
|
|
|
| 16 |
H |
0.078 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.906 |
2.003 |
0.000 |
2.198 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-46.611 |
-2.131 |
0.000 |
| y |
-2.131 |
-41.908 |
0.000 |
| z |
0.000 |
0.000 |
-57.287 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.987 |
-2.131 |
0.000 |
| y |
-2.131 |
10.041 |
0.000 |
| z |
0.000 |
0.000 |
-13.028 |
|
| Polar |
|---|
| 3z2-r2 | -26.057 |
|---|
| x2-y2 | -4.703 |
|---|
| xy | -2.131 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
18.954 |
-1.517 |
0.000 |
| y |
-1.517 |
15.330 |
0.000 |
| z |
0.000 |
0.000 |
6.885 |
<r2> (average value of r
2) Å
2
| <r2> |
279.369 |
| (<r2>)1/2 |
16.714 |