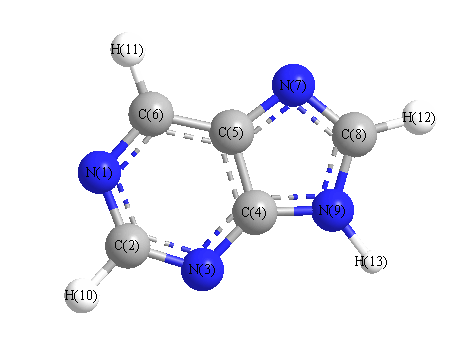
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3689 |
3531 |
92.05 |
|
|
|
| 2 |
A' |
3257 |
3117 |
0.63 |
|
|
|
| 3 |
A' |
3196 |
3059 |
14.73 |
|
|
|
| 4 |
A' |
3185 |
3048 |
15.28 |
|
|
|
| 5 |
A' |
1683 |
1611 |
66.90 |
|
|
|
| 6 |
A' |
1650 |
1579 |
85.04 |
|
|
|
| 7 |
A' |
1553 |
1487 |
27.98 |
|
|
|
| 8 |
A' |
1509 |
1444 |
2.15 |
|
|
|
| 9 |
A' |
1452 |
1390 |
69.66 |
|
|
|
| 10 |
A' |
1431 |
1370 |
26.92 |
|
|
|
| 11 |
A' |
1386 |
1326 |
55.61 |
|
|
|
| 12 |
A' |
1329 |
1272 |
13.85 |
|
|
|
| 13 |
A' |
1320 |
1264 |
13.46 |
|
|
|
| 14 |
A' |
1294 |
1239 |
34.89 |
|
|
|
| 15 |
A' |
1217 |
1165 |
6.56 |
|
|
|
| 16 |
A' |
1153 |
1104 |
6.62 |
|
|
|
| 17 |
A' |
1096 |
1049 |
14.11 |
|
|
|
| 18 |
A' |
951 |
910 |
1.16 |
|
|
|
| 19 |
A' |
914 |
875 |
12.23 |
|
|
|
| 20 |
A' |
815 |
780 |
14.96 |
|
|
|
| 21 |
A' |
660 |
631 |
0.43 |
|
|
|
| 22 |
A' |
568 |
543 |
4.11 |
|
|
|
| 23 |
A' |
448 |
429 |
12.32 |
|
|
|
| 24 |
A" |
1012 |
969 |
0.30 |
|
|
|
| 25 |
A" |
959 |
918 |
9.25 |
|
|
|
| 26 |
A" |
902 |
863 |
5.55 |
|
|
|
| 27 |
A" |
828 |
792 |
9.78 |
|
|
|
| 28 |
A" |
671 |
643 |
6.29 |
|
|
|
| 29 |
A" |
628 |
602 |
22.82 |
|
|
|
| 30 |
A" |
549 |
526 |
86.24 |
|
|
|
| 31 |
A" |
421 |
403 |
2.98 |
|
|
|
| 32 |
A" |
247 |
236 |
0.63 |
|
|
|
| 33 |
A" |
233 |
223 |
4.65 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21102.9 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 20197.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.193 |
|
|
|
| 2 |
C |
0.023 |
|
|
|
| 3 |
N |
-0.200 |
|
|
|
| 4 |
C |
0.124 |
|
|
|
| 5 |
C |
0.018 |
|
|
|
| 6 |
C |
-0.096 |
|
|
|
| 7 |
N |
-0.210 |
|
|
|
| 8 |
C |
-0.001 |
|
|
|
| 9 |
N |
-0.075 |
|
|
|
| 10 |
H |
0.133 |
|
|
|
| 11 |
H |
0.149 |
|
|
|
| 12 |
H |
0.151 |
|
|
|
| 13 |
H |
0.177 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.182 |
2.923 |
0.000 |
3.647 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-49.941 |
6.133 |
0.000 |
| y |
6.133 |
-46.848 |
0.000 |
| z |
0.000 |
0.000 |
-51.357 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.838 |
6.133 |
0.000 |
| y |
6.133 |
3.801 |
0.000 |
| z |
0.000 |
0.000 |
-2.962 |
|
| Polar |
|---|
| 3z2-r2 | -5.925 |
|---|
| x2-y2 | -3.093 |
|---|
| xy | 6.133 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
15.567 |
1.367 |
0.000 |
| y |
1.367 |
12.210 |
0.000 |
| z |
0.000 |
0.000 |
5.904 |
<r2> (average value of r
2) Å
2
| <r2> |
250.566 |
| (<r2>)1/2 |
15.829 |