return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H8O (Cyclopentanone)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1A
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-270.115067
Energy at 298.15K-270.124774
HF Energy-268.959055
Nuclear repulsion energy237.242807
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3170 3010 3.74      
2 A 3157 2997 18.90      
3 A 3093 2936 17.62      
4 A 3085 2929 0.02      
5 A 1808 1717 166.44      
6 A 1528 1451 1.51      
7 A 1470 1395 0.10      
8 A 1352 1284 0.15      
9 A 1319 1252 2.80      
10 A 1236 1173 0.31      
11 A 1188 1128 0.08      
12 A 1057 1004 0.71      
13 A 969 920 0.24      
14 A 929 882 0.55      
15 A 836 794 1.38      
16 A 722 685 1.43      
17 A 561 533 4.59      
18 A 263 250 0.12      
19 B 3170 3010 11.88      
20 B 3160 3000 22.85      
21 B 3100 2943 33.42      
22 B 3087 2931 3.27      
23 B 1515 1438 5.80      
24 B 1470 1395 19.74      
25 B 1352 1283 2.50      
26 B 1314 1248 8.47      
27 B 1267 1203 7.31      
28 B 1187 1127 59.02      
29 B 1170 1111 1.90      
30 B 990 940 10.36      
31 B 930 883 0.36      
32 B 864 821 9.95      
33 B 583 553 1.97      
34 B 477 453 5.67      
35 B 452 429 2.54      
36 B 97 92 4.19      

Unscaled Zero Point Vibrational Energy (zpe) 26964.1 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 25599.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
0.22533 0.11321 0.08177

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is C2

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
O1 0.000 0.000 2.120
C2 0.000 0.000 0.912
C3 0.000 1.229 0.023
C4 0.000 -1.229 0.023
C5 0.327 0.692 -1.362
C6 -0.327 -0.692 -1.362
H7 -1.010 1.636 0.048
H8 1.010 -1.636 0.048
H9 0.671 1.987 0.408
H10 -0.671 -1.987 0.408
H11 -0.022 1.327 -2.168
H12 0.022 -1.327 -2.168
H13 1.404 0.580 -1.468
H14 -1.404 -0.580 -1.468

Atom - Atom Distances (Å)
  O1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14
O11.20872.43132.43133.56563.56562.82732.82732.70752.70754.48874.48873.89673.8967
C21.20871.51701.51702.39912.39912.10822.10822.15682.15683.35323.35322.82312.8231
C32.43131.51702.45831.52112.39041.08953.03851.08313.30782.19283.36612.14812.7323
C42.43131.51702.45832.39041.52113.03851.08953.30781.08313.36612.19282.73232.1481
C53.56562.39911.52112.39041.52962.16062.80622.22043.36201.08352.19421.08792.1497
C63.56562.39912.39041.52111.52962.80622.16063.36202.22042.19421.08352.14971.0879
H72.82732.10821.08953.03852.16062.80623.84621.75493.65682.44573.84123.03992.7139
H82.82732.10823.03851.08952.80622.16063.84623.65681.75493.84122.44572.71393.0399
H92.70752.15681.08313.30782.22043.36201.75493.65684.19442.74804.24672.45683.7964
H102.70752.15683.30781.08313.36202.22043.65681.75494.19444.24672.74803.79642.4568
H114.48873.35322.19283.36611.08352.19422.44573.84122.74804.24672.65351.75502.4559
H124.48873.35323.36612.19282.19421.08353.84122.44574.24672.74802.65352.45591.7550
H133.89672.82312.14812.73231.08792.14973.03992.71392.45683.79641.75502.45593.0372
H143.89672.82312.73232.14812.14971.08792.71393.03993.79642.45682.45591.75503.0372

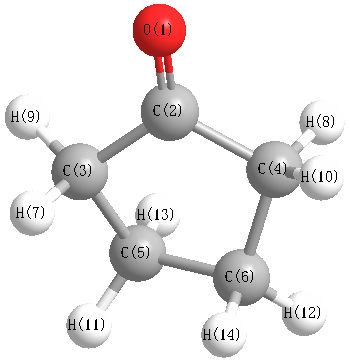
picture of Cyclopentanone state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
O1 C2 C3 125.876 O1 C2 C4 125.876
C2 C3 C5 104.310 C2 C3 H7 106.816
C2 C3 H9 111.002 C2 C4 C6 104.310
C2 C4 H8 106.816 C2 C4 H10 111.002
C3 C2 C4 108.248 C3 C5 C6 103.174
C3 C5 H11 113.621 C3 C5 H13 109.728
C4 C6 C5 103.174 C4 C6 H12 113.621
C4 C6 H14 109.728 C5 C3 H7 110.624
C5 C3 H9 115.981 C5 C6 H12 113.104
C5 C6 H14 109.264 C6 C4 H8 110.624
C6 C4 H10 115.981 C6 C5 H11 113.104
C6 C5 H13 109.264 H7 C3 H9 107.756
H8 C4 H10 107.756 H11 C5 H13 107.848
H12 C6 H14 107.848
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability