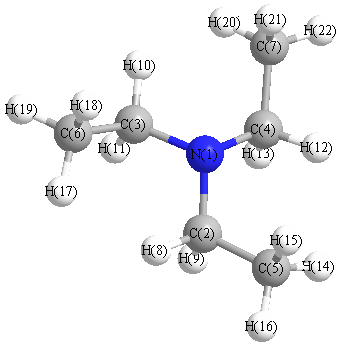
Vibrational Frequencies calculated at mPW1PW91/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3134 |
3006 |
51.42 |
|
|
|
| 2 |
A |
3123 |
2995 |
1.05 |
|
|
|
| 3 |
A |
3095 |
2969 |
0.01 |
|
|
|
| 4 |
A |
3055 |
2930 |
9.60 |
|
|
|
| 5 |
A |
2939 |
2819 |
207.11 |
|
|
|
| 6 |
A |
1515 |
1453 |
12.74 |
|
|
|
| 7 |
A |
1490 |
1429 |
2.64 |
|
|
|
| 8 |
A |
1485 |
1424 |
0.91 |
|
|
|
| 9 |
A |
1422 |
1364 |
14.90 |
|
|
|
| 10 |
A |
1403 |
1346 |
1.84 |
|
|
|
| 11 |
A |
1323 |
1269 |
5.31 |
|
|
|
| 12 |
A |
1174 |
1126 |
12.31 |
|
|
|
| 13 |
A |
1092 |
1048 |
1.84 |
|
|
|
| 14 |
A |
1044 |
1002 |
8.79 |
|
|
|
| 15 |
A |
814 |
781 |
0.04 |
|
|
|
| 16 |
A |
761 |
730 |
10.67 |
|
|
|
| 17 |
A |
434 |
417 |
2.98 |
|
|
|
| 18 |
A |
300 |
288 |
1.14 |
|
|
|
| 19 |
A |
184 |
176 |
0.62 |
|
|
|
| 20 |
A |
87 |
83 |
0.02 |
|
|
|
| 21 |
E |
3135 |
3007 |
23.03 |
|
|
|
| 21 |
E |
3135 |
3007 |
23.04 |
|
|
|
| 22 |
E |
3126 |
2998 |
54.70 |
|
|
|
| 22 |
E |
3126 |
2998 |
54.70 |
|
|
|
| 23 |
E |
3094 |
2968 |
14.32 |
|
|
|
| 23 |
E |
3094 |
2968 |
14.31 |
|
|
|
| 24 |
E |
3054 |
2930 |
28.27 |
|
|
|
| 24 |
E |
3054 |
2930 |
28.27 |
|
|
|
| 25 |
E |
2928 |
2809 |
19.17 |
|
|
|
| 25 |
E |
2928 |
2809 |
19.16 |
|
|
|
| 26 |
E |
1509 |
1448 |
6.24 |
|
|
|
| 26 |
E |
1509 |
1448 |
6.24 |
|
|
|
| 27 |
E |
1499 |
1438 |
1.65 |
|
|
|
| 27 |
E |
1499 |
1438 |
1.65 |
|
|
|
| 28 |
E |
1486 |
1425 |
1.85 |
|
|
|
| 28 |
E |
1486 |
1425 |
1.85 |
|
|
|
| 29 |
E |
1418 |
1361 |
22.68 |
|
|
|
| 29 |
E |
1418 |
1361 |
22.68 |
|
|
|
| 30 |
E |
1395 |
1338 |
0.88 |
|
|
|
| 30 |
E |
1395 |
1338 |
0.88 |
|
|
|
| 31 |
E |
1332 |
1277 |
22.20 |
|
|
|
| 31 |
E |
1332 |
1277 |
22.21 |
|
|
|
| 32 |
E |
1247 |
1196 |
24.29 |
|
|
|
| 32 |
E |
1247 |
1196 |
24.28 |
|
|
|
| 33 |
E |
1110 |
1065 |
20.46 |
|
|
|
| 33 |
E |
1110 |
1065 |
20.46 |
|
|
|
| 34 |
E |
1100 |
1056 |
9.55 |
|
|
|
| 34 |
E |
1100 |
1056 |
9.55 |
|
|
|
| 35 |
E |
937 |
899 |
1.00 |
|
|
|
| 35 |
E |
937 |
899 |
1.00 |
|
|
|
| 36 |
E |
801 |
768 |
3.48 |
|
|
|
| 36 |
E |
801 |
768 |
3.48 |
|
|
|
| 37 |
E |
477 |
457 |
2.12 |
|
|
|
| 37 |
E |
477 |
457 |
2.12 |
|
|
|
| 38 |
E |
306 |
293 |
0.36 |
|
|
|
| 38 |
E |
306 |
293 |
0.36 |
|
|
|
| 39 |
E |
209 |
200 |
0.04 |
|
|
|
| 39 |
E |
209 |
200 |
0.04 |
|
|
|
| 40 |
E |
86 |
82 |
0.21 |
|
|
|
| 40 |
E |
86 |
82 |
0.21 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 45185.3 cm
-1
Scaled (by 0.9592) Zero Point Vibrational Energy (zpe) 43341.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.191 |
|
|
|
| 2 |
C |
-0.069 |
|
|
|
| 3 |
C |
-0.069 |
|
|
|
| 4 |
C |
-0.069 |
|
|
|
| 5 |
C |
-0.256 |
|
|
|
| 6 |
C |
-0.256 |
|
|
|
| 7 |
C |
-0.256 |
|
|
|
| 8 |
H |
0.083 |
|
|
|
| 9 |
H |
0.054 |
|
|
|
| 10 |
H |
0.083 |
|
|
|
| 11 |
H |
0.054 |
|
|
|
| 12 |
H |
0.083 |
|
|
|
| 13 |
H |
0.054 |
|
|
|
| 14 |
H |
0.082 |
|
|
|
| 15 |
H |
0.088 |
|
|
|
| 16 |
H |
0.081 |
|
|
|
| 17 |
H |
0.082 |
|
|
|
| 18 |
H |
0.088 |
|
|
|
| 19 |
H |
0.081 |
|
|
|
| 20 |
H |
0.082 |
|
|
|
| 21 |
H |
0.088 |
|
|
|
| 22 |
H |
0.081 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.508 |
0.508 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-46.350 |
0.000 |
0.000 |
| y |
0.000 |
-46.350 |
0.000 |
| z |
0.000 |
0.000 |
-48.810 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.230 |
0.000 |
0.000 |
| y |
0.000 |
1.230 |
0.000 |
| z |
0.000 |
0.000 |
-2.460 |
|
| Polar |
|---|
| 3z2-r2 | -4.920 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
13.010 |
0.000 |
0.000 |
| y |
0.000 |
13.010 |
0.000 |
| z |
0.000 |
0.000 |
11.178 |
<r2> (average value of r
2) Å
2
| <r2> |
285.988 |
| (<r2>)1/2 |
16.911 |