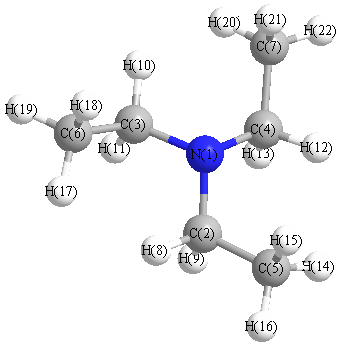
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3140 |
3005 |
52.76 |
|
|
|
| 2 |
A |
3128 |
2994 |
0.98 |
|
|
|
| 3 |
A |
3101 |
2968 |
0.01 |
|
|
|
| 4 |
A |
3059 |
2928 |
9.70 |
|
|
|
| 5 |
A |
2941 |
2815 |
212.65 |
|
|
|
| 6 |
A |
1513 |
1448 |
12.23 |
|
|
|
| 7 |
A |
1489 |
1425 |
3.06 |
|
|
|
| 8 |
A |
1484 |
1420 |
0.73 |
|
|
|
| 9 |
A |
1419 |
1358 |
16.45 |
|
|
|
| 10 |
A |
1399 |
1339 |
0.84 |
|
|
|
| 11 |
A |
1319 |
1263 |
6.22 |
|
|
|
| 12 |
A |
1172 |
1122 |
12.68 |
|
|
|
| 13 |
A |
1089 |
1042 |
1.24 |
|
|
|
| 14 |
A |
1044 |
1000 |
10.21 |
|
|
|
| 15 |
A |
810 |
775 |
0.22 |
|
|
|
| 16 |
A |
761 |
729 |
10.45 |
|
|
|
| 17 |
A |
430 |
412 |
2.88 |
|
|
|
| 18 |
A |
301 |
288 |
1.02 |
|
|
|
| 19 |
A |
183 |
176 |
0.56 |
|
|
|
| 20 |
A |
90 |
86 |
0.02 |
|
|
|
| 21 |
E |
3140 |
3005 |
23.62 |
|
|
|
| 21 |
E |
3140 |
3005 |
23.55 |
|
|
|
| 22 |
E |
3132 |
2997 |
55.24 |
|
|
|
| 22 |
E |
3132 |
2997 |
55.11 |
|
|
|
| 23 |
E |
3100 |
2967 |
14.13 |
|
|
|
| 23 |
E |
3100 |
2967 |
14.08 |
|
|
|
| 24 |
E |
3059 |
2928 |
29.06 |
|
|
|
| 24 |
E |
3059 |
2928 |
28.98 |
|
|
|
| 25 |
E |
2932 |
2806 |
17.96 |
|
|
|
| 25 |
E |
2932 |
2806 |
17.98 |
|
|
|
| 26 |
E |
1507 |
1443 |
4.98 |
|
|
|
| 26 |
E |
1507 |
1443 |
4.98 |
|
|
|
| 27 |
E |
1497 |
1433 |
2.16 |
|
|
|
| 27 |
E |
1497 |
1433 |
2.15 |
|
|
|
| 28 |
E |
1484 |
1420 |
1.68 |
|
|
|
| 28 |
E |
1484 |
1420 |
1.69 |
|
|
|
| 29 |
E |
1414 |
1354 |
22.72 |
|
|
|
| 29 |
E |
1414 |
1354 |
22.72 |
|
|
|
| 30 |
E |
1391 |
1331 |
1.71 |
|
|
|
| 30 |
E |
1391 |
1331 |
1.70 |
|
|
|
| 31 |
E |
1329 |
1272 |
22.28 |
|
|
|
| 31 |
E |
1329 |
1272 |
22.28 |
|
|
|
| 32 |
E |
1244 |
1191 |
24.25 |
|
|
|
| 32 |
E |
1244 |
1191 |
24.24 |
|
|
|
| 33 |
E |
1109 |
1062 |
21.40 |
|
|
|
| 33 |
E |
1109 |
1062 |
21.39 |
|
|
|
| 34 |
E |
1099 |
1052 |
7.67 |
|
|
|
| 34 |
E |
1099 |
1052 |
7.69 |
|
|
|
| 35 |
E |
936 |
896 |
1.20 |
|
|
|
| 35 |
E |
936 |
896 |
1.20 |
|
|
|
| 36 |
E |
797 |
763 |
3.21 |
|
|
|
| 36 |
E |
797 |
763 |
3.22 |
|
|
|
| 37 |
E |
473 |
453 |
2.04 |
|
|
|
| 37 |
E |
473 |
453 |
2.05 |
|
|
|
| 38 |
E |
303 |
290 |
0.39 |
|
|
|
| 38 |
E |
303 |
290 |
0.39 |
|
|
|
| 39 |
E |
205 |
197 |
0.05 |
|
|
|
| 39 |
E |
205 |
197 |
0.05 |
|
|
|
| 40 |
E |
87 |
83 |
0.22 |
|
|
|
| 40 |
E |
87 |
83 |
0.22 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 45174.4 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 43236.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.160 |
|
|
|
| 2 |
C |
-0.149 |
|
|
|
| 3 |
C |
-0.149 |
|
|
|
| 4 |
C |
-0.149 |
|
|
|
| 5 |
C |
-0.348 |
|
|
|
| 6 |
C |
-0.348 |
|
|
|
| 7 |
C |
-0.348 |
|
|
|
| 8 |
H |
0.117 |
|
|
|
| 9 |
H |
0.092 |
|
|
|
| 10 |
H |
0.117 |
|
|
|
| 11 |
H |
0.092 |
|
|
|
| 12 |
H |
0.117 |
|
|
|
| 13 |
H |
0.092 |
|
|
|
| 14 |
H |
0.111 |
|
|
|
| 15 |
H |
0.119 |
|
|
|
| 16 |
H |
0.111 |
|
|
|
| 17 |
H |
0.111 |
|
|
|
| 18 |
H |
0.119 |
|
|
|
| 19 |
H |
0.111 |
|
|
|
| 20 |
H |
0.111 |
|
|
|
| 21 |
H |
0.119 |
|
|
|
| 22 |
H |
0.111 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.505 |
0.505 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-46.393 |
0.000 |
0.000 |
| y |
0.000 |
-46.393 |
0.000 |
| z |
0.000 |
0.000 |
-48.908 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.257 |
0.000 |
0.000 |
| y |
0.000 |
1.257 |
0.000 |
| z |
0.000 |
0.000 |
-2.515 |
|
| Polar |
|---|
| 3z2-r2 | -5.029 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
13.025 |
0.000 |
-0.000 |
| y |
0.000 |
13.026 |
0.000 |
| z |
-0.000 |
0.000 |
11.212 |
<r2> (average value of r
2) Å
2
| <r2> |
284.845 |
| (<r2>)1/2 |
16.877 |