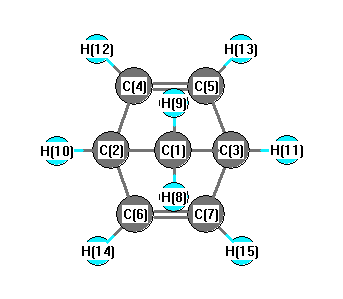
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3244 |
3105 |
7.81 |
|
|
|
| 2 |
A1 |
3138 |
3004 |
8.70 |
|
|
|
| 3 |
A1 |
3068 |
2936 |
45.13 |
|
|
|
| 4 |
A1 |
1672 |
1600 |
0.27 |
|
|
|
| 5 |
A1 |
1475 |
1411 |
2.04 |
|
|
|
| 6 |
A1 |
1262 |
1208 |
4.37 |
|
|
|
| 7 |
A1 |
1129 |
1081 |
0.28 |
|
|
|
| 8 |
A1 |
958 |
917 |
2.40 |
|
|
|
| 9 |
A1 |
904 |
865 |
7.95 |
|
|
|
| 10 |
A1 |
784 |
750 |
4.46 |
|
|
|
| 11 |
A1 |
756 |
723 |
52.31 |
|
|
|
| 12 |
A1 |
425 |
407 |
4.69 |
|
|
|
| 13 |
A2 |
3215 |
3077 |
0.00 |
|
|
|
| 14 |
A2 |
1298 |
1242 |
0.00 |
|
|
|
| 15 |
A2 |
1264 |
1210 |
0.00 |
|
|
|
| 16 |
A2 |
1134 |
1085 |
0.00 |
|
|
|
| 17 |
A2 |
943 |
903 |
0.00 |
|
|
|
| 18 |
A2 |
919 |
880 |
0.00 |
|
|
|
| 19 |
A2 |
719 |
689 |
0.00 |
|
|
|
| 20 |
A2 |
451 |
432 |
0.00 |
|
|
|
| 21 |
B1 |
3241 |
3102 |
17.97 |
|
|
|
| 22 |
B1 |
3131 |
2997 |
34.38 |
|
|
|
| 23 |
B1 |
1629 |
1559 |
9.94 |
|
|
|
| 24 |
B1 |
1233 |
1180 |
2.90 |
|
|
|
| 25 |
B1 |
1079 |
1033 |
0.26 |
|
|
|
| 26 |
B1 |
1024 |
980 |
0.44 |
|
|
|
| 27 |
B1 |
922 |
882 |
1.28 |
|
|
|
| 28 |
B1 |
683 |
654 |
36.05 |
|
|
|
| 29 |
B1 |
495 |
473 |
5.51 |
|
|
|
| 30 |
B2 |
3217 |
3079 |
12.23 |
|
|
|
| 31 |
B2 |
3136 |
3001 |
56.71 |
|
|
|
| 32 |
B2 |
1344 |
1287 |
25.48 |
|
|
|
| 33 |
B2 |
1274 |
1219 |
0.07 |
|
|
|
| 34 |
B2 |
1178 |
1127 |
0.97 |
|
|
|
| 35 |
B2 |
979 |
937 |
0.72 |
|
|
|
| 36 |
B2 |
948 |
908 |
3.08 |
|
|
|
| 37 |
B2 |
896 |
857 |
2.09 |
|
|
|
| 38 |
B2 |
820 |
785 |
7.98 |
|
|
|
| 39 |
B2 |
548 |
524 |
1.66 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28266.9 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 27054.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.186 |
|
|
|
| 2 |
C |
-0.028 |
|
|
|
| 3 |
C |
-0.028 |
|
|
|
| 4 |
C |
-0.170 |
|
|
|
| 5 |
C |
-0.170 |
|
|
|
| 6 |
C |
-0.170 |
|
|
|
| 7 |
C |
-0.170 |
|
|
|
| 8 |
H |
0.105 |
|
|
|
| 9 |
H |
0.105 |
|
|
|
| 10 |
H |
0.088 |
|
|
|
| 11 |
H |
0.088 |
|
|
|
| 12 |
H |
0.134 |
|
|
|
| 13 |
H |
0.134 |
|
|
|
| 14 |
H |
0.134 |
|
|
|
| 15 |
H |
0.134 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.038 |
0.038 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-42.301 |
0.000 |
0.000 |
| y |
0.000 |
-38.766 |
0.000 |
| z |
0.000 |
0.000 |
-42.652 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.593 |
0.000 |
0.000 |
| y |
0.000 |
3.711 |
0.000 |
| z |
0.000 |
0.000 |
-2.118 |
|
| Polar |
|---|
| 3z2-r2 | -4.236 |
|---|
| x2-y2 | -3.535 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.348 |
0.000 |
0.000 |
| y |
0.000 |
11.428 |
0.000 |
| z |
0.000 |
0.000 |
8.984 |
<r2> (average value of r
2) Å
2
| <r2> |
150.691 |
| (<r2>)1/2 |
12.276 |