Jump to
S1C2
S1C3
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -208.876612 |
| Energy at 298.15K | -208.882740 |
| HF Energy | -208.040763 |
| Nuclear repulsion energy | 119.821277 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3665 |
3479 |
39.87 |
|
|
|
| 2 |
A' |
3182 |
3021 |
11.12 |
|
|
|
| 3 |
A' |
3094 |
2937 |
46.97 |
|
|
|
| 4 |
A' |
3011 |
2858 |
76.90 |
|
|
|
| 5 |
A' |
1813 |
1722 |
449.10 |
|
|
|
| 6 |
A' |
1565 |
1486 |
14.61 |
|
|
|
| 7 |
A' |
1501 |
1425 |
9.36 |
|
|
|
| 8 |
A' |
1485 |
1409 |
10.95 |
|
|
|
| 9 |
A' |
1421 |
1349 |
17.62 |
|
|
|
| 10 |
A' |
1324 |
1257 |
103.95 |
|
|
|
| 11 |
A' |
1191 |
1131 |
25.78 |
|
|
|
| 12 |
A' |
1030 |
978 |
42.37 |
|
|
|
| 13 |
A' |
620 |
589 |
11.96 |
|
|
|
| 14 |
A' |
347 |
330 |
7.45 |
|
|
|
| 15 |
A" |
3155 |
2995 |
19.95 |
|
|
|
| 16 |
A" |
1517 |
1441 |
5.61 |
|
|
|
| 17 |
A" |
1174 |
1115 |
0.54 |
|
|
|
| 18 |
A" |
1066 |
1012 |
2.28 |
|
|
|
| 19 |
A" |
634 |
602 |
110.41 |
|
|
|
| 20 |
A" |
216 |
205 |
0.49 |
|
|
|
| 21 |
A" |
109 |
103 |
0.70 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16560.1 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 15722.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.284 |
-0.747 |
0.000 |
| O2 |
1.400 |
-1.224 |
0.000 |
| N3 |
0.000 |
0.575 |
0.000 |
| C4 |
-1.332 |
1.116 |
0.000 |
| H5 |
-0.627 |
-1.360 |
0.000 |
| H6 |
0.792 |
1.193 |
0.000 |
| H7 |
-2.037 |
0.292 |
0.000 |
| H8 |
-1.520 |
1.719 |
0.883 |
| H9 |
-1.520 |
1.719 |
-0.883 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.2139 | 1.3521 | 2.4664 | 1.0976 | 2.0056 | 2.5424 | 3.1804 | 3.1804 |
O2 | 1.2139 | | 2.2796 | 3.5975 | 2.0314 | 2.4924 | 3.7561 | 4.2389 | 4.2389 | N3 | 1.3521 | 2.2796 | | 1.4380 | 2.0335 | 1.0050 | 2.0561 | 2.0974 | 2.0974 | C4 | 2.4664 | 3.5975 | 1.4380 | | 2.5743 | 2.1260 | 1.0841 | 1.0855 | 1.0855 | H5 | 1.0976 | 2.0314 | 2.0335 | 2.5743 | | 2.9206 | 2.1714 | 3.3248 | 3.3248 | H6 | 2.0056 | 2.4924 | 1.0050 | 2.1260 | 2.9206 | | 2.9690 | 2.5306 | 2.5306 | H7 | 2.5424 | 3.7561 | 2.0561 | 1.0841 | 2.1714 | 2.9690 | | 1.7557 | 1.7557 | H8 | 3.1804 | 4.2389 | 2.0974 | 1.0855 | 3.3248 | 2.5306 | 1.7557 | | 1.7660 | H9 | 3.1804 | 4.2389 | 2.0974 | 1.0855 | 3.3248 | 2.5306 | 1.7557 | 1.7660 | |
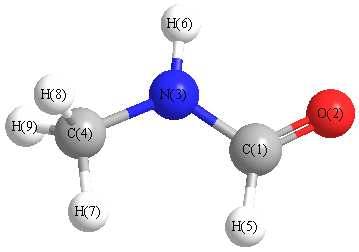 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
C4 |
124.231 |
|
C1 |
N3 |
H6 |
115.835 |
| O2 |
C1 |
N3 |
125.266 |
|
O2 |
C1 |
H5 |
122.929 |
| N3 |
C1 |
H5 |
111.805 |
|
N3 |
C4 |
H7 |
108.405 |
| N3 |
C4 |
H8 |
111.677 |
|
N3 |
C4 |
H9 |
111.677 |
| C4 |
N3 |
H6 |
119.934 |
|
H7 |
C4 |
H8 |
108.039 |
| H7 |
C4 |
H9 |
108.039 |
|
H8 |
C4 |
H9 |
108.871 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -208.876612 |
| Energy at 298.15K | -208.882740 |
| HF Energy | -208.040763 |
| Nuclear repulsion energy | 119.821277 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is Cs
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -208.878675 |
| Energy at 298.15K | -208.884774 |
| HF Energy | -208.042134 |
| Nuclear repulsion energy | 122.041684 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3704 |
3517 |
41.11 |
|
|
|
| 2 |
A |
3206 |
3044 |
0.15 |
|
|
|
| 3 |
A |
3164 |
3004 |
20.16 |
|
|
|
| 4 |
A |
3102 |
2945 |
39.47 |
|
|
|
| 5 |
A |
3030 |
2876 |
98.58 |
|
|
|
| 6 |
A |
1801 |
1710 |
316.40 |
|
|
|
| 7 |
A |
1580 |
1500 |
80.12 |
|
|
|
| 8 |
A |
1534 |
1457 |
6.55 |
|
|
|
| 9 |
A |
1522 |
1445 |
47.64 |
|
|
|
| 10 |
A |
1457 |
1384 |
19.77 |
|
|
|
| 11 |
A |
1445 |
1372 |
3.63 |
|
|
|
| 12 |
A |
1247 |
1183 |
72.17 |
|
|
|
| 13 |
A |
1177 |
1117 |
1.17 |
|
|
|
| 14 |
A |
1171 |
1111 |
15.93 |
|
|
|
| 15 |
A |
1050 |
997 |
0.10 |
|
|
|
| 16 |
A |
989 |
939 |
22.39 |
|
|
|
| 17 |
A |
776 |
737 |
0.64 |
|
|
|
| 18 |
A |
551 |
524 |
44.06 |
|
|
|
| 19 |
A |
302 |
286 |
13.12 |
|
|
|
| 20 |
A |
275 |
261 |
67.12 |
|
|
|
| 21 |
A |
74 |
70 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16578.2 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 15739.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.856 |
0.438 |
0.003 |
| O2 |
1.364 |
-0.669 |
0.000 |
| N3 |
-0.473 |
0.658 |
-0.020 |
| C4 |
-1.406 |
-0.444 |
0.005 |
| H5 |
1.440 |
1.362 |
0.018 |
| H6 |
-0.803 |
1.599 |
0.061 |
| H7 |
-2.399 |
-0.068 |
-0.205 |
| H8 |
-1.127 |
-1.167 |
-0.751 |
| H9 |
-1.404 |
-0.942 |
0.969 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.2180 | 1.3477 | 2.4283 | 1.0936 | 2.0259 | 3.3007 | 2.6608 | 2.8192 |
O2 | 1.2180 | | 2.2663 | 2.7793 | 2.0328 | 3.1376 | 3.8159 | 2.6493 | 2.9455 | N3 | 1.3477 | 2.2663 | | 1.4440 | 2.0398 | 1.0010 | 2.0661 | 2.0719 | 2.0989 | C4 | 2.4283 | 2.7793 | 1.4440 | | 3.3715 | 2.1315 | 1.0819 | 1.0828 | 1.0851 | H5 | 1.0936 | 2.0328 | 2.0398 | 3.3715 | | 2.2563 | 4.1033 | 3.6857 | 3.7826 | H6 | 2.0259 | 3.1376 | 1.0010 | 2.1315 | 2.2563 | | 2.3237 | 2.9017 | 2.7653 | H7 | 3.3007 | 3.8159 | 2.0661 | 1.0819 | 4.1033 | 2.3237 | | 1.7671 | 1.7696 | H8 | 2.6608 | 2.6493 | 2.0719 | 1.0828 | 3.6857 | 2.9017 | 1.7671 | | 1.7567 | H9 | 2.8192 | 2.9455 | 2.0989 | 1.0851 | 3.7826 | 2.7653 | 1.7696 | 1.7567 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
C4 |
120.839 |
|
C1 |
N3 |
H6 |
118.470 |
| O2 |
C1 |
N3 |
124.014 |
|
O2 |
C1 |
H5 |
123.050 |
| N3 |
C1 |
H5 |
112.935 |
|
N3 |
C4 |
H7 |
108.924 |
| N3 |
C4 |
H8 |
109.335 |
|
N3 |
C4 |
H9 |
111.389 |
| C4 |
N3 |
H6 |
120.251 |
|
H7 |
C4 |
H8 |
109.431 |
| H7 |
C4 |
H9 |
109.488 |
|
H8 |
C4 |
H9 |
108.250 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
