Jump to
S1C2
S1C3
Energy calculated at mPW1PW91/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -209.226091 |
| Energy at 298.15K | -209.232190 |
| HF Energy | -209.226091 |
| Nuclear repulsion energy | 119.857231 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3643 |
3495 |
27.61 |
|
|
|
| 2 |
A' |
3138 |
3010 |
13.54 |
|
|
|
| 3 |
A' |
3044 |
2920 |
51.56 |
|
|
|
| 4 |
A' |
2946 |
2826 |
85.30 |
|
|
|
| 5 |
A' |
1827 |
1752 |
496.38 |
|
|
|
| 6 |
A' |
1539 |
1477 |
21.08 |
|
|
|
| 7 |
A' |
1493 |
1433 |
7.55 |
|
|
|
| 8 |
A' |
1468 |
1409 |
5.90 |
|
|
|
| 9 |
A' |
1399 |
1342 |
10.92 |
|
|
|
| 10 |
A' |
1318 |
1264 |
106.62 |
|
|
|
| 11 |
A' |
1179 |
1131 |
25.17 |
|
|
|
| 12 |
A' |
1021 |
979 |
43.46 |
|
|
|
| 13 |
A' |
620 |
595 |
14.03 |
|
|
|
| 14 |
A' |
344 |
330 |
8.17 |
|
|
|
| 15 |
A" |
3100 |
2973 |
25.78 |
|
|
|
| 16 |
A" |
1485 |
1425 |
6.07 |
|
|
|
| 17 |
A" |
1154 |
1107 |
0.27 |
|
|
|
| 18 |
A" |
1050 |
1007 |
1.12 |
|
|
|
| 19 |
A" |
628 |
602 |
109.03 |
|
|
|
| 20 |
A" |
208 |
199 |
0.51 |
|
|
|
| 21 |
A" |
110 |
105 |
0.33 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16356.1 cm
-1
Scaled (by 0.9592) Zero Point Vibrational Energy (zpe) 15688.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.284 |
-0.751 |
0.000 |
| O2 |
1.396 |
-1.223 |
0.000 |
| N3 |
0.000 |
0.569 |
0.000 |
| C4 |
-1.328 |
1.122 |
0.000 |
| H5 |
-0.630 |
-1.374 |
0.000 |
| H6 |
0.794 |
1.188 |
0.000 |
| H7 |
-2.048 |
0.304 |
0.000 |
| H8 |
-1.512 |
1.731 |
0.886 |
| H9 |
-1.512 |
1.731 |
-0.886 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.2076 | 1.3506 | 2.4713 | 1.1062 | 2.0048 | 2.5597 | 3.1899 | 3.1899 |
O2 | 1.2076 | | 2.2716 | 3.5940 | 2.0320 | 2.4847 | 3.7671 | 4.2390 | 4.2390 | N3 | 1.3506 | 2.2716 | | 1.4382 | 2.0422 | 1.0064 | 2.0645 | 2.1028 | 2.1028 | C4 | 2.4713 | 3.5940 | 1.4382 | | 2.5908 | 2.1226 | 1.0890 | 1.0914 | 1.0914 | H5 | 1.1062 | 2.0320 | 2.0422 | 2.5908 | | 2.9305 | 2.1963 | 3.3470 | 3.3470 | H6 | 2.0048 | 2.4847 | 1.0064 | 2.1226 | 2.9305 | | 2.9754 | 2.5291 | 2.5291 | H7 | 2.5597 | 3.7671 | 2.0645 | 1.0890 | 2.1963 | 2.9754 | | 1.7632 | 1.7632 | H8 | 3.1899 | 4.2390 | 2.1028 | 1.0914 | 3.3470 | 2.5291 | 1.7632 | | 1.7726 | H9 | 3.1899 | 4.2390 | 2.1028 | 1.0914 | 3.3470 | 2.5291 | 1.7632 | 1.7726 | |
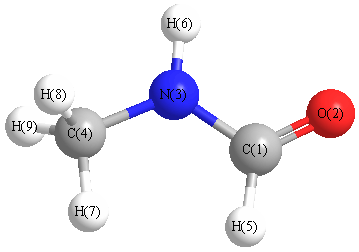 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
C4 |
124.754 |
|
C1 |
N3 |
H6 |
115.779 |
| O2 |
C1 |
N3 |
125.139 |
|
O2 |
C1 |
H5 |
122.794 |
| N3 |
C1 |
H5 |
112.067 |
|
N3 |
C4 |
H7 |
108.767 |
| N3 |
C4 |
H8 |
111.731 |
|
N3 |
C4 |
H9 |
111.731 |
| C4 |
N3 |
H6 |
119.467 |
|
H7 |
C4 |
H8 |
107.932 |
| H7 |
C4 |
H9 |
107.932 |
|
H8 |
C4 |
H9 |
108.605 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.157 |
|
|
|
| 2 |
O |
-0.335 |
|
|
|
| 3 |
N |
-0.102 |
|
|
|
| 4 |
C |
-0.139 |
|
|
|
| 5 |
H |
0.020 |
|
|
|
| 6 |
H |
0.142 |
|
|
|
| 7 |
H |
0.085 |
|
|
|
| 8 |
H |
0.087 |
|
|
|
| 9 |
H |
0.087 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.046 |
2.796 |
0.000 |
4.134 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-26.418 |
4.054 |
0.000 |
| y |
4.054 |
-24.261 |
0.000 |
| z |
0.000 |
0.000 |
-24.604 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.986 |
4.054 |
0.000 |
| y |
4.054 |
1.250 |
0.000 |
| z |
0.000 |
0.000 |
0.736 |
|
| Polar |
|---|
| 3z2-r2 | 1.471 |
|---|
| x2-y2 | -2.158 |
|---|
| xy | 4.054 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.093 |
-1.090 |
0.000 |
| y |
-1.090 |
6.080 |
0.000 |
| z |
0.000 |
0.000 |
3.741 |
<r2> (average value of r
2) Å
2
| <r2> |
88.243 |
| (<r2>)1/2 |
9.394 |
Jump to
S1C1
S1C3
Energy calculated at mPW1PW91/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -209.226091 |
| Energy at 298.15K | -209.232190 |
| HF Energy | -209.226091 |
| Nuclear repulsion energy | 119.857231 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/cc-pVTZ
Geometric Data calculated at mPW1PW91/cc-pVTZ
Point Group is Cs
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at mPW1PW91/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -209.227663 |
| Energy at 298.15K | -209.233752 |
| HF Energy | -209.227663 |
| Nuclear repulsion energy | 122.001800 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3681 |
3531 |
27.62 |
|
|
|
| 2 |
A |
3169 |
3040 |
0.58 |
|
|
|
| 3 |
A |
3109 |
2982 |
26.00 |
|
|
|
| 4 |
A |
3052 |
2927 |
42.90 |
|
|
|
| 5 |
A |
2964 |
2843 |
109.92 |
|
|
|
| 6 |
A |
1816 |
1742 |
344.36 |
|
|
|
| 7 |
A |
1569 |
1505 |
105.57 |
|
|
|
| 8 |
A |
1502 |
1441 |
7.16 |
|
|
|
| 9 |
A |
1496 |
1435 |
26.32 |
|
|
|
| 10 |
A |
1442 |
1383 |
20.13 |
|
|
|
| 11 |
A |
1426 |
1368 |
3.09 |
|
|
|
| 12 |
A |
1239 |
1188 |
71.70 |
|
|
|
| 13 |
A |
1164 |
1116 |
21.09 |
|
|
|
| 14 |
A |
1156 |
1109 |
0.71 |
|
|
|
| 15 |
A |
1033 |
991 |
0.00 |
|
|
|
| 16 |
A |
979 |
939 |
20.76 |
|
|
|
| 17 |
A |
774 |
743 |
0.79 |
|
|
|
| 18 |
A |
547 |
525 |
40.80 |
|
|
|
| 19 |
A |
298 |
286 |
14.02 |
|
|
|
| 20 |
A |
276 |
265 |
65.61 |
|
|
|
| 21 |
A |
79 |
76 |
0.08 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16384.5 cm
-1
Scaled (by 0.9592) Zero Point Vibrational Energy (zpe) 15716.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/cc-pVTZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.860 |
0.433 |
0.004 |
| O2 |
1.367 |
-0.669 |
-0.000 |
| N3 |
-0.468 |
0.654 |
-0.020 |
| C4 |
-1.416 |
-0.438 |
0.005 |
| H5 |
1.452 |
1.361 |
0.018 |
| H6 |
-0.794 |
1.599 |
0.056 |
| H7 |
-2.408 |
-0.051 |
-0.214 |
| H8 |
-1.143 |
-1.174 |
-0.748 |
| H9 |
-1.428 |
-0.933 |
0.976 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.2122 | 1.3466 | 2.4369 | 1.1011 | 2.0248 | 3.3110 | 2.6758 | 2.8367 |
O2 | 1.2122 | | 2.2620 | 2.7927 | 2.0316 | 3.1329 | 3.8314 | 2.6675 | 2.9724 | N3 | 1.3466 | 2.2620 | | 1.4462 | 2.0460 | 1.0025 | 2.0734 | 2.0805 | 2.1058 | C4 | 2.4369 | 2.7927 | 1.4462 | | 3.3853 | 2.1302 | 1.0871 | 1.0878 | 1.0906 | H5 | 1.1011 | 2.0316 | 2.0460 | 3.3853 | | 2.2586 | 4.1165 | 3.7078 | 3.8047 | H6 | 2.0248 | 3.1329 | 1.0025 | 2.1302 | 2.2586 | | 2.3237 | 2.9085 | 2.7678 | H7 | 3.3110 | 3.8314 | 2.0734 | 1.0871 | 4.1165 | 2.3237 | | 1.7743 | 1.7762 | H8 | 2.6758 | 2.6675 | 2.0805 | 1.0878 | 3.7078 | 2.9085 | 1.7743 | | 1.7637 | H9 | 2.8367 | 2.9724 | 2.1058 | 1.0906 | 3.8047 | 2.7678 | 1.7762 | 1.7637 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
C4 |
121.473 |
|
C1 |
N3 |
H6 |
118.329 |
| O2 |
C1 |
N3 |
124.175 |
|
O2 |
C1 |
H5 |
122.787 |
| N3 |
C1 |
H5 |
113.036 |
|
N3 |
C4 |
H7 |
109.037 |
| N3 |
C4 |
H8 |
109.561 |
|
N3 |
C4 |
H9 |
111.446 |
| C4 |
N3 |
H6 |
119.796 |
|
H7 |
C4 |
H8 |
109.340 |
| H7 |
C4 |
H9 |
109.309 |
|
H8 |
C4 |
H9 |
108.120 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.167 |
|
|
|
| 2 |
O |
-0.349 |
|
|
|
| 3 |
N |
-0.124 |
|
|
|
| 4 |
C |
-0.157 |
|
|
|
| 5 |
H |
0.034 |
|
|
|
| 6 |
H |
0.141 |
|
|
|
| 7 |
H |
0.086 |
|
|
|
| 8 |
H |
0.116 |
|
|
|
| 9 |
H |
0.086 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.919 |
2.588 |
-0.001 |
3.901 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.526 |
2.601 |
0.000 |
| y |
2.601 |
-22.110 |
-0.002 |
| z |
0.000 |
-0.002 |
-24.631 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.155 |
2.601 |
0.000 |
| y |
2.601 |
2.968 |
-0.002 |
| z |
0.000 |
-0.002 |
-0.813 |
|
| Polar |
|---|
| 3z2-r2 | -1.626 |
|---|
| x2-y2 | -3.416 |
|---|
| xy | 2.601 |
|---|
| xz | 0.000 |
|---|
| yz | -0.002 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.520 |
-0.046 |
-0.000 |
| y |
-0.046 |
5.439 |
-0.000 |
| z |
-0.000 |
-0.000 |
3.736 |
<r2> (average value of r
2) Å
2
| <r2> |
77.892 |
| (<r2>)1/2 |
8.826 |
