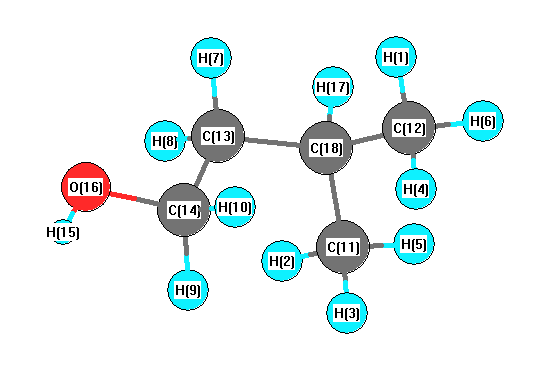
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3890 |
3723 |
30.85 |
|
|
|
| 2 |
A' |
3124 |
2990 |
49.37 |
|
|
|
| 3 |
A' |
3121 |
2987 |
73.65 |
|
|
|
| 4 |
A' |
3056 |
2925 |
29.02 |
|
|
|
| 5 |
A' |
3051 |
2920 |
30.51 |
|
|
|
| 6 |
A' |
3037 |
2907 |
10.02 |
|
|
|
| 7 |
A' |
3006 |
2877 |
42.11 |
|
|
|
| 8 |
A' |
1528 |
1462 |
1.27 |
|
|
|
| 9 |
A' |
1506 |
1442 |
6.64 |
|
|
|
| 10 |
A' |
1504 |
1440 |
12.38 |
|
|
|
| 11 |
A' |
1487 |
1423 |
2.63 |
|
|
|
| 12 |
A' |
1456 |
1393 |
2.78 |
|
|
|
| 13 |
A' |
1416 |
1355 |
4.36 |
|
|
|
| 14 |
A' |
1372 |
1313 |
0.82 |
|
|
|
| 15 |
A' |
1339 |
1281 |
27.88 |
|
|
|
| 16 |
A' |
1245 |
1191 |
25.24 |
|
|
|
| 17 |
A' |
1207 |
1156 |
3.97 |
|
|
|
| 18 |
A' |
1127 |
1079 |
1.36 |
|
|
|
| 19 |
A' |
1085 |
1039 |
106.07 |
|
|
|
| 20 |
A' |
1006 |
962 |
3.83 |
|
|
|
| 21 |
A' |
967 |
926 |
9.34 |
|
|
|
| 22 |
A' |
813 |
778 |
4.52 |
|
|
|
| 23 |
A' |
552 |
528 |
1.22 |
|
|
|
| 24 |
A' |
392 |
375 |
6.75 |
|
|
|
| 25 |
A' |
317 |
304 |
4.24 |
|
|
|
| 26 |
A' |
233 |
223 |
0.20 |
|
|
|
| 27 |
A' |
195 |
186 |
1.61 |
|
|
|
| 28 |
A" |
3122 |
2988 |
18.11 |
|
|
|
| 29 |
A" |
3116 |
2983 |
2.90 |
|
|
|
| 30 |
A" |
3097 |
2964 |
44.80 |
|
|
|
| 31 |
A" |
3049 |
2918 |
53.63 |
|
|
|
| 32 |
A" |
3036 |
2905 |
14.69 |
|
|
|
| 33 |
A" |
1493 |
1429 |
1.03 |
|
|
|
| 34 |
A" |
1483 |
1420 |
0.58 |
|
|
|
| 35 |
A" |
1400 |
1340 |
9.87 |
|
|
|
| 36 |
A" |
1369 |
1310 |
0.45 |
|
|
|
| 37 |
A" |
1318 |
1262 |
0.04 |
|
|
|
| 38 |
A" |
1246 |
1193 |
0.01 |
|
|
|
| 39 |
A" |
1180 |
1129 |
4.25 |
|
|
|
| 40 |
A" |
1011 |
967 |
0.28 |
|
|
|
| 41 |
A" |
964 |
923 |
0.00 |
|
|
|
| 42 |
A" |
923 |
883 |
0.66 |
|
|
|
| 43 |
A" |
789 |
756 |
0.09 |
|
|
|
| 44 |
A" |
354 |
339 |
0.00 |
|
|
|
| 45 |
A" |
281 |
269 |
103.64 |
|
|
|
| 46 |
A" |
219 |
210 |
1.33 |
|
|
|
| 47 |
A" |
96 |
92 |
0.00 |
|
|
|
| 48 |
A" |
45 |
44 |
8.38 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36310.0 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 34752.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.122 |
|
|
|
| 2 |
H |
0.122 |
|
|
|
| 3 |
H |
0.113 |
|
|
|
| 4 |
H |
0.113 |
|
|
|
| 5 |
H |
0.120 |
|
|
|
| 6 |
H |
0.120 |
|
|
|
| 7 |
H |
0.124 |
|
|
|
| 8 |
H |
0.124 |
|
|
|
| 9 |
H |
0.082 |
|
|
|
| 10 |
H |
0.082 |
|
|
|
| 11 |
C |
-0.378 |
|
|
|
| 12 |
C |
-0.378 |
|
|
|
| 13 |
C |
-0.220 |
|
|
|
| 14 |
C |
-0.046 |
|
|
|
| 15 |
H |
0.202 |
|
|
|
| 16 |
O |
-0.336 |
|
|
|
| 17 |
H |
0.123 |
|
|
|
| 18 |
C |
-0.090 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.280 |
-1.340 |
0.000 |
1.369 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-34.899 |
2.763 |
0.000 |
| y |
2.763 |
-43.594 |
0.000 |
| z |
0.000 |
0.000 |
-40.308 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
7.052 |
2.763 |
0.000 |
| y |
2.763 |
-5.990 |
0.000 |
| z |
0.000 |
0.000 |
-1.062 |
|
| Polar |
|---|
| 3z2-r2 | -2.123 |
|---|
| x2-y2 | 8.695 |
|---|
| xy | 2.763 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.257 |
-0.791 |
0.000 |
| y |
-0.791 |
9.420 |
0.000 |
| z |
0.000 |
0.000 |
9.350 |
<r2> (average value of r
2) Å
2
| <r2> |
220.212 |
| (<r2>)1/2 |
14.840 |