return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H8O2 (Acetylacetone)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1A
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-345.247065
Energy at 298.15K-345.255667
HF Energy-343.851609
Nuclear repulsion energy299.312748
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3212 3049 4.13      
2 A 3160 3000 2.22      
3 A 3130 2972 1.65      
4 A 3093 2936 0.28      
5 A 1785 1695 14.51      
6 A 1501 1425 20.35      
7 A 1497 1422 0.20      
8 A 1482 1407 12.22      
9 A 1402 1331 8.03      
10 A 1293 1228 19.28      
11 A 1150 1092 0.82      
12 A 1081 1026 0.25      
13 A 956 908 0.95      
14 A 815 773 0.25      
15 A 634 602 1.49      
16 A 494 469 6.03      
17 A 322 306 0.81      
18 A 182 173 0.02      
19 A 146 139 0.72      
20 A 64 60 6.84      
21 B 3212 3049 4.32      
22 B 3195 3033 2.86      
23 B 3160 3000 0.08      
24 B 3092 2936 1.87      
25 B 1761 1672 221.15      
26 B 1500 1424 1.97      
27 B 1489 1414 27.59      
28 B 1400 1329 98.24      
29 B 1257 1193 168.74      
30 B 1210 1149 66.39      
31 B 1070 1016 2.37      
32 B 1002 951 0.70      
33 B 915 869 13.16      
34 B 828 786 8.17      
35 B 551 523 23.77      
36 B 502 477 1.88      
37 B 417 395 1.84      
38 B 183 174 0.12      
39 B 70 67 11.61      

Unscaled Zero Point Vibrational Energy (zpe) 27106.1 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 25734.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
0.13781 0.06807 0.05337

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is C2

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 1.024
C2 0.000 1.206 0.105
C3 0.000 -1.206 0.105
C4 -1.348 1.713 -0.294
C5 1.348 -1.713 -0.294
O6 1.041 1.675 -0.306
O7 -1.041 -1.675 -0.306
H8 -0.896 -0.029 1.633
H9 0.896 0.029 1.633
H10 -1.256 2.450 -1.081
H11 -1.968 0.880 -0.613
H12 -1.830 2.157 0.573
H13 1.256 -2.450 -1.081
H14 1.968 -0.880 -0.613
H15 1.830 -2.157 0.573

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 O6 O7 H8 H9 H10 H11 H12 H13 H14 H15
C11.51621.51622.54792.54792.37922.37921.08331.08333.46592.70722.86493.46592.70722.8649
C21.51622.41211.49483.24051.21413.09112.15862.12642.12892.12042.11514.04392.95663.8575
C31.51622.41213.24051.49483.09111.21412.12642.15864.04392.95663.85752.12892.12042.1151
C42.54791.49483.24054.36042.38983.40242.63683.40401.08161.08661.08724.97354.22205.0830
C52.54793.24051.49484.36043.40242.38983.40402.63684.97354.22205.08301.08161.08661.0872
O62.37921.21413.09112.38983.40243.94483.22722.54792.54533.12793.04194.20282.73524.0102
O72.37923.09111.21413.40242.38983.94482.54793.22724.20282.73524.01022.54533.12793.0419
H81.08332.15862.12642.63683.40403.22722.54791.79263.69282.64932.60264.22583.73783.6175
H91.08332.12642.15863.40402.63682.54793.22721.79264.22583.73783.61753.69282.64932.6026
H103.46592.12894.04391.08164.97352.54534.20283.69284.22581.78651.77525.50674.65875.7870
H112.70722.12042.95661.08664.22203.12792.73522.64933.73781.78651.74854.65874.31195.0061
H122.86492.11513.85751.08725.08303.04194.01022.60263.61751.77521.74855.78705.00615.6582
H133.46594.04392.12894.97351.08164.20282.54534.22583.69285.50674.65875.78701.78651.7752
H142.70722.95662.12044.22201.08662.73523.12793.73782.64934.65874.31195.00611.78651.7485
H152.86493.85752.11515.08301.08724.01023.04193.61752.60265.78705.00615.65821.77521.7485

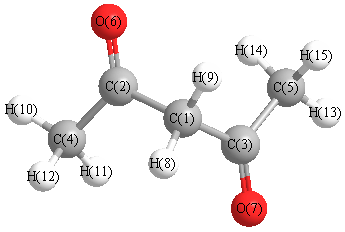
picture of Acetylacetone state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 115.598 C1 C2 O6 120.849
C1 C3 C5 115.598 C1 C3 O7 120.849
C2 C1 C3 105.394 C2 C1 H8 111.184
C2 C1 H9 108.624 C2 C4 H10 110.417
C2 C4 H11 109.434 C2 C4 H12 108.982
C3 C1 H8 108.624 C3 C1 H9 111.184
C3 C5 H13 110.417 C3 C5 H14 109.434
C3 C5 H15 108.982 C4 C2 O6 123.488
C5 C3 O7 123.488 H8 C1 H9 111.664
H10 C4 H11 110.962 H10 C4 H12 109.874
H11 C4 H12 107.094 H13 C5 H14 110.962
H13 C5 H15 109.874 H14 C5 H15 107.094
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability