return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CHOCH2CH2CH3 (Butanal)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-232.063397
Energy at 298.15K-232.072095
HF Energy-231.070967
Nuclear repulsion energy173.181819
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3168 3008 24.26      
2 A' 3114 2956 13.77      
3 A' 3088 2932 20.46      
4 A' 3067 2912 17.86      
5 A' 2962 2812 129.35      
6 A' 1786 1695 105.18      
7 A' 1532 1455 6.49      
8 A' 1513 1437 2.81      
9 A' 1471 1397 12.03      
10 A' 1444 1371 7.05      
11 A' 1428 1356 6.73      
12 A' 1415 1343 13.41      
13 A' 1324 1257 6.42      
14 A' 1160 1101 8.88      
15 A' 1087 1032 0.10      
16 A' 986 936 6.21      
17 A' 881 837 7.89      
18 A' 708 672 13.77      
19 A' 349 331 0.77      
20 A' 195 185 7.80      
21 A" 3161 3001 39.21      
22 A" 3138 2979 0.48      
23 A" 3091 2935 5.48      
24 A" 1530 1453 7.42      
25 A" 1342 1274 0.20      
26 A" 1269 1205 0.07      
27 A" 1169 1110 0.08      
28 A" 967 918 0.04      
29 A" 801 761 1.00      
30 A" 680 646 3.26      
31 A" 265 252 0.01      
32 A" 184 174 1.90      
33 A" 88 84 1.86      

Unscaled Zero Point Vibrational Energy (zpe) 25182.4 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 23908.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
0.50912 0.08667 0.07716

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 2.259 -0.338 0.000
H2 2.790 -1.284 0.000
H3 2.570 0.221 0.878
H4 2.570 0.221 -0.878
C5 0.759 -0.556 0.000
H6 0.463 -1.136 -0.869
H7 0.463 -1.136 0.869
C8 0.000 0.750 0.000
H9 0.260 1.358 0.868
H10 0.260 1.358 -0.868
C11 -1.489 0.587 0.000
H12 -2.053 1.535 0.000
O13 -2.062 -0.474 0.000

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 H6 H7 C8 H9 H10 C11 H12 O13
C11.08501.08631.08631.51592.14822.14822.50752.76122.76123.86054.70084.3228
H21.08501.75611.75612.15812.48802.48803.45333.75963.75964.67065.60374.9193
H31.08631.75611.75552.15783.05462.50612.76732.57493.11084.16954.88574.7656
H41.08631.75611.75552.15782.50613.05462.76733.11082.57494.16954.88574.7656
C51.51592.15812.15782.15781.08551.08551.51092.15972.15972.52183.50362.8217
H62.14822.48803.05462.50611.08551.73732.12833.04582.50232.74543.77102.7515
H72.14822.48802.50613.05461.08551.73732.12832.50233.04582.74543.77102.7515
C82.50753.45332.76732.76731.51092.12832.12831.09061.09061.49822.19752.3981
H92.76123.75962.57493.11082.15973.04582.50231.09061.73582.09932.47673.0822
H102.76123.75963.11082.57492.15972.50233.04581.09061.73582.09932.47673.0822
C113.86054.67064.16954.16952.52182.74542.74541.49822.09932.09931.10261.2057
H124.70085.60374.88574.88573.50363.77103.77102.19752.47672.47671.10262.0088
O134.32284.91934.76564.76562.82172.75152.75152.39813.08223.08221.20572.0088

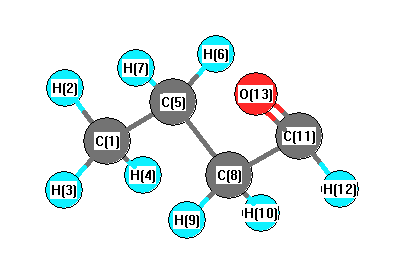
picture of Butanal state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C5 H6 110.236 C1 C5 H7 110.236
C1 C5 C8 111.881 H2 C1 H3 107.953
H2 C1 H4 107.953 H2 C1 C5 111.065
H3 C1 H4 107.806 H3 C1 C5 110.962
H4 C1 C5 110.962 C5 C8 H9 111.205
C5 C8 H10 111.205 C5 C8 C11 113.873
H6 C5 H7 106.298 H6 C5 C8 109.011
H7 C5 C8 109.011 C8 C11 H12 114.469
C8 C11 O13 124.625 H9 C8 H10 105.452
H9 C8 C11 107.326 H10 C8 C11 107.326
H12 C11 O13 120.905
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability