return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H9N (Pyrrolidine)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-212.199169
Energy at 298.15K-212.210712
HF Energy-211.220763
Nuclear repulsion energy191.860252
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3568 3388 0.86      
2 A' 3177 3016 40.97      
3 A' 3145 2986 3.95      
4 A' 3128 2969 34.76      
5 A' 2996 2845 121.38      
6 A' 1548 1470 0.84      
7 A' 1525 1447 6.14      
8 A' 1403 1332 0.99      
9 A' 1332 1264 1.52      
10 A' 1263 1199 2.55      
11 A' 1239 1177 6.24      
12 A' 1091 1035 0.26      
13 A' 1021 969 1.97      
14 A' 967 918 4.96      
15 A' 930 883 2.42      
16 A' 897 852 61.73      
17 A' 775 735 26.18      
18 A' 582 552 40.18      
19 A' 320 303 3.49      
20 A" 3157 2997 2.10      
21 A" 3140 2981 58.61      
22 A" 3117 2960 12.55      
23 A" 2996 2845 32.37      
24 A" 1529 1452 0.12      
25 A" 1504 1428 0.92      
26 A" 1440 1368 6.49      
27 A" 1336 1268 7.84      
28 A" 1315 1249 11.07      
29 A" 1258 1195 6.71      
30 A" 1215 1153 1.27      
31 A" 1153 1094 12.01      
32 A" 1115 1058 0.09      
33 A" 958 910 0.30      
34 A" 880 836 2.48      
35 A" 625 593 0.56      
36 A" 51 49 0.09      

Unscaled Zero Point Vibrational Energy (zpe) 28847.1 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 27387.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
0.23354 0.23081 0.13321

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 0.561 -1.072 0.000
H2 0.469 -2.079 0.000
C3 -0.109 -0.468 1.142
C4 -0.109 -0.468 -1.142
C5 -0.109 1.012 0.771
C6 -0.109 1.012 -0.771
H7 -1.142 -0.818 1.250
H8 -1.142 -0.818 -1.250
H9 0.421 -0.676 2.065
H10 0.421 -0.676 -2.065
H11 0.789 1.488 1.148
H12 0.789 1.488 -1.148
H13 -0.963 1.531 1.189
H14 -0.963 1.531 -1.189

Atom - Atom Distances (Å)
  N1 H2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14
N11.01081.45551.45552.32122.32122.12802.12802.10752.10752.81502.81503.24273.2427
H21.01082.05722.05723.23773.23772.39772.39772.49692.49693.76073.76074.06174.0617
C31.45552.05722.28471.52652.41951.09592.62941.08423.25762.15273.14312.17503.1882
C41.45552.05722.28472.41951.52652.62941.09593.25761.08423.14312.15273.18822.1750
C52.32123.23771.52652.41951.54232.15542.91582.19223.34301.08352.17121.08372.2006
C62.32123.23772.41951.52651.54232.91582.15543.34302.19222.17121.08352.20061.0837
H72.12802.39771.09592.62942.15542.91582.50041.76843.66813.00923.84652.35653.3912
H82.12802.39772.62941.09592.91582.15542.50043.66811.76843.84653.00923.39122.3565
H92.10752.49691.08423.25762.19223.34301.76843.66814.13042.37933.89132.74884.1689
H102.10752.49693.25761.08423.34302.19223.66811.76844.13043.89132.37934.16892.7488
H112.81503.76072.15273.14311.08352.17123.00923.84652.37933.89132.29561.75302.9211
H122.81503.76073.14312.15272.17121.08353.84653.00923.89132.37932.29562.92111.7530
H133.24274.06172.17503.18821.08372.20062.35653.39122.74884.16891.75302.92112.3784
H143.24274.06173.18822.17502.20061.08373.39122.35654.16892.74882.92111.75302.3784

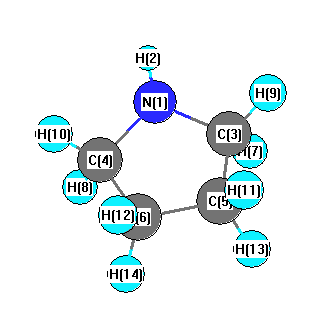
picture of Pyrrolidine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C3 C5 102.205 N1 C3 H7 112.275
N1 C3 H9 111.321 N1 C4 C6 102.205
N1 C4 H8 112.275 N1 C4 H10 111.321
H2 N1 C3 111.785 H2 N1 C4 111.785
C3 N1 C4 103.418 C3 C5 C6 104.075
C3 C5 H11 109.979 C3 C5 H13 111.756
C4 C6 C5 104.075 C4 C6 H12 109.979
C4 C6 H14 111.756 C5 C3 H7 109.462
C5 C3 H9 113.125 C5 C6 H12 110.342
C5 C6 H14 112.691 C6 C4 H8 109.462
C6 C4 H10 113.125 C6 C5 H11 110.342
C6 C5 H13 112.691 H7 C3 H9 108.416
H8 C4 H10 108.416 H11 C5 H13 107.981
H12 C6 H14 107.981
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability