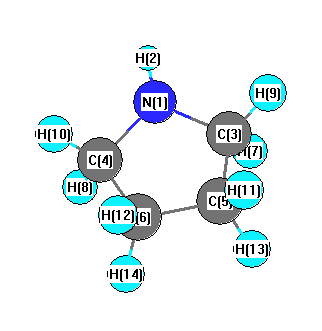
Vibrational Frequencies calculated at mPW1PW91/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3575 |
3429 |
0.22 |
|
|
|
| 2 |
A' |
3138 |
3010 |
54.05 |
|
|
|
| 3 |
A' |
3098 |
2972 |
14.23 |
|
|
|
| 4 |
A' |
3085 |
2959 |
36.61 |
|
|
|
| 5 |
A' |
2948 |
2828 |
141.51 |
|
|
|
| 6 |
A' |
1535 |
1472 |
0.82 |
|
|
|
| 7 |
A' |
1507 |
1446 |
5.83 |
|
|
|
| 8 |
A' |
1403 |
1345 |
0.89 |
|
|
|
| 9 |
A' |
1323 |
1269 |
1.59 |
|
|
|
| 10 |
A' |
1255 |
1204 |
1.47 |
|
|
|
| 11 |
A' |
1232 |
1182 |
6.97 |
|
|
|
| 12 |
A' |
1082 |
1038 |
0.36 |
|
|
|
| 13 |
A' |
1013 |
972 |
1.38 |
|
|
|
| 14 |
A' |
960 |
920 |
7.76 |
|
|
|
| 15 |
A' |
921 |
883 |
1.69 |
|
|
|
| 16 |
A' |
883 |
847 |
54.84 |
|
|
|
| 17 |
A' |
769 |
738 |
25.39 |
|
|
|
| 18 |
A' |
580 |
556 |
48.61 |
|
|
|
| 19 |
A' |
296 |
284 |
3.58 |
|
|
|
| 20 |
A" |
3117 |
2989 |
0.27 |
|
|
|
| 21 |
A" |
3090 |
2964 |
88.22 |
|
|
|
| 22 |
A" |
3078 |
2952 |
4.30 |
|
|
|
| 23 |
A" |
2945 |
2825 |
46.53 |
|
|
|
| 24 |
A" |
1515 |
1453 |
0.25 |
|
|
|
| 25 |
A" |
1488 |
1427 |
0.85 |
|
|
|
| 26 |
A" |
1449 |
1390 |
8.69 |
|
|
|
| 27 |
A" |
1333 |
1279 |
3.76 |
|
|
|
| 28 |
A" |
1315 |
1261 |
18.52 |
|
|
|
| 29 |
A" |
1254 |
1202 |
5.83 |
|
|
|
| 30 |
A" |
1211 |
1162 |
0.50 |
|
|
|
| 31 |
A" |
1151 |
1104 |
13.52 |
|
|
|
| 32 |
A" |
1116 |
1071 |
0.24 |
|
|
|
| 33 |
A" |
949 |
911 |
0.25 |
|
|
|
| 34 |
A" |
878 |
842 |
2.58 |
|
|
|
| 35 |
A" |
641 |
615 |
0.71 |
|
|
|
| 36 |
A" |
84 |
81 |
0.07 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28606.9 cm
-1
Scaled (by 0.9592) Zero Point Vibrational Energy (zpe) 27439.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.217 |
|
|
|
| 2 |
H |
0.098 |
|
|
|
| 3 |
C |
-0.060 |
|
|
|
| 4 |
C |
-0.060 |
|
|
|
| 5 |
C |
-0.151 |
|
|
|
| 6 |
C |
-0.151 |
|
|
|
| 7 |
H |
0.048 |
|
|
|
| 8 |
H |
0.048 |
|
|
|
| 9 |
H |
0.067 |
|
|
|
| 10 |
H |
0.067 |
|
|
|
| 11 |
H |
0.079 |
|
|
|
| 12 |
H |
0.079 |
|
|
|
| 13 |
H |
0.076 |
|
|
|
| 14 |
H |
0.076 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.837 |
-0.256 |
0.000 |
0.875 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.938 |
0.724 |
0.000 |
| y |
0.724 |
-30.396 |
0.000 |
| z |
0.000 |
0.000 |
-31.632 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.924 |
0.724 |
0.000 |
| y |
0.724 |
2.389 |
0.000 |
| z |
0.000 |
0.000 |
0.535 |
|
| Polar |
|---|
| 3z2-r2 | 1.070 |
|---|
| x2-y2 | -3.542 |
|---|
| xy | 0.724 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.096 |
-0.084 |
0.000 |
| y |
-0.084 |
8.061 |
0.000 |
| z |
0.000 |
0.000 |
8.482 |
<r2> (average value of r
2) Å
2
| <r2> |
108.254 |
| (<r2>)1/2 |
10.404 |