All results from a given calculation for C6H12 ((E)-hex-3-ene)
using model chemistry: MP2=FULL/cc-pVTZ
19 10 17 12 22
States and conformations
| State |
Conformation |
minimum conformation |
conformer description |
state description |
| 1 |
1 |
yes |
CI |
1AG |
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -235.391826 |
| Energy at 298.15K | |
| Nuclear repulsion energy | |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is Ci
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-0.666 |
| C2 |
0.000 |
0.000 |
0.666 |
| C3 |
1.218 |
0.000 |
-1.521 |
| C4 |
-1.218 |
0.000 |
1.521 |
| C5 |
1.324 |
-1.268 |
-2.357 |
| C6 |
-1.324 |
1.268 |
2.357 |
| H7 |
-0.952 |
-0.025 |
-1.179 |
| H8 |
0.952 |
0.025 |
1.179 |
| H9 |
1.197 |
0.862 |
-2.187 |
| H10 |
2.100 |
0.106 |
-0.893 |
| H11 |
-1.197 |
-0.862 |
2.187 |
| H12 |
-2.100 |
-0.106 |
0.893 |
| H13 |
2.200 |
-1.247 |
-2.998 |
| H14 |
1.389 |
-2.141 |
-1.715 |
| H15 |
0.448 |
-1.385 |
-2.989 |
| H16 |
-2.200 |
1.247 |
2.998 |
| H17 |
-1.389 |
2.141 |
1.715 |
| H18 |
-0.448 |
1.385 |
2.989 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
| C1 | | 1.3312 | 1.4889 | 2.5034 | 2.4945 | 3.5353 | 1.0820 | 2.0761 | 2.1191 | 2.1148 | 3.2114 | 2.6174 | 3.4401 | 2.7593 | 2.7420 | 4.4516 | 3.4900 | 3.9341 |
C2 | 1.3312 | | 2.5034 | 1.4889 | 3.5353 | 2.4945 | 2.0761 | 1.0820 | 3.2114 | 2.6174 | 2.1191 | 2.1148 | 4.4516 | 3.4900 | 3.9341 | 3.4401 | 2.7593 | 2.7420 | C3 | 1.4889 | 2.5034 | | 3.8981 | 1.5226 | 4.8077 | 2.1974 | 2.7137 | 1.0891 | 1.0873 | 4.5088 | 4.1052 | 2.1677 | 2.1563 | 2.1605 | 5.8021 | 4.6750 | 5.0040 | C4 | 2.5034 | 1.4889 | 3.8981 | | 4.8077 | 1.5226 | 2.7137 | 2.1974 | 4.5088 | 4.1052 | 1.0891 | 1.0873 | 5.8021 | 4.6750 | 5.0040 | 2.1677 | 2.1563 | 2.1605 | C5 | 2.4945 | 3.5353 | 1.5226 | 4.8077 | | 5.9724 | 2.8483 | 3.7838 | 2.1404 | 2.1527 | 5.2126 | 4.8619 | 1.0856 | 1.0854 | 1.0862 | 6.8864 | 5.9636 | 6.2261 | C6 | 3.5353 | 2.4945 | 4.8077 | 1.5226 | 5.9724 | | 3.7838 | 2.8483 | 5.2126 | 4.8619 | 2.1404 | 2.1527 | 6.8864 | 5.9636 | 6.2261 | 1.0856 | 1.0854 | 1.0862 | H7 | 1.0820 | 2.0761 | 2.1974 | 2.7137 | 2.8483 | 3.7838 | | 3.0315 | 2.5340 | 3.0680 | 3.4773 | 2.3705 | 3.8387 | 3.2008 | 2.6621 | 4.5414 | 3.6411 | 4.4292 | H8 | 2.0761 | 1.0820 | 2.7137 | 2.1974 | 3.7838 | 2.8483 | 3.0315 | | 3.4773 | 2.3705 | 2.5340 | 3.0680 | 4.5414 | 3.6411 | 4.4292 | 3.8387 | 3.2008 | 2.6621 | H9 | 2.1191 | 3.2114 | 1.0891 | 4.5088 | 2.1404 | 5.2126 | 2.5340 | 3.4773 | | 1.7490 | 5.2759 | 4.6147 | 2.4716 | 3.0453 | 2.5008 | 6.2108 | 4.8530 | 5.4565 | H10 | 2.1148 | 2.6174 | 1.0873 | 4.1052 | 2.1527 | 4.8619 | 3.0680 | 2.3705 | 1.7490 | | 4.6147 | 4.5688 | 2.5042 | 2.4957 | 3.0570 | 5.9103 | 4.8079 | 4.8168 | H11 | 3.2114 | 2.1191 | 4.5088 | 1.0891 | 5.2126 | 2.1404 | 3.4773 | 2.5340 | 5.2759 | 4.6147 | | 1.7490 | 6.2108 | 4.8530 | 5.4565 | 2.4716 | 3.0453 | 2.5008 | H12 | 2.6174 | 2.1148 | 4.1052 | 1.0873 | 4.8619 | 2.1527 | 2.3705 | 3.0680 | 4.6147 | 4.5688 | 1.7490 | | 5.9103 | 4.8079 | 4.8168 | 2.5042 | 2.4957 | 3.0570 | H13 | 3.4401 | 4.4516 | 2.1677 | 5.8021 | 1.0856 | 6.8864 | 3.8387 | 4.5414 | 2.4716 | 2.5042 | 6.2108 | 5.9103 | | 1.7614 | 1.7572 | 7.8441 | 6.8243 | 7.0560 | H14 | 2.7593 | 3.4900 | 2.1563 | 4.6750 | 1.0854 | 5.9636 | 3.2008 | 3.6411 | 3.0453 | 2.4957 | 4.8530 | 4.8079 | 1.7614 | | 1.7549 | 6.8243 | 6.1494 | 6.1594 | H15 | 2.7420 | 3.9341 | 2.1605 | 5.0040 | 1.0862 | 6.2261 | 2.6621 | 4.4292 | 2.5008 | 3.0570 | 5.4565 | 4.8168 | 1.7572 | 1.7549 | | 7.0560 | 6.1594 | 6.6497 | H16 | 4.4516 | 3.4401 | 5.8021 | 2.1677 | 6.8864 | 1.0856 | 4.5414 | 3.8387 | 6.2108 | 5.9103 | 2.4716 | 2.5042 | 7.8441 | 6.8243 | 7.0560 | | 1.7614 | 1.7572 | H17 | 3.4900 | 2.7593 | 4.6750 | 2.1563 | 5.9636 | 1.0854 | 3.6411 | 3.2008 | 4.8530 | 4.8079 | 3.0453 | 2.4957 | 6.8243 | 6.1494 | 6.1594 | 1.7614 | | 1.7549 | H18 | 3.9341 | 2.7420 | 5.0040 | 2.1605 | 6.2261 | 1.0862 | 4.4292 | 2.6621 | 5.4565 | 4.8168 | 2.5008 | 3.0570 | 7.0560 | 6.1594 | 6.6497 | 1.7572 | 1.7549 | |
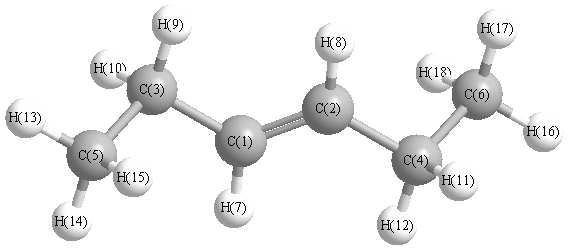 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C4 |
125.077 |
|
C1 |
C2 |
H8 |
118.336 |
| C1 |
C3 |
C5 |
111.849 |
|
C1 |
C3 |
H9 |
109.596 |
| C1 |
C3 |
H10 |
109.360 |
|
C2 |
C1 |
C3 |
125.077 |
| C2 |
C1 |
H7 |
118.336 |
|
C2 |
C4 |
C6 |
111.849 |
| C2 |
C4 |
H11 |
109.596 |
|
C2 |
C4 |
H12 |
109.360 |
| C3 |
C1 |
H7 |
116.570 |
|
C3 |
C5 |
H13 |
111.324 |
| C3 |
C5 |
H14 |
110.415 |
|
C3 |
C5 |
H15 |
110.701 |
| C4 |
C2 |
H8 |
116.570 |
|
C4 |
C6 |
H16 |
111.324 |
| C4 |
C6 |
H17 |
110.415 |
|
C4 |
C6 |
H18 |
110.701 |
| C5 |
C3 |
H9 |
108.943 |
|
C5 |
C3 |
H10 |
110.017 |
| C6 |
C4 |
H11 |
108.943 |
|
C6 |
C4 |
H12 |
110.017 |
| H9 |
C3 |
H10 |
106.952 |
|
H11 |
C4 |
H12 |
106.952 |
| H13 |
C5 |
H14 |
108.452 |
|
H13 |
C5 |
H15 |
108.017 |
| H14 |
C5 |
H15 |
107.816 |
|
H16 |
C6 |
H17 |
108.452 |
| H16 |
C6 |
H18 |
108.017 |
|
H17 |
C6 |
H18 |
107.816 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
