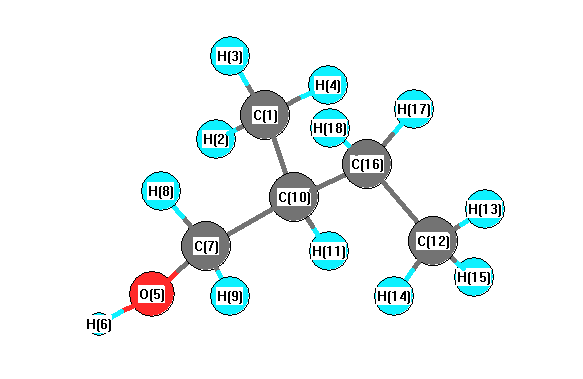
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3893 |
3726 |
35.02 |
|
|
|
| 2 |
A |
3148 |
3013 |
16.65 |
|
|
|
| 3 |
A |
3131 |
2996 |
43.68 |
|
|
|
| 4 |
A |
3124 |
2990 |
38.72 |
|
|
|
| 5 |
A |
3115 |
2981 |
36.09 |
|
|
|
| 6 |
A |
3076 |
2944 |
19.34 |
|
|
|
| 7 |
A |
3057 |
2926 |
64.64 |
|
|
|
| 8 |
A |
3050 |
2919 |
68.38 |
|
|
|
| 9 |
A |
3045 |
2915 |
1.14 |
|
|
|
| 10 |
A |
3037 |
2907 |
19.99 |
|
|
|
| 11 |
A |
3029 |
2899 |
10.67 |
|
|
|
| 12 |
A |
2985 |
2857 |
48.51 |
|
|
|
| 13 |
A |
1519 |
1454 |
2.48 |
|
|
|
| 14 |
A |
1509 |
1444 |
7.10 |
|
|
|
| 15 |
A |
1505 |
1440 |
6.19 |
|
|
|
| 16 |
A |
1499 |
1435 |
7.44 |
|
|
|
| 17 |
A |
1492 |
1428 |
6.63 |
|
|
|
| 18 |
A |
1483 |
1420 |
1.36 |
|
|
|
| 19 |
A |
1456 |
1393 |
3.18 |
|
|
|
| 20 |
A |
1412 |
1351 |
3.29 |
|
|
|
| 21 |
A |
1401 |
1341 |
8.02 |
|
|
|
| 22 |
A |
1389 |
1329 |
0.32 |
|
|
|
| 23 |
A |
1363 |
1304 |
3.23 |
|
|
|
| 24 |
A |
1326 |
1269 |
1.30 |
|
|
|
| 25 |
A |
1299 |
1244 |
10.26 |
|
|
|
| 26 |
A |
1274 |
1219 |
5.07 |
|
|
|
| 27 |
A |
1247 |
1193 |
24.34 |
|
|
|
| 28 |
A |
1191 |
1140 |
2.21 |
|
|
|
| 29 |
A |
1175 |
1125 |
4.53 |
|
|
|
| 30 |
A |
1133 |
1085 |
1.13 |
|
|
|
| 31 |
A |
1095 |
1048 |
92.10 |
|
|
|
| 32 |
A |
1059 |
1013 |
4.56 |
|
|
|
| 33 |
A |
1025 |
981 |
15.68 |
|
|
|
| 34 |
A |
974 |
933 |
5.38 |
|
|
|
| 35 |
A |
925 |
886 |
1.49 |
|
|
|
| 36 |
A |
924 |
884 |
1.55 |
|
|
|
| 37 |
A |
832 |
796 |
3.85 |
|
|
|
| 38 |
A |
763 |
730 |
4.59 |
|
|
|
| 39 |
A |
492 |
471 |
6.65 |
|
|
|
| 40 |
A |
436 |
417 |
1.53 |
|
|
|
| 41 |
A |
377 |
360 |
1.58 |
|
|
|
| 42 |
A |
267 |
256 |
5.00 |
|
|
|
| 43 |
A |
255 |
244 |
23.72 |
|
|
|
| 44 |
A |
235 |
225 |
55.77 |
|
|
|
| 45 |
A |
228 |
218 |
11.04 |
|
|
|
| 46 |
A |
216 |
207 |
9.68 |
|
|
|
| 47 |
A |
117 |
112 |
4.64 |
|
|
|
| 48 |
A |
70 |
67 |
4.20 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36324.2 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 34765.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.371 |
|
|
|
| 2 |
H |
0.130 |
|
|
|
| 3 |
H |
0.106 |
|
|
|
| 4 |
H |
0.112 |
|
|
|
| 5 |
O |
-0.336 |
|
|
|
| 6 |
H |
0.204 |
|
|
|
| 7 |
C |
-0.040 |
|
|
|
| 8 |
H |
0.077 |
|
|
|
| 9 |
H |
0.089 |
|
|
|
| 10 |
C |
-0.075 |
|
|
|
| 11 |
H |
0.119 |
|
|
|
| 12 |
C |
-0.374 |
|
|
|
| 13 |
H |
0.121 |
|
|
|
| 14 |
H |
0.112 |
|
|
|
| 15 |
H |
0.117 |
|
|
|
| 16 |
C |
-0.220 |
|
|
|
| 17 |
H |
0.116 |
|
|
|
| 18 |
H |
0.112 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.424 |
-0.944 |
0.947 |
1.403 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.025 |
3.103 |
-2.408 |
| y |
3.103 |
-37.712 |
-1.566 |
| z |
-2.408 |
-1.566 |
-39.995 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.171 |
3.103 |
-2.408 |
| y |
3.103 |
2.298 |
-1.566 |
| z |
-2.408 |
-1.566 |
-1.127 |
|
| Polar |
|---|
| 3z2-r2 | -2.254 |
|---|
| x2-y2 | -2.313 |
|---|
| xy | 3.103 |
|---|
| xz | -2.408 |
|---|
| yz | -1.566 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.684 |
-0.158 |
-0.036 |
| y |
-0.158 |
9.871 |
-0.072 |
| z |
-0.036 |
-0.072 |
8.632 |
<r2> (average value of r
2) Å
2
| <r2> |
213.068 |
| (<r2>)1/2 |
14.597 |