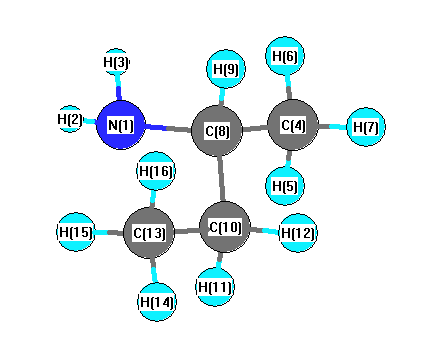
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3611 |
3456 |
0.24 |
|
|
|
| 2 |
A |
3531 |
3379 |
0.53 |
|
|
|
| 3 |
A |
3135 |
3001 |
34.35 |
|
|
|
| 4 |
A |
3126 |
2992 |
35.53 |
|
|
|
| 5 |
A |
3119 |
2985 |
32.41 |
|
|
|
| 6 |
A |
3118 |
2985 |
40.03 |
|
|
|
| 7 |
A |
3087 |
2954 |
8.58 |
|
|
|
| 8 |
A |
3051 |
2920 |
41.03 |
|
|
|
| 9 |
A |
3048 |
2917 |
22.90 |
|
|
|
| 10 |
A |
3046 |
2915 |
19.79 |
|
|
|
| 11 |
A |
2942 |
2815 |
69.04 |
|
|
|
| 12 |
A |
1640 |
1569 |
40.76 |
|
|
|
| 13 |
A |
1506 |
1442 |
7.48 |
|
|
|
| 14 |
A |
1498 |
1434 |
3.29 |
|
|
|
| 15 |
A |
1494 |
1430 |
7.46 |
|
|
|
| 16 |
A |
1488 |
1425 |
3.78 |
|
|
|
| 17 |
A |
1474 |
1411 |
0.92 |
|
|
|
| 18 |
A |
1422 |
1361 |
6.48 |
|
|
|
| 19 |
A |
1402 |
1342 |
13.00 |
|
|
|
| 20 |
A |
1399 |
1339 |
4.93 |
|
|
|
| 21 |
A |
1381 |
1322 |
10.98 |
|
|
|
| 22 |
A |
1332 |
1275 |
2.25 |
|
|
|
| 23 |
A |
1298 |
1242 |
1.46 |
|
|
|
| 24 |
A |
1256 |
1202 |
1.09 |
|
|
|
| 25 |
A |
1201 |
1149 |
2.36 |
|
|
|
| 26 |
A |
1165 |
1115 |
9.90 |
|
|
|
| 27 |
A |
1074 |
1028 |
0.91 |
|
|
|
| 28 |
A |
1036 |
992 |
3.42 |
|
|
|
| 29 |
A |
1012 |
969 |
4.49 |
|
|
|
| 30 |
A |
980 |
938 |
2.65 |
|
|
|
| 31 |
A |
949 |
908 |
8.18 |
|
|
|
| 32 |
A |
858 |
821 |
106.22 |
|
|
|
| 33 |
A |
817 |
782 |
5.85 |
|
|
|
| 34 |
A |
767 |
734 |
2.50 |
|
|
|
| 35 |
A |
471 |
451 |
6.37 |
|
|
|
| 36 |
A |
449 |
429 |
6.04 |
|
|
|
| 37 |
A |
361 |
346 |
3.64 |
|
|
|
| 38 |
A |
265 |
253 |
19.38 |
|
|
|
| 39 |
A |
227 |
217 |
1.11 |
|
|
|
| 40 |
A |
220 |
210 |
2.29 |
|
|
|
| 41 |
A |
199 |
191 |
22.75 |
|
|
|
| 42 |
A |
98 |
94 |
0.74 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 32774.7 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 31368.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.348 |
|
|
|
| 2 |
H |
0.144 |
|
|
|
| 3 |
H |
0.139 |
|
|
|
| 4 |
C |
-0.360 |
|
|
|
| 5 |
H |
0.121 |
|
|
|
| 6 |
H |
0.107 |
|
|
|
| 7 |
H |
0.108 |
|
|
|
| 8 |
C |
-0.008 |
|
|
|
| 9 |
H |
0.101 |
|
|
|
| 10 |
C |
-0.188 |
|
|
|
| 11 |
H |
0.115 |
|
|
|
| 12 |
H |
0.107 |
|
|
|
| 13 |
C |
-0.389 |
|
|
|
| 14 |
H |
0.122 |
|
|
|
| 15 |
H |
0.115 |
|
|
|
| 16 |
H |
0.115 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.002 |
0.152 |
1.139 |
1.149 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.268 |
0.473 |
0.919 |
| y |
0.473 |
-32.446 |
1.842 |
| z |
0.919 |
1.842 |
-35.520 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.715 |
0.473 |
0.919 |
| y |
0.473 |
1.947 |
1.842 |
| z |
0.919 |
1.842 |
-2.662 |
|
| Polar |
|---|
| 3z2-r2 | -5.324 |
|---|
| x2-y2 | -0.822 |
|---|
| xy | 0.473 |
|---|
| xz | 0.919 |
|---|
| yz | 1.842 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.723 |
-0.072 |
-0.004 |
| y |
-0.072 |
8.584 |
0.035 |
| z |
-0.004 |
0.035 |
7.669 |
<r2> (average value of r
2) Å
2
| <r2> |
148.745 |
| (<r2>)1/2 |
12.196 |